return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Methyl cyclopropyl ketone)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-269.703865
Energy at 298.15K-269.712578
HF Energy-268.832180
Nuclear repulsion energy229.976888
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3273 3117 7.56      
2 A' 3198 3045 9.43      
3 A' 3187 3035 8.39      
4 A' 3179 3028 9.75      
5 A' 3067 2920 3.60      
6 A' 1806 1720 98.28      
7 A' 1550 1476 11.44      
8 A' 1519 1447 11.63      
9 A' 1467 1397 99.36      
10 A' 1431 1363 6.43      
11 A' 1259 1199 9.26      
12 A' 1233 1174 74.68      
13 A' 1141 1086 10.92      
14 A' 1109 1057 9.81      
15 A' 1007 959 35.50      
16 A' 951 906 13.21      
17 A' 830 790 1.48      
18 A' 772 735 0.13      
19 A' 600 571 13.25      
20 A' 379 361 3.91      
21 A' 248 236 4.45      
22 A" 3261 3106 0.40      
23 A" 3175 3024 12.98      
24 A" 3135 2986 9.77      
25 A" 1524 1451 9.62      
26 A" 1501 1430 2.86      
27 A" 1243 1184 0.16      
28 A" 1161 1106 2.18      
29 A" 1130 1076 1.99      
30 A" 1083 1031 3.47      
31 A" 924 880 0.27      
32 A" 864 823 5.33      
33 A" 611 582 2.22      
34 A" 268 255 0.51      
35 A" 142 136 0.02      
36 A" 71 68 1.80      

Unscaled Zero Point Vibrational Energy (zpe) 26649.7 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 25378.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.23928 0.08751 0.07647

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.484 -0.551 0.000
C2 -0.276 -0.759 0.000
C3 0.298 -2.160 0.000
C4 0.703 0.372 0.000
C5 0.298 1.630 0.746
C6 0.298 1.630 -0.746
H7 -0.514 -2.891 0.000
H8 0.932 -2.307 -0.885
H9 0.932 -2.307 0.885
H10 1.758 0.105 0.000
H11 -0.665 1.593 1.247
H12 1.080 2.173 1.272
H13 -0.665 1.593 -1.247
H14 1.080 2.173 -1.272

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.22612.40152.37342.91362.91362.53343.11523.11523.30782.61153.95092.61153.9509
C21.22611.51421.49552.56782.56782.14552.15362.15362.20972.69043.47152.69043.4715
C32.40151.51422.56423.86303.86301.09321.09821.09822.69474.07064.58294.07064.5829
C42.37341.49552.56421.51751.51753.48282.83082.83081.08882.21762.23672.21762.2367
C52.91362.56783.86301.51751.49154.65404.30853.99042.23921.08701.08742.21392.2308
C62.91362.56783.86301.51751.49154.65403.99044.30852.23922.21392.23081.08701.0874
H72.53342.14551.09323.48284.65404.65401.79331.79333.76054.65675.45934.65675.4593
H83.11522.15361.09822.83084.30853.99041.79331.76982.69894.72314.97434.23014.4991
H93.11522.15361.09822.83083.99044.30851.79331.76982.69894.23014.49914.72314.9743
H103.30782.20972.69471.08882.23922.23923.76052.69892.69893.10552.52073.10552.5207
H112.61152.69044.07062.21761.08702.21394.65674.72314.23013.10551.83962.49403.1190
H123.95093.47154.58292.23671.08742.23085.45934.97434.49912.52071.83963.11902.5437
H132.61152.69044.07062.21762.21391.08704.65674.23014.72313.10552.49403.11901.8396
H143.95093.47154.58292.23672.23081.08745.45934.49914.97432.52073.11902.54371.8396

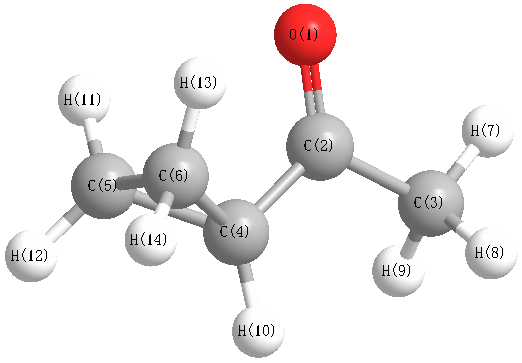
picture of Methyl cyclopropyl ketone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 122.060 O1 C2 C4 121.081
C2 C3 H7 109.688 C2 C3 H8 110.035
C2 C3 H9 110.035 C2 C4 C5 116.917
C2 C4 C6 116.917 C2 C4 H10 116.658
C3 C2 C4 116.859 C4 C5 C6 60.564
C4 C5 H11 115.754 C4 C5 H12 117.378
C4 C6 C5 60.564 C4 C6 H13 115.754
C4 C6 H14 117.378 C5 C4 C6 58.872
C5 C4 H10 117.504 C5 C6 H13 117.458
C5 C6 H14 118.934 C6 C4 H10 117.504
C6 C5 H11 117.458 C6 C5 H12 118.934
H7 C3 H8 109.838 H7 C3 H9 109.838
H8 C3 H9 107.376 H11 C5 H12 115.566
H13 C6 H14 115.566
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability