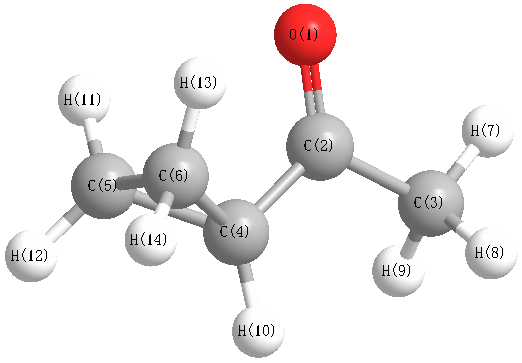
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3186 |
3133 |
5.80 |
|
|
|
| 2 |
A' |
3112 |
3060 |
12.80 |
|
|
|
| 3 |
A' |
3108 |
3057 |
7.86 |
|
|
|
| 4 |
A' |
3088 |
3037 |
10.32 |
|
|
|
| 5 |
A' |
2981 |
2932 |
2.80 |
|
|
|
| 6 |
A' |
1736 |
1708 |
104.62 |
|
|
|
| 7 |
A' |
1461 |
1437 |
9.03 |
|
|
|
| 8 |
A' |
1445 |
1421 |
13.55 |
|
|
|
| 9 |
A' |
1382 |
1359 |
109.87 |
|
|
|
| 10 |
A' |
1347 |
1325 |
11.66 |
|
|
|
| 11 |
A' |
1209 |
1189 |
3.23 |
|
|
|
| 12 |
A' |
1167 |
1148 |
91.37 |
|
|
|
| 13 |
A' |
1084 |
1066 |
17.42 |
|
|
|
| 14 |
A' |
1028 |
1011 |
11.42 |
|
|
|
| 15 |
A' |
948 |
932 |
54.40 |
|
|
|
| 16 |
A' |
916 |
901 |
4.94 |
|
|
|
| 17 |
A' |
790 |
777 |
3.73 |
|
|
|
| 18 |
A' |
738 |
726 |
0.15 |
|
|
|
| 19 |
A' |
576 |
566 |
7.58 |
|
|
|
| 20 |
A' |
364 |
358 |
4.21 |
|
|
|
| 21 |
A' |
235 |
231 |
4.25 |
|
|
|
| 22 |
A" |
3172 |
3120 |
0.48 |
|
|
|
| 23 |
A" |
3088 |
3037 |
14.59 |
|
|
|
| 24 |
A" |
3045 |
2995 |
7.45 |
|
|
|
| 25 |
A" |
1454 |
1430 |
10.08 |
|
|
|
| 26 |
A" |
1424 |
1401 |
4.01 |
|
|
|
| 27 |
A" |
1184 |
1164 |
0.11 |
|
|
|
| 28 |
A" |
1095 |
1077 |
1.84 |
|
|
|
| 29 |
A" |
1050 |
1033 |
1.23 |
|
|
|
| 30 |
A" |
1019 |
1002 |
6.98 |
|
|
|
| 31 |
A" |
853 |
839 |
0.02 |
|
|
|
| 32 |
A" |
820 |
806 |
3.90 |
|
|
|
| 33 |
A" |
585 |
575 |
0.74 |
|
|
|
| 34 |
A" |
247 |
243 |
0.21 |
|
|
|
| 35 |
A" |
130 |
127 |
0.23 |
|
|
|
| 36 |
A" |
73 |
72 |
2.04 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25568.4 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 25146.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.435 |
|
|
|
| 2 |
C |
0.444 |
|
|
|
| 3 |
C |
-0.582 |
|
|
|
| 4 |
C |
-0.216 |
|
|
|
| 5 |
C |
-0.321 |
|
|
|
| 6 |
C |
-0.321 |
|
|
|
| 7 |
H |
0.190 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.183 |
|
|
|
| 10 |
H |
0.151 |
|
|
|
| 11 |
H |
0.192 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.192 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.670 |
0.052 |
0.000 |
2.671 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.799 |
-0.851 |
0.000 |
| y |
-0.851 |
-33.787 |
0.000 |
| z |
0.000 |
0.000 |
-35.538 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.137 |
-0.851 |
0.000 |
| y |
-0.851 |
3.381 |
0.000 |
| z |
0.000 |
0.000 |
0.755 |
|
| Polar |
|---|
| 3z2-r2 | 1.511 |
|---|
| x2-y2 | -5.012 |
|---|
| xy | -0.851 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.875 |
0.257 |
0.000 |
| y |
0.257 |
9.621 |
0.000 |
| z |
0.000 |
0.000 |
6.586 |
<r2> (average value of r
2) Å
2
| <r2> |
169.420 |
| (<r2>)1/2 |
13.016 |