Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -235.029252 |
| Energy at 298.15K | -235.041495 |
| HF Energy | -234.175316 |
| Nuclear repulsion energy | 229.498097 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3173 |
3022 |
63.99 |
|
|
|
| 2 |
A |
3160 |
3009 |
5.31 |
|
|
|
| 3 |
A |
3144 |
2994 |
16.50 |
|
|
|
| 4 |
A |
3135 |
2986 |
38.13 |
|
|
|
| 5 |
A |
3132 |
2982 |
54.03 |
|
|
|
| 6 |
A |
3114 |
2965 |
24.51 |
|
|
|
| 7 |
A |
3109 |
2961 |
17.51 |
|
|
|
| 8 |
A |
3088 |
2940 |
6.64 |
|
|
|
| 9 |
A |
3059 |
2913 |
21.54 |
|
|
|
| 10 |
A |
3058 |
2912 |
29.28 |
|
|
|
| 11 |
A |
3054 |
2908 |
35.64 |
|
|
|
| 12 |
A |
3041 |
2896 |
18.15 |
|
|
|
| 13 |
A |
1770 |
1686 |
0.90 |
|
|
|
| 14 |
A |
1561 |
1486 |
5.68 |
|
|
|
| 15 |
A |
1550 |
1476 |
4.05 |
|
|
|
| 16 |
A |
1549 |
1475 |
6.41 |
|
|
|
| 17 |
A |
1544 |
1470 |
0.55 |
|
|
|
| 18 |
A |
1537 |
1464 |
1.73 |
|
|
|
| 19 |
A |
1535 |
1462 |
3.46 |
|
|
|
| 20 |
A |
1483 |
1412 |
6.49 |
|
|
|
| 21 |
A |
1466 |
1396 |
3.00 |
|
|
|
| 22 |
A |
1456 |
1387 |
2.85 |
|
|
|
| 23 |
A |
1427 |
1359 |
0.46 |
|
|
|
| 24 |
A |
1358 |
1293 |
0.09 |
|
|
|
| 25 |
A |
1333 |
1269 |
2.92 |
|
|
|
| 26 |
A |
1329 |
1266 |
1.29 |
|
|
|
| 27 |
A |
1292 |
1230 |
0.11 |
|
|
|
| 28 |
A |
1201 |
1144 |
0.70 |
|
|
|
| 29 |
A |
1148 |
1093 |
0.94 |
|
|
|
| 30 |
A |
1095 |
1043 |
1.96 |
|
|
|
| 31 |
A |
1093 |
1041 |
0.17 |
|
|
|
| 32 |
A |
1091 |
1039 |
0.85 |
|
|
|
| 33 |
A |
1033 |
983 |
4.12 |
|
|
|
| 34 |
A |
998 |
951 |
0.24 |
|
|
|
| 35 |
A |
940 |
896 |
2.52 |
|
|
|
| 36 |
A |
924 |
880 |
4.58 |
|
|
|
| 37 |
A |
894 |
851 |
3.48 |
|
|
|
| 38 |
A |
761 |
725 |
1.89 |
|
|
|
| 39 |
A |
729 |
694 |
29.02 |
|
|
|
| 40 |
A |
575 |
548 |
6.26 |
|
|
|
| 41 |
A |
474 |
451 |
0.67 |
|
|
|
| 42 |
A |
372 |
354 |
0.01 |
|
|
|
| 43 |
A |
297 |
283 |
0.03 |
|
|
|
| 44 |
A |
249 |
237 |
0.00 |
|
|
|
| 45 |
A |
202 |
192 |
0.01 |
|
|
|
| 46 |
A |
110 |
105 |
0.21 |
|
|
|
| 47 |
A |
93 |
88 |
0.00 |
|
|
|
| 48 |
A |
41 |
39 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36887.8 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 35128.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.483 |
0.951 |
-0.071 |
| C2 |
1.998 |
-0.473 |
-0.144 |
| C3 |
0.769 |
-0.939 |
0.132 |
| C4 |
-0.444 |
-0.158 |
0.565 |
| C5 |
-1.582 |
-0.236 |
-0.463 |
| C6 |
-2.839 |
0.507 |
-0.007 |
| H7 |
1.695 |
1.652 |
0.222 |
| H8 |
3.305 |
1.044 |
0.652 |
| H9 |
2.876 |
1.274 |
-1.045 |
| H10 |
2.753 |
-1.198 |
-0.457 |
| H11 |
0.602 |
-2.014 |
0.017 |
| H12 |
-0.192 |
0.895 |
0.747 |
| H13 |
-0.806 |
-0.558 |
1.526 |
| H14 |
-1.228 |
0.178 |
-1.418 |
| H15 |
-1.825 |
-1.291 |
-0.656 |
| H16 |
-3.638 |
0.441 |
-0.757 |
| H17 |
-3.224 |
0.088 |
0.932 |
| H18 |
-2.627 |
1.571 |
0.164 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5060 | 2.5598 | 3.1939 | 4.2529 | 5.3412 | 1.0943 | 1.0988 | 1.0987 | 2.1999 | 3.5126 | 2.7983 | 3.9558 | 4.0227 | 4.8916 | 6.1800 | 5.8585 | 5.1526 |
C2 | 1.5060 | | 1.3429 | 2.5620 | 3.6021 | 4.9373 | 2.1767 | 2.1546 | 2.1531 | 1.0925 | 2.0856 | 2.7316 | 3.2647 | 3.5290 | 3.9428 | 5.7419 | 5.3608 | 5.0653 | C3 | 2.5598 | 1.3429 | | 1.5068 | 2.5257 | 3.8902 | 2.7529 | 3.2609 | 3.2749 | 2.0851 | 1.0935 | 2.1608 | 2.1380 | 2.7643 | 2.7339 | 4.7028 | 4.2001 | 4.2231 | C4 | 3.1939 | 2.5620 | 1.5068 | | 1.5358 | 2.5509 | 2.8224 | 3.9374 | 3.9579 | 3.5135 | 2.2000 | 1.0977 | 1.1021 | 2.1589 | 2.1640 | 3.5080 | 2.8149 | 2.8128 | C5 | 4.2529 | 3.6021 | 2.5257 | 1.5358 | | 1.5295 | 3.8433 | 5.1732 | 4.7428 | 4.4404 | 2.8573 | 2.1620 | 2.1590 | 1.0995 | 1.1004 | 2.1837 | 2.1786 | 2.1787 | C6 | 5.3412 | 4.9373 | 3.8902 | 2.5509 | 1.5295 | | 4.6823 | 6.2025 | 5.8593 | 5.8633 | 4.2659 | 2.7795 | 2.7598 | 2.1670 | 2.1643 | 1.0973 | 1.0981 | 1.0981 | H7 | 1.0943 | 2.1767 | 2.7529 | 2.8224 | 3.8433 | 4.6823 | | 1.7731 | 1.7730 | 3.1142 | 3.8305 | 2.1000 | 3.5830 | 3.6613 | 4.6713 | 5.5554 | 5.2102 | 4.3230 | H8 | 1.0988 | 2.1546 | 3.2609 | 3.9374 | 5.1732 | 6.2025 | 1.7731 | | 1.7659 | 2.5617 | 4.1304 | 3.5014 | 4.4977 | 5.0577 | 5.7859 | 7.1094 | 6.6041 | 5.9745 | H9 | 1.0987 | 2.1531 | 3.2749 | 3.9579 | 4.7428 | 5.8593 | 1.7730 | 1.7659 | | 2.5440 | 4.1366 | 3.5737 | 4.8504 | 4.2641 | 5.3694 | 6.5731 | 6.5213 | 5.6420 | H10 | 2.1999 | 1.0925 | 2.0851 | 3.5135 | 4.4404 | 5.8633 | 3.1142 | 2.5617 | 2.5440 | | 2.3487 | 3.8084 | 4.1241 | 4.3200 | 4.5827 | 6.6038 | 6.2691 | 6.0818 | H11 | 3.5126 | 2.0856 | 1.0935 | 2.2000 | 2.8573 | 4.2659 | 3.8305 | 4.1304 | 4.1366 | 2.3487 | | 3.1028 | 2.5260 | 3.1960 | 2.6200 | 4.9597 | 4.4600 | 4.8265 | H12 | 2.7983 | 2.7316 | 2.1608 | 1.0977 | 2.1620 | 2.7795 | 2.1000 | 3.5014 | 3.5737 | 3.8084 | 3.1028 | | 1.7592 | 2.5047 | 3.0682 | 3.7865 | 3.1427 | 2.5925 | H13 | 3.9558 | 3.2647 | 2.1380 | 1.1021 | 2.1590 | 2.7598 | 3.5830 | 4.4977 | 4.8504 | 4.1241 | 2.5260 | 1.7592 | | 3.0638 | 2.5170 | 3.7715 | 2.5717 | 3.1140 | H14 | 4.0227 | 3.5290 | 2.7643 | 2.1589 | 1.0995 | 2.1670 | 3.6613 | 5.0577 | 4.2641 | 4.3200 | 3.1960 | 2.5047 | 3.0638 | | 1.7599 | 2.5125 | 3.0848 | 2.5298 | H15 | 4.8916 | 3.9428 | 2.7339 | 2.1640 | 1.1004 | 2.1643 | 4.6713 | 5.7859 | 5.3694 | 4.5827 | 2.6200 | 3.0682 | 2.5170 | 1.7599 | | 2.5093 | 2.5261 | 3.0833 | H16 | 6.1800 | 5.7419 | 4.7028 | 3.5080 | 2.1837 | 1.0973 | 5.5554 | 7.1094 | 6.5731 | 6.6038 | 4.9597 | 3.7865 | 3.7715 | 2.5125 | 2.5093 | | 1.7745 | 1.7741 | H17 | 5.8585 | 5.3608 | 4.2001 | 2.8149 | 2.1786 | 1.0981 | 5.2102 | 6.6041 | 6.5213 | 6.2691 | 4.4600 | 3.1427 | 2.5717 | 3.0848 | 2.5261 | 1.7745 | | 1.7734 | H18 | 5.1526 | 5.0653 | 4.2231 | 2.8128 | 2.1787 | 1.0981 | 4.3230 | 5.9745 | 5.6420 | 6.0818 | 4.8265 | 2.5925 | 3.1140 | 2.5298 | 3.0833 | 1.7741 | 1.7734 | |
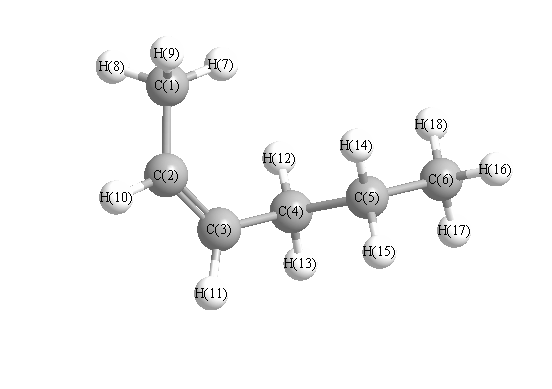 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.844 |
|
C1 |
C2 |
H10 |
114.753 |
| C2 |
C1 |
H7 |
112.708 |
|
C2 |
C1 |
H8 |
110.643 |
| C2 |
C1 |
H9 |
110.530 |
|
C2 |
C3 |
C4 |
127.970 |
| C2 |
C3 |
H11 |
117.384 |
|
C3 |
C2 |
H10 |
117.403 |
| C3 |
C4 |
C5 |
112.215 |
|
C3 |
C4 |
H12 |
111.154 |
| C3 |
C4 |
H13 |
109.084 |
|
C4 |
C3 |
H11 |
114.636 |
| C4 |
C5 |
C6 |
112.645 |
|
C4 |
C5 |
H14 |
108.880 |
| C4 |
C5 |
H15 |
109.232 |
|
C5 |
C4 |
H12 |
109.228 |
| C5 |
C4 |
H13 |
108.747 |
|
C5 |
C6 |
H16 |
111.405 |
| C5 |
C6 |
H17 |
110.946 |
|
C5 |
C6 |
H18 |
110.952 |
| C6 |
C5 |
H14 |
109.942 |
|
C6 |
C5 |
H15 |
109.681 |
| H7 |
C1 |
H8 |
107.898 |
|
H7 |
C1 |
H9 |
107.892 |
| H8 |
C1 |
H9 |
106.946 |
|
H12 |
C4 |
H13 |
106.208 |
| H14 |
C5 |
H16 |
94.003 |
|
H16 |
C6 |
H17 |
107.861 |
| H16 |
C6 |
H18 |
107.822 |
|
H17 |
C6 |
H18 |
107.695 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
