return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for Si3H8 (trisilane)

using model chemistry: MP4/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP4/6-31G*
 hartrees
Energy at 0K-871.688708
Energy at 298.15K-871.696371
HF Energy-871.385648
Nuclear repulsion energy192.261846
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 2266 2164        
2 A1 2250 2149        
3 A1 2228 2127        
4 A1 962 919        
5 A1 938 896        
6 A1 906 865        
7 A1 581 555        
8 A1 402 384        
9 A1 100 95        
10 A2 2262 2160        
11 A2 952 909        
12 A2 725 692        
13 A2 428 409        
14 A2 77 73        
15 B1 2266 2164        
16 B1 2240 2139        
17 B1 955 912        
18 B1 614 586        
19 B1 318 304        
20 B1 97 93        
21 B2 2265 2162        
22 B2 2246 2144        
23 B2 954 911        
24 B2 897 857        
25 B2 746 712        
26 B2 485 463        
27 B2 448 427        

Unscaled Zero Point Vibrational Energy (zpe) 14803.2 cm-1
Scaled (by 0.9548) Zero Point Vibrational Energy (zpe) 14134.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/6-31G*
ABC
0.31180 0.06574 0.05758

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/6-31G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.887
Si2 0.000 1.945 -0.417
Si3 0.000 -1.945 -0.417
H4 1.208 0.000 1.772
H5 -1.208 0.000 1.772
H6 0.000 3.166 0.443
H7 0.000 -3.166 0.443
H8 1.211 1.977 -1.292
H9 -1.211 1.977 -1.292
H10 -1.211 -1.977 -1.292
H11 1.211 -1.977 -1.292

Atom - Atom Distances (Å)
  Si1 Si2 Si3 H4 H5 H6 H7 H8 H9 H10 H11
Si12.34182.34181.49751.49753.19673.19673.18153.18153.18153.1815
Si22.34183.88953.16803.16801.49335.18221.49371.49374.19654.1965
Si32.34183.88953.16803.16805.18221.49334.19654.19651.49371.4937
H41.49753.16803.16802.41573.63993.63993.64664.37574.37573.6466
H51.49753.16803.16802.41573.63993.63994.37573.64663.64664.3757
H63.19671.49335.18223.63993.63996.33122.42612.42615.56065.5606
H73.19675.18221.49333.63993.63996.33125.56065.56062.42612.4261
H83.18151.49374.19653.64664.37572.42615.56062.42154.63673.9542
H93.18151.49374.19654.37573.64662.42615.56062.42153.95424.6367
H103.18154.19651.49374.37573.64665.56062.42614.63673.95422.4215
H113.18154.19651.49373.64664.37575.56062.42613.95424.63672.4215

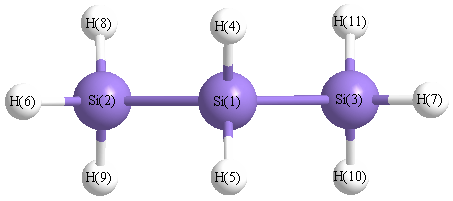
picture of trisilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 H6 110.986 S1 S2 H8 110.123
S1 S2 H9 110.123 S1 S3 H7 110.986
S1 S3 H10 110.123 S1 S3 H11 110.123
S2 S1 S3 112.290 S2 S1 H4 109.227
S2 S1 H5 109.227 S3 S1 H4 109.227
S3 S1 H5 109.227 H4 S1 H5 107.525
H6 S2 H8 108.621 H6 S2 H9 108.621
H7 S3 H10 108.621 H7 S3 H11 108.621
H8 S2 H9 108.299 H10 S3 H11 108.299
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability