return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Butane, 2-methyl-)

using model chemistry: B3PW91/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B3PW91/6-31G*
 hartrees
Energy at 0K-197.701314
Energy at 298.15K-197.693143
Nuclear repulsion energy 
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-31G*
Rotational Constants (cm-1) from geometry optimized at B3PW91/6-31G*
ABC
0.24342 0.11157 0.08529

See section I.F.4 to change rotational constant units
Geometric Data calculated at B3PW91/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.936 -0.597 0.305
H2 -2.214 -0.018 -1.203
H3 -2.185 0.999 0.245
C4 -2.094 -0.033 -0.112
H5 -0.770 -1.724 -0.003
H6 -0.662 -0.652 1.383
C7 -0.761 -0.664 0.287
H8 0.348 -0.003 -1.424
C9 0.481 -0.003 -0.330
H10 -0.196 2.077 -0.163
H11 1.554 1.894 -0.324
H12 0.764 1.508 1.213
C13 0.656 1.450 0.121
H14 2.627 -0.380 -0.477
H15 1.645 -1.851 -0.364
H16 1.915 -0.852 1.073
C17 1.737 -0.817 -0.010

Atom - Atom Distances (Å)
  H1 H2 H3 C4 H5 H6 C7 H8 C9 H10 H11 H12 C13 H14 H15 H16 C17
H11.76921.76501.09602.46062.51722.17573.75863.52583.85705.17294.35244.13795.62184.79664.91764.6883
H21.76921.76981.09732.53653.08182.17882.57192.83293.08904.31584.12703.48474.90864.35434.78784.2038
H31.76501.76981.09563.07852.51792.18933.19472.90522.29873.88633.14492.87865.05754.81304.57344.3292
C41.09601.09731.09562.15052.16111.52772.77242.58442.83824.13093.50673.13264.74794.16574.25983.9116
H52.46062.53653.07852.15051.75581.09932.49612.15313.84754.31213.77873.48173.68412.44573.02162.6660
H62.51723.08182.51792.16111.75581.10053.05272.15913.17103.78212.59362.78293.78823.13242.60272.7783
C72.17572.17882.18931.52771.09931.10052.14341.53662.83453.50362.81072.55003.48502.76142.79552.5202
H83.75862.57193.19472.77242.49613.05272.14341.10172.49242.50243.06712.14312.49622.49433.06752.1426
C93.52582.83292.90522.58442.15312.15911.53661.10172.19342.17892.17761.53092.18402.18512.17861.5309
H103.85703.08902.29872.83823.84753.17102.83452.49242.19341.76701.77181.09543.75594.34313.81593.4838
H115.17294.31583.88634.13094.31213.78213.50362.50242.17891.76701.77041.09662.51923.74643.10152.7355
H124.35244.12703.14493.50673.77872.59362.81073.06712.17761.77181.77041.09833.14523.81392.62882.8013
C134.13793.48472.87863.13263.48172.78292.55002.14311.53091.09541.09661.09832.75573.48022.79072.5152
H145.62184.90865.05754.74793.68413.78823.48502.49622.18403.75592.51923.14522.75571.77211.77031.0965
H154.79664.35434.81304.16572.44573.13242.76142.49432.18514.34313.74643.81393.48021.77211.77131.0967
H164.91764.78784.57344.25983.02162.60272.79553.06752.17863.81593.10152.62882.79071.77031.77131.0981
C174.68834.20384.32923.91162.66602.77832.52022.14261.53093.48382.73552.80132.51521.09651.09671.0981

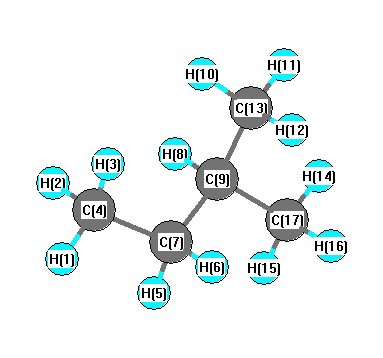
picture of Butane, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H2 107.531 H1 C4 H3 107.282
H1 C4 C7 110.968 H2 C4 H3 107.615
H2 C4 C7 111.139 H3 C4 C7 112.090
C4 C7 H5 108.802 C4 C7 H6 109.549
C4 C7 C9 114.997 H5 C7 H6 105.906
H5 C7 C9 108.403 H6 C7 C9 108.793
C7 C9 H8 107.521 C7 C9 C13 112.464
C7 C9 C17 110.490 H8 C9 C13 107.875
H8 C9 C17 107.839 C9 C13 H10 112.201
C9 C13 H11 110.971 C9 C13 H12 110.762
C9 C17 H14 111.381 C9 C17 H15 111.459
C9 C17 H16 110.854 H10 C13 H11 107.437
H10 C13 H12 107.738 H11 C13 H12 107.531
C13 C9 C17 110.465 H14 C17 H15 107.804
H14 C17 H16 107.543 H15 C17 H16 107.617
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1       0.086
2       0.071
3       0.083
4       -0.317
5       0.015
6       0.019
7       0.003
8       -0.018
9       0.357
10       0.096
11       0.102
12       0.109
13       -0.435
14       0.140
15       0.117
16       0.126
17       -0.551


Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section VII.A.3)
  x y z Total
  -0.026 -0.027 -0.049 0.061
CHELPG        
AIM        
ESP -0.028 -0.001 -0.030 0.041


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000