All results from a given calculation for C5H12 (Butane, 2-methyl-)
using model chemistry: QCISD/6-31G*
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C1 |
1A |
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -197.061504 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.091 |
-0.034 |
-0.107 |
| C2 |
-0.756 |
-0.676 |
0.279 |
| C3 |
0.482 |
-0.003 |
-0.335 |
| C4 |
0.640 |
1.451 |
0.122 |
| C5 |
1.744 |
-0.806 |
-0.005 |
| H6 |
-2.933 |
-0.617 |
0.289 |
| H7 |
-2.201 |
0.014 |
-1.199 |
| H8 |
-2.183 |
0.986 |
0.285 |
| H9 |
-0.766 |
-1.733 |
-0.029 |
| H10 |
-0.649 |
-0.678 |
1.376 |
| H11 |
0.354 |
-0.004 |
-1.430 |
| H12 |
-0.211 |
2.073 |
-0.179 |
| H13 |
1.546 |
1.898 |
-0.307 |
| H14 |
0.726 |
1.502 |
1.217 |
| H15 |
2.633 |
-0.360 |
-0.471 |
| H16 |
1.661 |
-1.842 |
-0.358 |
| H17 |
1.912 |
-0.832 |
1.081 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.5308 | 2.5827 | 3.1168 | 3.9134 | 1.0974 | 1.0985 | 1.0964 | 2.1556 | 2.1658 | 2.7798 | 2.8248 | 4.1233 | 3.4713 | 4.7487 | 4.1721 | 4.2512 |
C2 | 1.5308 | | 1.5362 | 2.5488 | 2.5194 | 2.1778 | 2.1788 | 2.1905 | 1.1006 | 1.1019 | 2.1452 | 2.8399 | 3.5023 | 2.7967 | 3.4847 | 2.7576 | 2.7903 | C3 | 2.5827 | 1.5362 | | 1.5325 | 1.5322 | 3.5245 | 2.8182 | 2.9088 | 2.1541 | 2.1586 | 1.1031 | 2.1945 | 2.1789 | 2.1756 | 2.1846 | 2.1850 | 2.1766 | C4 | 3.1168 | 2.5488 | 1.5325 | | 2.5164 | 4.1310 | 3.4466 | 2.8653 | 3.4834 | 2.7870 | 2.1472 | 1.0963 | 1.0979 | 1.0996 | 2.7576 | 3.4816 | 2.7843 | C5 | 3.9134 | 2.5194 | 1.5322 | 2.5164 | | 4.6900 | 4.2024 | 4.3264 | 2.6756 | 2.7663 | 2.1465 | 3.4851 | 2.7279 | 2.8034 | 1.0979 | 1.0979 | 1.0992 | H6 | 1.0974 | 2.1778 | 3.5245 | 4.1310 | 4.6900 | | 1.7736 | 1.7690 | 2.4580 | 2.5296 | 3.7587 | 3.8549 | 5.1708 | 4.3289 | 5.6227 | 4.7978 | 4.9140 | H7 | 1.0985 | 2.1788 | 2.8182 | 3.4466 | 4.2024 | 1.7736 | | 1.7738 | 2.5449 | 3.0844 | 2.5648 | 3.0396 | 4.2878 | 4.0767 | 4.9020 | 4.3662 | 4.7780 | H8 | 1.0964 | 2.1905 | 2.9088 | 2.8653 | 4.3264 | 1.7690 | 1.7738 | | 3.0816 | 2.5120 | 3.2180 | 2.2990 | 3.8845 | 3.0983 | 5.0568 | 4.8151 | 4.5507 | H9 | 2.1556 | 1.1006 | 2.1541 | 3.4834 | 2.6756 | 2.4580 | 2.5449 | 3.0816 | | 1.7604 | 2.4902 | 3.8489 | 4.3130 | 3.7741 | 3.6916 | 2.4509 | 3.0356 | H10 | 2.1658 | 1.1019 | 2.1586 | 2.7870 | 2.7663 | 2.5296 | 3.0844 | 2.5120 | 1.7604 | | 3.0548 | 3.1907 | 3.7799 | 2.5831 | 3.7794 | 3.1138 | 2.5831 | H11 | 2.7798 | 2.1452 | 1.1031 | 2.1472 | 2.1465 | 3.7587 | 2.5648 | 3.2180 | 2.4902 | 3.0548 | | 2.4903 | 2.5107 | 3.0688 | 2.4979 | 2.4974 | 3.0692 | H12 | 2.8248 | 2.8399 | 2.1945 | 1.0963 | 3.4851 | 3.8549 | 3.0396 | 2.2990 | 3.8489 | 3.1907 | 2.4903 | | 1.7710 | 1.7761 | 3.7542 | 4.3440 | 3.8130 | H13 | 4.1233 | 3.5023 | 2.1789 | 1.0979 | 2.7279 | 5.1708 | 4.2878 | 3.8845 | 4.3130 | 3.7799 | 2.5107 | 1.7710 | | 1.7751 | 2.5109 | 3.7425 | 3.0841 | H14 | 3.4713 | 2.7967 | 2.1756 | 1.0996 | 2.8034 | 4.3289 | 4.0767 | 3.0983 | 3.7741 | 2.5831 | 3.0688 | 1.7761 | 1.7751 | | 3.1548 | 3.8133 | 2.6219 | H15 | 4.7487 | 3.4847 | 2.1846 | 2.7576 | 1.0979 | 5.6227 | 4.9020 | 5.0568 | 3.6916 | 3.7794 | 2.4979 | 3.7542 | 2.5109 | 3.1548 | | 1.7765 | 1.7751 | H16 | 4.1721 | 2.7576 | 2.1850 | 3.4816 | 1.0979 | 4.7978 | 4.3662 | 4.8151 | 2.4509 | 3.1138 | 2.4974 | 4.3440 | 3.7425 | 3.8133 | 1.7765 | | 1.7759 | H17 | 4.2512 | 2.7903 | 2.1766 | 2.7843 | 1.0992 | 4.9140 | 4.7780 | 4.5507 | 3.0356 | 2.5831 | 3.0692 | 3.8130 | 3.0841 | 2.6219 | 1.7751 | 1.7759 | |
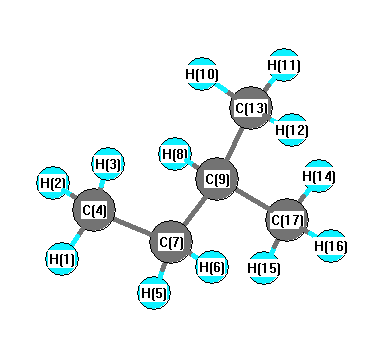 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| H1 |
C4 |
H2 |
29.213 |
|
H1 |
C4 |
H3 |
55.633 |
| H1 |
C4 |
C7 |
18.397 |
|
H2 |
C4 |
H3 |
33.887 |
| H2 |
C4 |
C7 |
39.135 |
|
H3 |
C4 |
C7 |
53.490 |
| C4 |
C7 |
H5 |
36.761 |
|
C4 |
C7 |
H6 |
99.608 |
| C4 |
C7 |
C9 |
69.229 |
|
H5 |
C7 |
H6 |
94.579 |
| H5 |
C7 |
C9 |
37.463 |
|
H6 |
C7 |
C9 |
66.635 |
| C7 |
C9 |
H8 |
35.138 |
|
C7 |
C9 |
C13 |
72.249 |
| C7 |
C9 |
C17 |
117.515 |
|
H8 |
C9 |
C13 |
60.700 |
| H8 |
C9 |
C17 |
96.130 |
|
C9 |
C13 |
H10 |
23.982 |
| C9 |
C13 |
H11 |
30.270 |
|
C9 |
C13 |
H12 |
63.049 |
| C9 |
C17 |
H14 |
83.341 |
|
C9 |
C17 |
H15 |
96.722 |
| C9 |
C17 |
H16 |
53.839 |
|
H10 |
C13 |
H11 |
53.616 |
| H10 |
C13 |
H12 |
57.247 |
|
H11 |
C13 |
H12 |
68.645 |
| C13 |
C9 |
C17 |
45.649 |
|
H14 |
C17 |
H15 |
89.550 |
| H14 |
C17 |
H16 |
118.985 |
|
H15 |
C17 |
H16 |
60.036 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
