return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12 (Butane, 2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-197.490820
Energy at 298.15K-197.503763
HF Energy-197.273203
Nuclear repulsion energy192.194960
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3161 3000 35.20      
2 A 3152 2992 58.08      
3 A 3146 2986 59.74      
4 A 3146 2986 28.08      
5 A 3141 2981 49.90      
6 A 3136 2976 3.89      
7 A 3092 2934 25.32      
8 A 3078 2921 19.28      
9 A 3072 2916 30.25      
10 A 3067 2911 29.46      
11 A 3053 2898 32.36      
12 A 3039 2885 11.18      
13 A 1565 1485 3.46      
14 A 1558 1479 8.82      
15 A 1555 1476 2.28      
16 A 1551 1472 9.99      
17 A 1546 1468 1.27      
18 A 1539 1461 0.19      
19 A 1532 1454 0.19      
20 A 1467 1392 3.91      
21 A 1459 1385 2.90      
22 A 1449 1375 4.76      
23 A 1424 1352 0.26      
24 A 1405 1334 2.27      
25 A 1359 1290 3.64      
26 A 1329 1262 1.52      
27 A 1237 1174 0.17      
28 A 1225 1163 1.45      
29 A 1198 1137 2.33      
30 A 1074 1020 0.13      
31 A 1050 997 2.31      
32 A 1007 955 3.71      
33 A 989 939 0.21      
34 A 951 902 0.93      
35 A 939 891 0.51      
36 A 816 775 0.51      
37 A 781 741 1.91      
38 A 462 439 0.21      
39 A 421 400 0.17      
40 A 373 354 0.00      
41 A 273 259 0.00      
42 A 267 253 0.01      
43 A 231 220 0.01      
44 A 220 209 0.03      
45 A 97 92 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 35814.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 33995.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.24363 0.11167 0.08535

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.929 -0.609 0.299
H2 -2.206 -0.005 -1.197
H3 -2.178 0.987 0.267
C4 -2.090 -0.037 -0.109
H5 -0.764 -1.731 -0.022
H6 -0.652 -0.673 1.374
C7 -0.755 -0.677 0.281
H8 0.353 -0.004 -1.425
C9 0.482 -0.003 -0.334
H10 -0.221 2.066 -0.168
H11 1.531 1.904 -0.317
H12 0.736 1.506 1.213
C13 0.636 1.452 0.122
H14 2.631 -0.357 -0.471
H15 1.666 -1.837 -0.360
H16 1.917 -0.834 1.076
C17 1.747 -0.805 -0.006

Atom - Atom Distances (Å)
  H1 H2 H3 C4 H5 H6 C7 H8 C9 H10 H11 H12 C13 H14 H15 H16 C17
H11.76791.76431.09422.45952.51922.17533.75603.52163.83505.15674.32884.12215.61894.80174.91364.6899
H21.76791.76811.09522.53763.07812.17782.56922.82353.04794.28844.09203.45604.90444.36524.78134.2055
H31.76431.76811.09353.07672.51152.18873.20082.90032.27623.86503.10652.85565.04714.81044.55394.3225
C41.09421.09521.09352.15322.16151.53112.77532.58212.81414.11433.48013.11514.74614.17314.25453.9146
H52.45952.53763.07672.15321.75591.09712.48902.15263.83824.30913.77503.48033.68992.45553.03342.6763
H62.51923.07812.51152.16151.75591.09823.04832.15643.17223.77692.58782.78223.77933.12032.59162.7707
C72.17532.17782.18871.53111.09711.09822.14221.53642.82983.49922.80222.54783.48322.76022.79262.5214
H83.75602.56923.20082.77532.48903.04832.14221.09912.48932.50213.06342.14382.49532.49413.06482.1443
C93.52162.82352.90032.58212.15262.15641.53641.09912.19152.17712.17531.53272.18262.18362.17681.5330
H103.83503.04792.27622.81413.83823.17222.82982.48932.19151.76631.77031.09363.75484.34003.81173.4842
H115.15674.28843.86504.11434.30913.77693.49922.50212.17711.76631.76931.09472.51923.74433.09602.7350
H124.32884.09203.10653.48013.77502.58782.80223.06342.17531.77031.76931.09623.14623.81012.62482.8011
C134.12213.45602.85563.11513.48032.78222.54782.14381.53271.09361.09471.09622.75773.48052.78862.5185
H145.61894.90445.04714.74613.68993.77933.48322.49532.18263.75482.51923.14622.75771.77081.76921.0947
H154.80174.36524.81044.17312.45553.12032.76022.49412.18364.34003.74433.81013.48051.77081.77001.0948
H164.91364.78134.55394.25453.03342.59162.79263.06482.17683.81173.09602.62482.78861.76921.77001.0959
C174.68994.20554.32253.91462.67632.77072.52142.14431.53303.48422.73502.80112.51851.09471.09481.0959

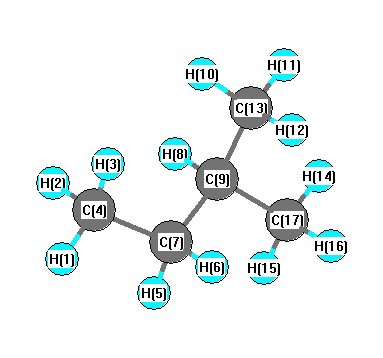
picture of Butane, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 C4 H2 107.703 H1 C4 H3 107.501
H1 C4 C7 110.810 H2 C4 H3 107.772
H2 C4 C7 110.948 H3 C4 C7 111.924
C4 C7 H5 108.905 C4 C7 H6 109.488
C4 C7 C9 114.653 H5 C7 H6 106.235
H5 C7 C9 108.497 H6 C7 C9 108.728
C7 C9 H8 107.586 C7 C9 C13 112.228
C7 C9 C17 110.463 H8 C9 C13 107.951
H8 C9 C17 107.975 C9 C13 H10 112.034
C9 C13 H11 110.806 C9 C13 H12 110.584
C9 C17 H14 111.228 C9 C17 H15 111.298
C9 C17 H16 110.692 H10 C13 H11 107.635
H10 C13 H12 107.889 H11 C13 H12 107.724
C13 C9 C17 110.469 H14 C17 H15 107.950
H14 C17 H16 107.727 H15 C17 H16 107.790
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 H 0.151      
2 H 0.148      
3 H 0.153      
4 C -0.466      
5 H 0.140      
6 H 0.137      
7 C -0.265      
8 H 0.133      
9 C -0.099      
10 H 0.153      
11 H 0.148      
12 H 0.144      
13 C -0.461      
14 H 0.148      
15 H 0.148      
16 H 0.145      
17 C -0.457      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.643 0.038 -0.029
y 0.038 7.856 -0.005
z -0.029 -0.005 7.266


<r2> (average value of r2) Å2
<r2> 157.873
(<r2>)1/2 12.565