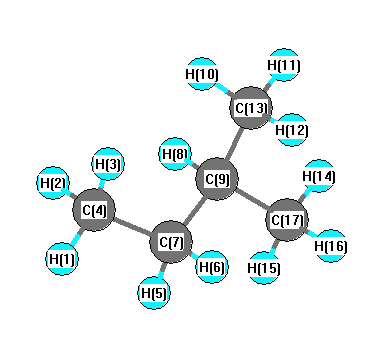
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3151 |
2997 |
30.38 |
|
|
|
| 2 |
A |
3145 |
2991 |
62.11 |
|
|
|
| 3 |
A |
3142 |
2988 |
17.97 |
|
|
|
| 4 |
A |
3140 |
2986 |
54.23 |
|
|
|
| 5 |
A |
3135 |
2981 |
50.08 |
|
|
|
| 6 |
A |
3130 |
2977 |
5.48 |
|
|
|
| 7 |
A |
3083 |
2932 |
24.04 |
|
|
|
| 8 |
A |
3066 |
2916 |
21.64 |
|
|
|
| 9 |
A |
3061 |
2911 |
31.68 |
|
|
|
| 10 |
A |
3056 |
2907 |
31.10 |
|
|
|
| 11 |
A |
3043 |
2894 |
33.83 |
|
|
|
| 12 |
A |
3030 |
2881 |
9.14 |
|
|
|
| 13 |
A |
1541 |
1466 |
4.99 |
|
|
|
| 14 |
A |
1535 |
1460 |
8.39 |
|
|
|
| 15 |
A |
1532 |
1457 |
5.45 |
|
|
|
| 16 |
A |
1527 |
1452 |
12.28 |
|
|
|
| 17 |
A |
1523 |
1448 |
1.82 |
|
|
|
| 18 |
A |
1514 |
1440 |
0.32 |
|
|
|
| 19 |
A |
1506 |
1433 |
0.13 |
|
|
|
| 20 |
A |
1444 |
1374 |
6.17 |
|
|
|
| 21 |
A |
1436 |
1366 |
3.73 |
|
|
|
| 22 |
A |
1429 |
1359 |
8.06 |
|
|
|
| 23 |
A |
1407 |
1338 |
0.77 |
|
|
|
| 24 |
A |
1386 |
1318 |
1.19 |
|
|
|
| 25 |
A |
1339 |
1273 |
2.38 |
|
|
|
| 26 |
A |
1311 |
1247 |
1.53 |
|
|
|
| 27 |
A |
1220 |
1161 |
0.70 |
|
|
|
| 28 |
A |
1214 |
1155 |
0.77 |
|
|
|
| 29 |
A |
1190 |
1131 |
2.67 |
|
|
|
| 30 |
A |
1078 |
1026 |
0.07 |
|
|
|
| 31 |
A |
1043 |
992 |
3.22 |
|
|
|
| 32 |
A |
994 |
945 |
5.03 |
|
|
|
| 33 |
A |
980 |
932 |
0.42 |
|
|
|
| 34 |
A |
940 |
894 |
0.37 |
|
|
|
| 35 |
A |
937 |
891 |
1.66 |
|
|
|
| 36 |
A |
811 |
771 |
0.27 |
|
|
|
| 37 |
A |
777 |
739 |
2.93 |
|
|
|
| 38 |
A |
458 |
436 |
0.27 |
|
|
|
| 39 |
A |
416 |
396 |
0.24 |
|
|
|
| 40 |
A |
369 |
351 |
0.00 |
|
|
|
| 41 |
A |
267 |
254 |
0.00 |
|
|
|
| 42 |
A |
266 |
253 |
0.02 |
|
|
|
| 43 |
A |
229 |
217 |
0.02 |
|
|
|
| 44 |
A |
216 |
205 |
0.04 |
|
|
|
| 45 |
A |
97 |
92 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35555.2 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 33813.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.169 |
|
|
|
| 2 |
H |
0.166 |
|
|
|
| 3 |
H |
0.170 |
|
|
|
| 4 |
C |
-0.519 |
|
|
|
| 5 |
H |
0.157 |
|
|
|
| 6 |
H |
0.154 |
|
|
|
| 7 |
C |
-0.298 |
|
|
|
| 8 |
H |
0.152 |
|
|
|
| 9 |
C |
-0.120 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.161 |
|
|
|
| 13 |
C |
-0.510 |
|
|
|
| 14 |
H |
0.165 |
|
|
|
| 15 |
H |
0.165 |
|
|
|
| 16 |
H |
0.162 |
|
|
|
| 17 |
C |
-0.508 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.026 |
-0.027 |
-0.050 |
0.063 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.032 |
0.228 |
-0.173 |
| y |
0.228 |
-34.828 |
0.056 |
| z |
-0.173 |
0.056 |
-34.373 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.431 |
0.228 |
-0.173 |
| y |
0.228 |
-0.126 |
0.056 |
| z |
-0.173 |
0.056 |
0.557 |
|
| Polar |
|---|
| 3z2-r2 | 1.113 |
|---|
| x2-y2 | -0.204 |
|---|
| xy | 0.228 |
|---|
| xz | -0.173 |
|---|
| yz | 0.056 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.781 |
0.032 |
-0.024 |
| y |
0.032 |
7.969 |
-0.008 |
| z |
-0.024 |
-0.008 |
7.346 |
<r2> (average value of r
2) Å
2
| <r2> |
156.945 |
| (<r2>)1/2 |
12.528 |