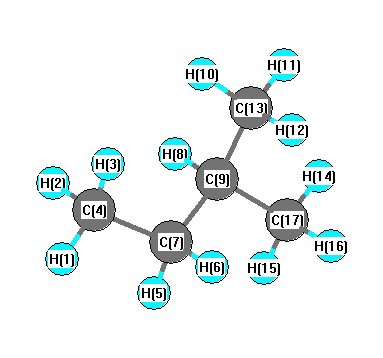
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3062 |
3011 |
31.56 |
|
|
|
| 2 |
A |
3056 |
3006 |
57.24 |
|
|
|
| 3 |
A |
3051 |
3001 |
20.56 |
|
|
|
| 4 |
A |
3051 |
3000 |
53.25 |
|
|
|
| 5 |
A |
3045 |
2995 |
48.61 |
|
|
|
| 6 |
A |
3040 |
2990 |
6.96 |
|
|
|
| 7 |
A |
2991 |
2942 |
24.94 |
|
|
|
| 8 |
A |
2980 |
2931 |
23.52 |
|
|
|
| 9 |
A |
2975 |
2926 |
29.91 |
|
|
|
| 10 |
A |
2970 |
2921 |
32.41 |
|
|
|
| 11 |
A |
2953 |
2904 |
33.75 |
|
|
|
| 12 |
A |
2936 |
2887 |
11.08 |
|
|
|
| 13 |
A |
1496 |
1472 |
3.48 |
|
|
|
| 14 |
A |
1490 |
1466 |
9.14 |
|
|
|
| 15 |
A |
1487 |
1462 |
3.70 |
|
|
|
| 16 |
A |
1482 |
1458 |
12.05 |
|
|
|
| 17 |
A |
1477 |
1453 |
1.50 |
|
|
|
| 18 |
A |
1469 |
1445 |
0.18 |
|
|
|
| 19 |
A |
1460 |
1436 |
0.10 |
|
|
|
| 20 |
A |
1396 |
1373 |
4.19 |
|
|
|
| 21 |
A |
1389 |
1366 |
4.11 |
|
|
|
| 22 |
A |
1377 |
1354 |
5.86 |
|
|
|
| 23 |
A |
1354 |
1331 |
0.34 |
|
|
|
| 24 |
A |
1338 |
1316 |
2.26 |
|
|
|
| 25 |
A |
1295 |
1273 |
3.57 |
|
|
|
| 26 |
A |
1270 |
1249 |
1.93 |
|
|
|
| 27 |
A |
1179 |
1159 |
0.59 |
|
|
|
| 28 |
A |
1173 |
1154 |
1.19 |
|
|
|
| 29 |
A |
1149 |
1130 |
2.90 |
|
|
|
| 30 |
A |
1041 |
1024 |
0.10 |
|
|
|
| 31 |
A |
1009 |
993 |
3.38 |
|
|
|
| 32 |
A |
963 |
947 |
5.40 |
|
|
|
| 33 |
A |
948 |
932 |
0.31 |
|
|
|
| 34 |
A |
908 |
893 |
0.84 |
|
|
|
| 35 |
A |
907 |
892 |
1.24 |
|
|
|
| 36 |
A |
786 |
773 |
0.43 |
|
|
|
| 37 |
A |
754 |
742 |
2.96 |
|
|
|
| 38 |
A |
445 |
438 |
0.31 |
|
|
|
| 39 |
A |
404 |
398 |
0.28 |
|
|
|
| 40 |
A |
356 |
350 |
0.00 |
|
|
|
| 41 |
A |
263 |
259 |
0.00 |
|
|
|
| 42 |
A |
258 |
254 |
0.02 |
|
|
|
| 43 |
A |
224 |
220 |
0.02 |
|
|
|
| 44 |
A |
215 |
212 |
0.04 |
|
|
|
| 45 |
A |
96 |
94 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 34483.1 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 33914.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.156 |
|
|
|
| 2 |
H |
0.154 |
|
|
|
| 3 |
H |
0.159 |
|
|
|
| 4 |
C |
-0.489 |
|
|
|
| 5 |
H |
0.142 |
|
|
|
| 6 |
H |
0.140 |
|
|
|
| 7 |
C |
-0.268 |
|
|
|
| 8 |
H |
0.135 |
|
|
|
| 9 |
C |
-0.085 |
|
|
|
| 10 |
H |
0.157 |
|
|
|
| 11 |
H |
0.153 |
|
|
|
| 12 |
H |
0.150 |
|
|
|
| 13 |
C |
-0.481 |
|
|
|
| 14 |
H |
0.152 |
|
|
|
| 15 |
H |
0.152 |
|
|
|
| 16 |
H |
0.151 |
|
|
|
| 17 |
C |
-0.480 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.028 |
-0.024 |
-0.048 |
0.060 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.090 |
0.215 |
-0.166 |
| y |
0.215 |
-34.909 |
0.051 |
| z |
-0.166 |
0.051 |
-34.484 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.394 |
0.215 |
-0.166 |
| y |
0.215 |
-0.122 |
0.051 |
| z |
-0.166 |
0.051 |
0.516 |
|
| Polar |
|---|
| 3z2-r2 | 1.032 |
|---|
| x2-y2 | -0.181 |
|---|
| xy | 0.215 |
|---|
| xz | -0.166 |
|---|
| yz | 0.051 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.131 |
0.030 |
-0.026 |
| y |
0.030 |
8.219 |
-0.011 |
| z |
-0.026 |
-0.011 |
7.512 |
<r2> (average value of r
2) Å
2
| <r2> |
159.013 |
| (<r2>)1/2 |
12.610 |