return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Butadiene, 2-methyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-195.057862
Energy at 298.15K-195.065515
HF Energy-194.845262
Nuclear repulsion energy160.233547
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3285 3118 16.07      
2 A' 3274 3107 14.36      
3 A' 3205 3042 3.95      
4 A' 3191 3029 10.95      
5 A' 3183 3022 10.65      
6 A' 3169 3008 17.80      
7 A' 3075 2919 18.11      
8 A' 1730 1642 1.46      
9 A' 1685 1599 14.98      
10 A' 1551 1472 6.44      
11 A' 1500 1424 0.51      
12 A' 1468 1393 2.31      
13 A' 1453 1379 4.01      
14 A' 1361 1291 1.04      
15 A' 1350 1281 0.30      
16 A' 1111 1055 4.26      
17 A' 1033 980 0.22      
18 A' 983 933 0.44      
19 A' 805 764 0.16      
20 A' 537 510 0.14      
21 A' 428 406 1.50      
22 A' 282 268 0.84      
23 A" 3135 2976 19.80      
24 A" 1531 1453 6.91      
25 A" 1097 1041 0.14      
26 A" 1035 982 16.84      
27 A" 926 879 28.72      
28 A" 918 872 51.56      
29 A" 791 751 0.48      
30 A" 648 616 0.08      
31 A" 403 383 5.76      
32 A" 209 198 0.36      
33 A" 157 149 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 25253.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23970.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.28463 0.13893 0.09501

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.587 1.735 0.000
C2 0.000 0.525 0.000
C3 -0.828 -0.686 0.000
C4 -0.372 -1.948 0.000
C5 1.500 0.375 0.000
H6 -1.668 1.841 0.000
H7 -0.008 2.652 0.000
H8 -1.904 -0.518 0.000
H9 0.687 -2.182 0.000
H10 -1.053 -2.791 0.000
H11 1.991 1.350 0.000
H12 1.840 -0.180 0.881
H13 1.840 -0.180 -0.881

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.34482.43303.68872.49081.08561.08482.60964.11934.54942.60643.21423.2142
C21.34481.46742.50041.50702.12422.12702.17092.79333.47902.15492.15802.1580
C32.43301.46741.34142.55822.66283.43761.08852.12952.11643.47752.85482.8548
C43.68872.50041.34142.98254.00394.61392.09521.08451.08414.05652.96502.9650
C52.49081.50702.55822.98253.49002.73093.51852.68334.06661.09191.09501.0950
H61.08562.12422.66284.00393.49001.84732.37094.66164.67203.69104.14244.1424
H71.08482.12703.43764.61392.73091.84733.69394.88425.54212.38523.49403.4940
H82.60962.17091.08852.09523.51852.37093.69393.07912.42624.31943.86053.8605
H94.11932.79332.12951.08452.68334.66164.88423.07911.84353.76542.47292.4729
H104.54943.47902.11641.08414.06664.67205.54212.42621.84355.13913.99523.9952
H112.60642.15493.47754.05651.09193.69102.38524.31943.76545.13911.77171.7717
H123.21422.15802.85482.96501.09504.14243.49403.86052.47293.99521.77171.7616
H133.21422.15802.85482.96501.09504.14243.49403.86052.47293.99521.77171.7616

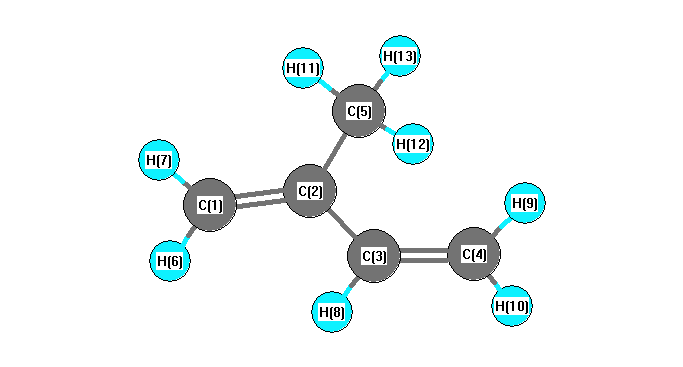
picture of 1,3-Butadiene, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.746 C1 C2 C5 121.612
C2 C1 H6 121.495 C2 C1 H7 121.831
C2 C3 C4 125.739 C2 C3 H8 115.489
C2 C5 H11 111.010 C2 C5 H12 111.072
C2 C5 H13 111.072 C3 C2 C5 118.643
C3 C4 H9 122.408 C3 C4 H10 121.149
C4 C3 H8 118.772 H6 C1 H7 116.674
H9 C4 H10 116.443 H11 C5 H12 108.218
H11 C5 H13 108.218 H12 C5 H13 107.098
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.424      
2 C 0.178      
3 C -0.123      
4 C -0.381      
5 C -0.530      
6 H 0.156      
7 H 0.155      
8 H 0.152      
9 H 0.158      
10 H 0.160      
11 H 0.163      
12 H 0.167      
13 H 0.167      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.674 -0.839 0.000
y -0.839 12.084 0.000
z 0.000 0.000 3.719


<r2> (average value of r2) Å2
<r2> 131.573
(<r2>)1/2 11.471