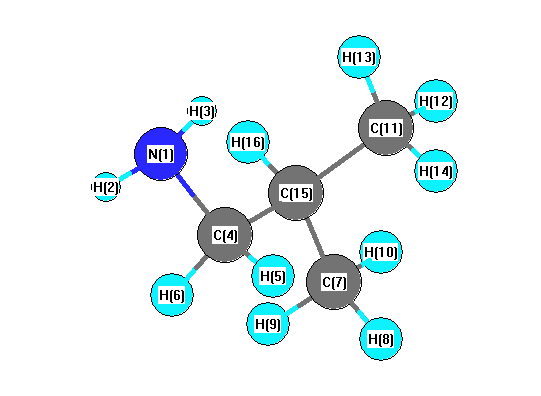
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3606 |
3429 |
0.03 |
|
|
|
| 2 |
A |
3518 |
3346 |
1.17 |
|
|
|
| 3 |
A |
3145 |
2991 |
39.30 |
|
|
|
| 4 |
A |
3143 |
2989 |
26.26 |
|
|
|
| 5 |
A |
3134 |
2980 |
63.43 |
|
|
|
| 6 |
A |
3129 |
2975 |
11.61 |
|
|
|
| 7 |
A |
3092 |
2941 |
36.81 |
|
|
|
| 8 |
A |
3067 |
2917 |
20.96 |
|
|
|
| 9 |
A |
3058 |
2909 |
18.01 |
|
|
|
| 10 |
A |
3053 |
2903 |
26.69 |
|
|
|
| 11 |
A |
2952 |
2807 |
79.51 |
|
|
|
| 12 |
A |
1704 |
1621 |
33.41 |
|
|
|
| 13 |
A |
1544 |
1468 |
15.31 |
|
|
|
| 14 |
A |
1533 |
1458 |
3.50 |
|
|
|
| 15 |
A |
1529 |
1454 |
2.15 |
|
|
|
| 16 |
A |
1520 |
1445 |
2.26 |
|
|
|
| 17 |
A |
1514 |
1440 |
1.22 |
|
|
|
| 18 |
A |
1455 |
1384 |
11.36 |
|
|
|
| 19 |
A |
1438 |
1367 |
1.26 |
|
|
|
| 20 |
A |
1426 |
1356 |
9.79 |
|
|
|
| 21 |
A |
1391 |
1323 |
1.04 |
|
|
|
| 22 |
A |
1365 |
1298 |
0.26 |
|
|
|
| 23 |
A |
1327 |
1262 |
6.95 |
|
|
|
| 24 |
A |
1249 |
1187 |
0.32 |
|
|
|
| 25 |
A |
1214 |
1155 |
2.36 |
|
|
|
| 26 |
A |
1189 |
1131 |
4.72 |
|
|
|
| 27 |
A |
1117 |
1062 |
14.84 |
|
|
|
| 28 |
A |
1108 |
1054 |
1.78 |
|
|
|
| 29 |
A |
982 |
934 |
0.48 |
|
|
|
| 30 |
A |
959 |
912 |
3.77 |
|
|
|
| 31 |
A |
944 |
898 |
49.65 |
|
|
|
| 32 |
A |
937 |
891 |
8.75 |
|
|
|
| 33 |
A |
865 |
823 |
90.56 |
|
|
|
| 34 |
A |
816 |
776 |
3.63 |
|
|
|
| 35 |
A |
488 |
464 |
7.55 |
|
|
|
| 36 |
A |
417 |
396 |
0.64 |
|
|
|
| 37 |
A |
363 |
346 |
0.78 |
|
|
|
| 38 |
A |
287 |
272 |
24.15 |
|
|
|
| 39 |
A |
267 |
254 |
3.21 |
|
|
|
| 40 |
A |
238 |
227 |
15.38 |
|
|
|
| 41 |
A |
221 |
210 |
19.81 |
|
|
|
| 42 |
A |
117 |
111 |
4.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33209.1 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 31581.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.756 |
|
|
|
| 2 |
H |
0.317 |
|
|
|
| 3 |
H |
0.315 |
|
|
|
| 4 |
C |
-0.184 |
|
|
|
| 5 |
H |
0.130 |
|
|
|
| 6 |
H |
0.162 |
|
|
|
| 7 |
C |
-0.514 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.166 |
|
|
|
| 10 |
H |
0.168 |
|
|
|
| 11 |
C |
-0.523 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.173 |
|
|
|
| 14 |
H |
0.157 |
|
|
|
| 15 |
C |
-0.112 |
|
|
|
| 16 |
H |
0.169 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.253 |
-0.169 |
1.237 |
1.273 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.834 |
0.689 |
-3.407 |
| y |
0.689 |
-32.709 |
0.540 |
| z |
-3.407 |
0.540 |
-35.097 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.069 |
0.689 |
-3.407 |
| y |
0.689 |
0.756 |
0.540 |
| z |
-3.407 |
0.540 |
-2.825 |
|
| Polar |
|---|
| 3z2-r2 | -5.651 |
|---|
| x2-y2 | 0.876 |
|---|
| xy | 0.689 |
|---|
| xz | -3.407 |
|---|
| yz | 0.540 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.233 |
-0.001 |
-0.009 |
| y |
-0.001 |
7.587 |
0.028 |
| z |
-0.009 |
0.028 |
6.558 |
<r2> (average value of r
2) Å
2
| <r2> |
149.217 |
| (<r2>)1/2 |
12.215 |