return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(NH2)H2C(CH3)HCH3 (1-Propanamine, 2-methyl-)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-213.058213
Energy at 298.15K-213.071054
HF Energy-212.315413
Nuclear repulsion energy193.632231
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3616 3394 0.06      
2 A 3532 3315 0.59      
3 A 3181 2986 28.73      
4 A 3173 2978 34.58      
5 A 3166 2971 58.72      
6 A 3161 2967 8.83      
7 A 3126 2934 32.61      
8 A 3102 2911 15.72      
9 A 3090 2900 16.35      
10 A 3086 2897 22.69      
11 A 3011 2826 60.42      
12 A 1739 1632 35.74      
13 A 1578 1481 13.40      
14 A 1568 1472 1.40      
15 A 1565 1469 4.73      
16 A 1555 1460 2.55      
17 A 1549 1454 0.95      
18 A 1487 1396 9.57      
19 A 1474 1384 1.65      
20 A 1465 1375 9.34      
21 A 1426 1339 0.34      
22 A 1395 1310 0.04      
23 A 1359 1275 6.10      
24 A 1276 1198 0.21      
25 A 1242 1166 2.75      
26 A 1218 1143 6.38      
27 A 1130 1061 3.75      
28 A 1125 1056 10.00      
29 A 1004 942 0.38      
30 A 982 921 40.54      
31 A 973 913 42.70      
32 A 958 899 1.43      
33 A 895 840 63.98      
34 A 829 778 1.78      
35 A 497 467 7.02      
36 A 430 404 0.61      
37 A 375 352 1.16      
38 A 298 279 21.45      
39 A 274 257 9.61      
40 A 245 230 20.16      
41 A 229 215 12.14      
42 A 125 118 4.99      

Unscaled Zero Point Vibrational Energy (zpe) 33752.7 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 31680.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.25056 0.11589 0.08734

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -2.004 -0.025 -0.191
H2 -2.802 -0.608 0.053
H3 -2.124 0.838 0.334
C4 -0.766 -0.692 0.234
H5 -0.653 -0.756 1.332
H6 -0.801 -1.722 -0.140
C7 1.735 -0.780 0.028
H8 1.877 -0.790 1.115
H9 1.679 -1.819 -0.315
H10 2.623 -0.324 -0.422
C11 0.572 1.455 0.109
H12 1.476 1.925 -0.291
H13 -0.285 2.041 -0.236
H14 0.619 1.516 1.203
C15 0.471 -0.003 -0.346
H16 0.356 -0.013 -1.438

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 C7 H8 H9 H10 C11 H12 H13 H14 C15 H16
N11.01771.01741.46882.16292.08073.82064.16624.09844.64242.98593.99082.68823.34712.47972.6695
H21.01771.62192.04512.50462.29774.53964.80154.65555.45263.95484.98373.66564.18833.35163.5423
H31.01741.62192.04802.38752.92044.19564.39014.68434.94522.77493.81262.27032.95682.81123.1650
C41.46882.04512.04801.10521.09632.51112.78832.74773.47152.53273.48602.81452.78131.53012.1249
H52.16292.50462.38751.10521.76612.72092.54013.04613.74072.80813.78913.22772.60772.15543.0397
H62.08072.29772.92041.09631.76612.71003.10092.48763.70893.46954.30193.79933.78242.14792.4375
C73.82064.53964.19562.51112.72092.71001.09671.09551.09492.52072.73593.47972.81081.53032.1529
H84.16624.80154.39012.78832.54013.10091.09671.77291.77032.78513.08373.81022.62882.17573.0713
H94.09844.65554.68432.74773.04612.48761.09551.77291.77173.48183.74954.33183.81492.18142.5038
H104.64245.45264.94523.47153.74073.70891.09491.77031.77172.76602.52743.75283.16872.17732.5033
C112.98593.95482.77492.53272.80813.46952.52072.78513.48182.76601.09501.09401.09761.53052.1434
H123.99084.98373.81263.48603.78914.30192.73593.08373.74952.52741.09501.76621.77082.17512.5153
H132.68823.66562.27032.81453.22773.79933.47973.81024.33183.75281.09401.76621.77912.18212.4653
H143.34714.18832.95682.78132.60773.78242.81082.62883.81493.16871.09761.77081.77912.17533.0636
C152.47973.35162.81121.53012.15542.14791.53032.17572.18142.17731.53052.17512.18212.17531.0975
H162.66953.54233.16502.12493.03972.43752.15293.07132.50382.50332.14342.51532.46533.06361.0975

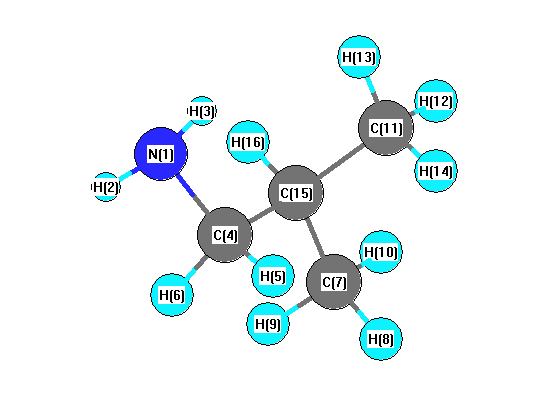
picture of 1-Propanamine, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C4 H5 113.585 N1 C4 H6 107.528
N1 C4 C15 111.540 H2 N1 H3 105.679
H2 N1 C4 109.320 H3 N1 C4 109.577
C4 C15 C7 110.274 C4 C15 C11 111.688
C4 C15 H16 106.778 H5 C4 H6 106.683
H5 C4 C15 108.675 H6 C4 C15 108.608
C7 C15 C11 110.882 C7 C15 H16 108.911
H8 C7 H9 107.946 H8 C7 H10 107.755
H8 C7 C15 110.753 H9 C7 H10 107.972
H9 C7 C15 111.280 H10 C7 C15 110.991
C11 C15 H16 108.162 H12 C11 H13 107.580
H12 C11 H14 107.730 H12 C11 C15 110.792
H13 C11 H14 108.542 H13 C11 C15 111.408
H14 C11 C15 110.653
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability