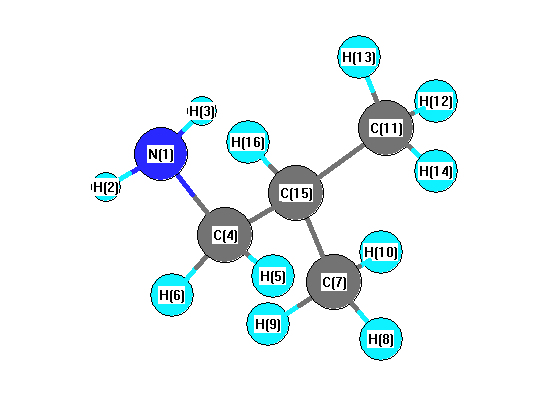
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3465 |
3408 |
0.92 |
|
|
|
| 2 |
A |
3379 |
3323 |
3.86 |
|
|
|
| 3 |
A |
3055 |
3005 |
40.26 |
|
|
|
| 4 |
A |
3053 |
3003 |
24.85 |
|
|
|
| 5 |
A |
3044 |
2994 |
62.31 |
|
|
|
| 6 |
A |
3039 |
2989 |
10.87 |
|
|
|
| 7 |
A |
2998 |
2948 |
36.73 |
|
|
|
| 8 |
A |
2975 |
2926 |
32.86 |
|
|
|
| 9 |
A |
2970 |
2921 |
11.69 |
|
|
|
| 10 |
A |
2967 |
2918 |
23.78 |
|
|
|
| 11 |
A |
2850 |
2803 |
84.58 |
|
|
|
| 12 |
A |
1652 |
1625 |
26.57 |
|
|
|
| 13 |
A |
1498 |
1473 |
14.00 |
|
|
|
| 14 |
A |
1487 |
1463 |
3.57 |
|
|
|
| 15 |
A |
1481 |
1457 |
1.53 |
|
|
|
| 16 |
A |
1474 |
1450 |
2.20 |
|
|
|
| 17 |
A |
1468 |
1444 |
1.24 |
|
|
|
| 18 |
A |
1405 |
1382 |
11.36 |
|
|
|
| 19 |
A |
1385 |
1362 |
0.46 |
|
|
|
| 20 |
A |
1374 |
1352 |
7.72 |
|
|
|
| 21 |
A |
1345 |
1323 |
2.18 |
|
|
|
| 22 |
A |
1320 |
1298 |
0.43 |
|
|
|
| 23 |
A |
1280 |
1258 |
7.52 |
|
|
|
| 24 |
A |
1208 |
1188 |
0.51 |
|
|
|
| 25 |
A |
1172 |
1153 |
2.98 |
|
|
|
| 26 |
A |
1150 |
1131 |
4.04 |
|
|
|
| 27 |
A |
1074 |
1056 |
2.22 |
|
|
|
| 28 |
A |
1068 |
1051 |
9.55 |
|
|
|
| 29 |
A |
950 |
935 |
0.23 |
|
|
|
| 30 |
A |
927 |
911 |
11.73 |
|
|
|
| 31 |
A |
913 |
898 |
36.20 |
|
|
|
| 32 |
A |
907 |
892 |
13.23 |
|
|
|
| 33 |
A |
845 |
831 |
81.85 |
|
|
|
| 34 |
A |
788 |
775 |
3.42 |
|
|
|
| 35 |
A |
473 |
465 |
7.55 |
|
|
|
| 36 |
A |
404 |
397 |
0.73 |
|
|
|
| 37 |
A |
352 |
346 |
1.04 |
|
|
|
| 38 |
A |
282 |
278 |
24.68 |
|
|
|
| 39 |
A |
261 |
256 |
4.72 |
|
|
|
| 40 |
A |
232 |
228 |
20.17 |
|
|
|
| 41 |
A |
218 |
214 |
9.92 |
|
|
|
| 42 |
A |
119 |
117 |
5.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32153.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 31622.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.702 |
|
|
|
| 2 |
H |
0.297 |
|
|
|
| 3 |
H |
0.297 |
|
|
|
| 4 |
C |
-0.177 |
|
|
|
| 5 |
H |
0.117 |
|
|
|
| 6 |
H |
0.147 |
|
|
|
| 7 |
C |
-0.484 |
|
|
|
| 8 |
H |
0.150 |
|
|
|
| 9 |
H |
0.153 |
|
|
|
| 10 |
H |
0.155 |
|
|
|
| 11 |
C |
-0.493 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.160 |
|
|
|
| 14 |
H |
0.146 |
|
|
|
| 15 |
C |
-0.076 |
|
|
|
| 16 |
H |
0.151 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.252 |
-0.143 |
1.222 |
1.256 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.954 |
0.637 |
-3.413 |
| y |
0.637 |
-32.854 |
0.508 |
| z |
-3.413 |
0.508 |
-35.220 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.083 |
0.637 |
-3.413 |
| y |
0.637 |
0.733 |
0.508 |
| z |
-3.413 |
0.508 |
-2.816 |
|
| Polar |
|---|
| 3z2-r2 | -5.633 |
|---|
| x2-y2 | 0.900 |
|---|
| xy | 0.637 |
|---|
| xz | -3.413 |
|---|
| yz | 0.508 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.616 |
0.001 |
-0.021 |
| y |
0.001 |
7.863 |
0.018 |
| z |
-0.021 |
0.018 |
6.740 |
<r2> (average value of r
2) Å
2
| <r2> |
151.231 |
| (<r2>)1/2 |
12.298 |