return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: CID/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CID/6-31G*
 hartrees
Energy at 0K-210.591505
Energy at 298.15K-210.598944
HF Energy-209.997173
Nuclear repulsion energy159.490307
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3260 3011 22.12      
2 A' 3255 3007 37.22      
3 A' 3177 2935 5.00      
4 A' 3172 2930 13.43      
5 A' 2488 2299 5.66      
6 A' 1603 1481 10.86      
7 A' 1595 1473 7.08      
8 A' 1524 1407 4.07      
9 A' 1441 1331 7.14      
10 A' 1269 1172 2.78      
11 A' 1196 1105 6.14      
12 A' 988 913 0.31      
13 A' 811 749 1.57      
14 A' 580 536 1.40      
15 A' 372 344 0.30      
16 A' 304 281 1.23      
17 A' 233 215 2.65      
18 A" 3258 3010 13.57      
19 A" 3250 3002 0.26      
20 A" 3169 2927 15.84      
21 A" 1584 1463 3.72      
22 A" 1582 1461 0.13      
23 A" 1503 1388 4.48      
24 A" 1419 1311 0.15      
25 A" 1207 1115 1.59      
26 A" 1033 954 0.42      
27 A" 993 917 1.88      
28 A" 599 553 0.00      
29 A" 242 224 0.02      
30 A" 197 182 5.50      

Unscaled Zero Point Vibrational Energy (zpe) 23651.2 cm-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 21846.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CID/6-31G*
ABC
0.26686 0.13260 0.09707

See section I.F.4 to change rotational constant units
Geometric Data calculated at CID/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.402 -2.173 0.000
C2 0.027 -1.084 0.000
C3 -0.440 0.312 0.000
C4 0.027 1.026 1.270
C5 0.027 1.026 -1.270
H6 -1.530 0.273 0.000
H7 -0.360 2.043 1.282
H8 -0.323 0.513 2.162
H9 1.114 1.072 1.305
H10 -0.360 2.043 -1.282
H11 -0.323 0.513 -2.162
H12 1.114 1.072 -1.305

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.15142.62423.46203.46203.11694.47213.52373.56914.47213.52373.5691
C21.15141.47282.46282.46282.06563.40202.71112.74473.40202.71112.7447
C32.62421.47281.52961.52961.09092.15512.17492.16702.15512.17492.1670
C43.46202.46281.52962.53932.14611.08871.08771.08852.77373.48782.7950
C53.46202.46281.52962.53932.14612.77373.48782.79501.08871.08771.0885
H63.11692.06561.09092.14612.14612.47922.48823.05542.47922.48823.0554
H74.47213.40202.15511.08872.77372.47921.76611.76592.56303.76883.1317
H83.52372.71112.17491.08773.48782.48821.76611.76463.76884.32493.7949
H93.56912.74472.16701.08852.79503.05541.76591.76463.13173.79492.6099
H104.47213.40202.15512.77371.08872.47922.56303.76883.13171.76611.7659
H113.52372.71112.17493.48781.08772.48823.76884.32493.79491.76611.7646
H123.56912.74472.16702.79501.08853.05543.13173.79492.60991.76591.7646

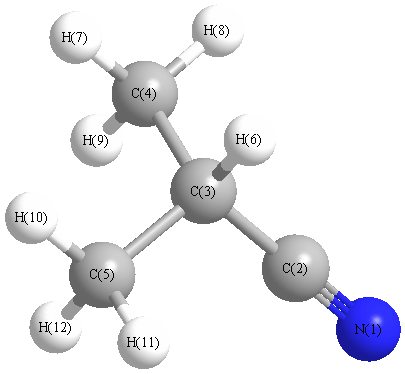
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.493 C2 C3 C4 110.207
C2 C3 C5 110.207 C2 C3 H6 106.409
C3 C4 H7 109.644 C3 C4 H8 111.275
C3 C4 H9 110.597 C3 C5 H10 109.644
C3 C5 H11 111.275 C3 C5 H12 110.597
C4 C3 C5 112.206 C4 C3 H6 108.811
C5 C3 H6 108.811 H7 C4 H8 108.484
H7 C4 H9 108.406 H8 C4 H9 108.359
H10 C5 H11 108.484 H10 C5 H12 108.406
H11 C5 H12 108.359
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability