return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-210.714362
Energy at 298.15K-210.721660
HF Energy-209.994456
Nuclear repulsion energy158.422472
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3161 3010 22.73      
2 A' 3156 3005 38.70      
3 A' 3076 2929 14.83      
4 A' 3072 2926 5.07      
5 A' 2341 2229 1.13      
6 A' 1558 1483 10.50      
7 A' 1550 1476 6.69      
8 A' 1476 1405 3.80      
9 A' 1395 1328 6.27      
10 A' 1232 1173 2.39      
11 A' 1159 1104 5.24      
12 A' 960 914 0.36      
13 A' 786 749 1.02      
14 A' 554 527 1.07      
15 A' 363 345 0.23      
16 A' 295 281 0.84      
17 A' 223 212 2.49      
18 A" 3159 3008 14.14      
19 A" 3152 3001 0.22      
20 A" 3073 2926 15.89      
21 A" 1539 1466 3.29      
22 A" 1537 1464 0.37      
23 A" 1456 1386 4.55      
24 A" 1378 1312 0.28      
25 A" 1172 1116 1.83      
26 A" 1004 956 0.33      
27 A" 964 918 1.77      
28 A" 566 539 0.03      
29 A" 239 227 0.01      
30 A" 186 177 4.80      

Unscaled Zero Point Vibrational Energy (zpe) 22889.0 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 21797.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.26467 0.13060 0.09582

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.407 -2.192 0.000
C2 0.027 -1.088 0.000
C3 -0.443 0.316 0.000
C4 0.027 1.033 1.274
C5 0.027 1.033 -1.274
H6 -1.541 0.277 0.000
H7 -0.363 2.058 1.285
H8 -0.326 0.517 2.174
H9 1.122 1.078 1.308
H10 -0.363 2.058 -1.285
H11 -0.326 0.517 -2.174
H12 1.122 1.078 -1.308

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.16832.64823.48873.48873.14564.50673.54993.59404.50673.54993.5940
C21.16831.47992.47402.47402.07903.42032.72472.75673.42032.72472.7567
C32.64821.47991.53571.53571.09952.16642.18632.17702.16642.18632.1770
C43.48872.47401.53572.54852.15741.09671.09571.09652.78443.50442.8054
C53.48872.47401.53572.54852.15742.78443.50442.80541.09671.09571.0965
H63.14562.07901.09952.15742.15742.49242.50213.07332.49242.50213.0733
H74.50673.42032.16641.09672.78442.49241.77971.77922.57023.78713.1448
H83.54992.72472.18631.09573.50442.50211.77971.77783.78714.34793.8126
H93.59402.75672.17701.09652.80543.07331.77921.77783.14483.81262.6163
H104.50673.42032.16642.78441.09672.49242.57023.78713.14481.77971.7792
H113.54992.72472.18633.50441.09572.50213.78714.34793.81261.77971.7778
H123.59402.75672.17702.80541.09653.07333.14483.81262.61631.77921.7778

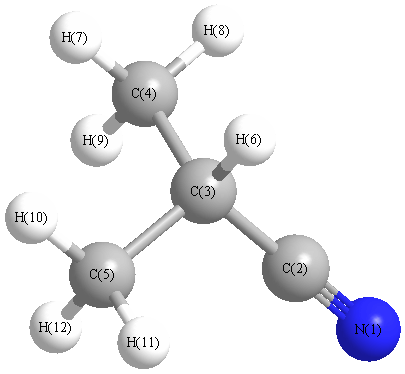
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.517 C2 C3 C4 110.236
C2 C3 C5 110.236 C2 C3 H6 106.484
C3 C4 H7 109.640 C3 C4 H8 111.275
C3 C4 H9 110.489 C3 C5 H10 109.640
C3 C5 H11 111.275 C3 C5 H12 110.489
C4 C3 C5 112.148 C4 C3 H6 108.776
C5 C3 H6 108.776 H7 C4 H8 108.541
H7 C4 H9 108.435 H8 C4 H9 108.389
H10 C5 H11 108.541 H10 C5 H12 108.435
H11 C5 H12 108.389
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability