return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-210.698211
Energy at 298.15K-210.705536
HF Energy-209.995977
Nuclear repulsion energy158.824936
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3201 3004 18.73      
2 A' 3195 2999 32.98      
3 A' 3113 2922 12.69      
4 A' 3111 2920 4.36      
5 A' 2442 2292 3.35      
6 A' 1568 1472 11.57      
7 A' 1560 1464 7.59      
8 A' 1485 1394 4.45      
9 A' 1404 1318 6.36      
10 A' 1240 1164 2.65      
11 A' 1167 1095 5.77      
12 A' 969 909 0.42      
13 A' 795 746 1.13      
14 A' 560 526 1.16      
15 A' 364 342 0.23      
16 A' 295 277 0.92      
17 A' 223 210 2.48      
18 A" 3199 3003 12.00      
19 A" 3192 2996 0.15      
20 A" 3110 2919 13.48      
21 A" 1549 1454 3.60      
22 A" 1548 1453 0.49      
23 A" 1464 1375 5.45      
24 A" 1385 1300 0.20      
25 A" 1179 1107 1.85      
26 A" 1011 949 0.38      
27 A" 969 910 1.98      
28 A" 575 540 0.03      
29 A" 238 223 0.02      
30 A" 188 176 4.91      

Unscaled Zero Point Vibrational Energy (zpe) 23149.8 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 21728.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.26518 0.13143 0.09630

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.406 -2.183 0.000
C2 0.027 -1.087 0.000
C3 -0.442 0.314 0.000
C4 0.027 1.030 1.274
C5 0.027 1.030 -1.274
H6 -1.537 0.274 0.000
H7 -0.363 2.052 1.283
H8 -0.325 0.514 2.170
H9 1.119 1.076 1.307
H10 -0.363 2.052 -1.283
H11 -0.325 0.514 -2.170
H12 1.119 1.076 -1.307

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.15982.63673.47763.47763.13254.49223.53843.58304.49223.53843.5830
C21.15981.47692.47112.47112.07353.41412.72002.75283.41412.72002.7528
C32.63671.47691.53471.53471.09662.16272.18282.17352.16272.18282.1735
C43.47762.47111.53472.54722.15441.09401.09301.09382.78123.50022.8022
C53.47762.47111.53472.54722.15442.78123.50022.80221.09401.09301.0938
H63.13252.07351.09662.15442.15442.48792.49773.06732.48792.49773.0673
H74.49223.41412.16271.09402.78122.48791.77611.77552.56693.78113.1401
H83.53842.72002.18281.09303.50022.49771.77611.77403.78114.34083.8066
H93.58302.75282.17351.09382.80223.06731.77551.77403.14013.80662.6131
H104.49223.41412.16272.78121.09402.48792.56693.78113.14011.77611.7755
H113.53842.72002.18283.50021.09302.49773.78114.34083.80661.77611.7740
H123.58302.75282.17352.80221.09383.06733.14013.80662.61311.77551.7740

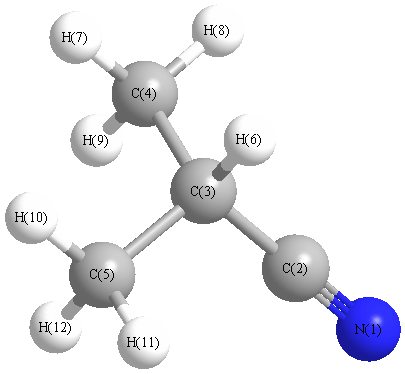
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.440 C2 C3 C4 110.253
C2 C3 C5 110.253 C2 C3 H6 106.418
C3 C4 H7 109.574 C3 C4 H8 111.223
C3 C4 H9 110.436 C3 C5 H10 109.574
C3 C5 H11 111.223 C3 C5 H12 110.436
C4 C3 C5 112.166 C4 C3 H6 108.781
C5 C3 H6 108.781 H7 C4 H8 108.613
H7 C4 H9 108.493 H8 C4 H9 108.436
H10 C5 H11 108.613 H10 C5 H12 108.493
H11 C5 H12 108.436
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability