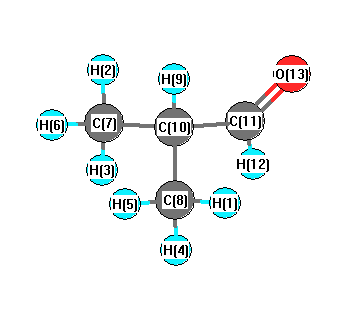
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3142 |
3006 |
23.62 |
|
|
|
| 2 |
A' |
3133 |
2997 |
52.25 |
|
|
|
| 3 |
A' |
3090 |
2956 |
5.64 |
|
|
|
| 4 |
A' |
3056 |
2924 |
17.87 |
|
|
|
| 5 |
A' |
2873 |
2749 |
119.66 |
|
|
|
| 6 |
A' |
1851 |
1771 |
182.95 |
|
|
|
| 7 |
A' |
1536 |
1469 |
16.74 |
|
|
|
| 8 |
A' |
1529 |
1463 |
8.78 |
|
|
|
| 9 |
A' |
1446 |
1384 |
2.04 |
|
|
|
| 10 |
A' |
1430 |
1368 |
3.48 |
|
|
|
| 11 |
A' |
1326 |
1269 |
1.08 |
|
|
|
| 12 |
A' |
1193 |
1142 |
0.21 |
|
|
|
| 13 |
A' |
1190 |
1138 |
8.54 |
|
|
|
| 14 |
A' |
934 |
893 |
20.41 |
|
|
|
| 15 |
A' |
855 |
818 |
23.51 |
|
|
|
| 16 |
A' |
546 |
523 |
5.91 |
|
|
|
| 17 |
A' |
355 |
339 |
6.97 |
|
|
|
| 18 |
A' |
330 |
315 |
0.22 |
|
|
|
| 19 |
A' |
233 |
223 |
0.83 |
|
|
|
| 20 |
A" |
3141 |
3005 |
16.21 |
|
|
|
| 21 |
A" |
3128 |
2992 |
2.80 |
|
|
|
| 22 |
A" |
3053 |
2921 |
23.75 |
|
|
|
| 23 |
A" |
1516 |
1451 |
1.77 |
|
|
|
| 24 |
A" |
1512 |
1447 |
1.28 |
|
|
|
| 25 |
A" |
1422 |
1360 |
3.99 |
|
|
|
| 26 |
A" |
1358 |
1300 |
0.98 |
|
|
|
| 27 |
A" |
1148 |
1098 |
0.39 |
|
|
|
| 28 |
A" |
998 |
955 |
0.16 |
|
|
|
| 29 |
A" |
962 |
921 |
0.83 |
|
|
|
| 30 |
A" |
942 |
902 |
1.54 |
|
|
|
| 31 |
A" |
325 |
311 |
0.66 |
|
|
|
| 32 |
A" |
216 |
207 |
0.07 |
|
|
|
| 33 |
A" |
67 |
64 |
4.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24918.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 23839.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.187 |
|
|
0.116 |
| 2 |
H |
0.187 |
|
|
0.116 |
| 3 |
H |
0.172 |
|
|
0.110 |
| 4 |
H |
0.172 |
|
|
0.110 |
| 5 |
H |
0.181 |
|
|
0.129 |
| 6 |
H |
0.181 |
|
|
0.129 |
| 7 |
C |
-0.524 |
|
|
-0.461 |
| 8 |
C |
-0.524 |
|
|
-0.461 |
| 9 |
H |
0.187 |
|
|
-0.011 |
| 10 |
C |
-0.203 |
|
|
0.304 |
| 11 |
C |
0.261 |
|
|
0.370 |
| 12 |
H |
0.113 |
|
|
-0.014 |
| 13 |
O |
-0.390 |
|
|
-0.433 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.377 |
1.471 |
0.000 |
2.795 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-2.413 |
1.474 |
0.000 |
2.828 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.432 |
3.408 |
0.000 |
| y |
3.408 |
-32.512 |
0.000 |
| z |
0.000 |
0.000 |
-30.348 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.002 |
3.408 |
0.000 |
| y |
3.408 |
0.377 |
0.000 |
| z |
0.000 |
0.000 |
3.625 |
|
| Polar |
|---|
| 3z2-r2 | 7.249 |
|---|
| x2-y2 | -2.920 |
|---|
| xy | 3.408 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
134.712 |
| (<r2>)1/2 |
11.607 |