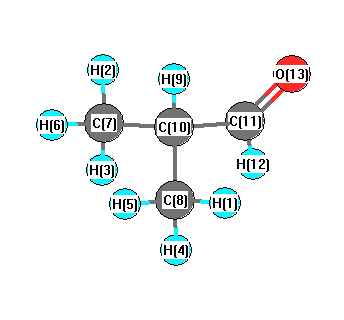
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3160 |
3003 |
20.94 |
|
|
|
| 2 |
A' |
3149 |
2993 |
46.13 |
|
|
|
| 3 |
A' |
3107 |
2953 |
5.02 |
|
|
|
| 4 |
A' |
3069 |
2917 |
16.80 |
|
|
|
| 5 |
A' |
2892 |
2749 |
113.20 |
|
|
|
| 6 |
A' |
1873 |
1779 |
185.52 |
|
|
|
| 7 |
A' |
1538 |
1461 |
17.67 |
|
|
|
| 8 |
A' |
1530 |
1454 |
9.36 |
|
|
|
| 9 |
A' |
1447 |
1375 |
2.39 |
|
|
|
| 10 |
A' |
1432 |
1361 |
4.02 |
|
|
|
| 11 |
A' |
1329 |
1263 |
0.80 |
|
|
|
| 12 |
A' |
1198 |
1139 |
0.02 |
|
|
|
| 13 |
A' |
1195 |
1135 |
8.82 |
|
|
|
| 14 |
A' |
944 |
897 |
19.56 |
|
|
|
| 15 |
A' |
860 |
817 |
23.49 |
|
|
|
| 16 |
A' |
550 |
523 |
6.44 |
|
|
|
| 17 |
A' |
351 |
333 |
6.74 |
|
|
|
| 18 |
A' |
331 |
315 |
0.27 |
|
|
|
| 19 |
A' |
234 |
222 |
0.87 |
|
|
|
| 20 |
A" |
3159 |
3002 |
14.42 |
|
|
|
| 21 |
A" |
3144 |
2988 |
3.03 |
|
|
|
| 22 |
A" |
3067 |
2915 |
21.22 |
|
|
|
| 23 |
A" |
1516 |
1441 |
2.20 |
|
|
|
| 24 |
A" |
1513 |
1438 |
1.09 |
|
|
|
| 25 |
A" |
1423 |
1352 |
5.01 |
|
|
|
| 26 |
A" |
1361 |
1293 |
0.72 |
|
|
|
| 27 |
A" |
1157 |
1100 |
0.29 |
|
|
|
| 28 |
A" |
1006 |
956 |
0.23 |
|
|
|
| 29 |
A" |
965 |
917 |
0.78 |
|
|
|
| 30 |
A" |
942 |
895 |
1.70 |
|
|
|
| 31 |
A" |
326 |
309 |
0.65 |
|
|
|
| 32 |
A" |
213 |
203 |
0.09 |
|
|
|
| 33 |
A" |
64 |
61 |
5.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25022.8 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 23779.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.191 |
|
|
|
| 2 |
H |
0.191 |
|
|
|
| 3 |
H |
0.176 |
|
|
|
| 4 |
H |
0.176 |
|
|
|
| 5 |
H |
0.186 |
|
|
|
| 6 |
H |
0.186 |
|
|
|
| 7 |
C |
-0.534 |
|
|
|
| 8 |
C |
-0.534 |
|
|
|
| 9 |
H |
0.193 |
|
|
|
| 10 |
C |
-0.222 |
|
|
|
| 11 |
C |
0.262 |
|
|
|
| 12 |
H |
0.117 |
|
|
|
| 13 |
O |
-0.389 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.446 |
2.753 |
0.000 |
2.789 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.458 |
0.954 |
0.000 |
| y |
0.954 |
-37.605 |
0.000 |
| z |
0.000 |
0.000 |
-30.429 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.559 |
0.954 |
0.000 |
| y |
0.954 |
-7.161 |
0.000 |
| z |
0.000 |
0.000 |
3.602 |
|
| Polar |
|---|
| 3z2-r2 | 7.205 |
|---|
| x2-y2 | 7.147 |
|---|
| xy | 0.954 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.235 |
-0.119 |
0.000 |
| y |
-0.119 |
7.482 |
0.000 |
| z |
0.000 |
0.000 |
6.051 |
<r2> (average value of r
2) Å
2
| <r2> |
134.216 |
| (<r2>)1/2 |
11.585 |