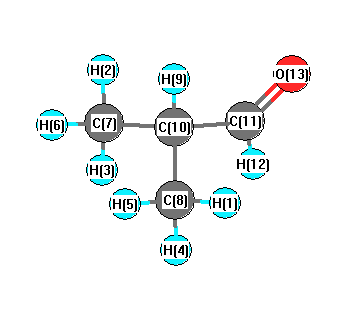
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3068 |
3018 |
20.10 |
|
|
|
| 2 |
A' |
3057 |
3006 |
47.95 |
|
|
|
| 3 |
A' |
3011 |
2961 |
7.57 |
|
|
|
| 4 |
A' |
2982 |
2933 |
17.41 |
|
|
|
| 5 |
A' |
2757 |
2711 |
131.69 |
|
|
|
| 6 |
A' |
1781 |
1752 |
169.66 |
|
|
|
| 7 |
A' |
1491 |
1467 |
14.99 |
|
|
|
| 8 |
A' |
1485 |
1460 |
9.69 |
|
|
|
| 9 |
A' |
1398 |
1375 |
1.74 |
|
|
|
| 10 |
A' |
1382 |
1359 |
2.91 |
|
|
|
| 11 |
A' |
1278 |
1257 |
0.98 |
|
|
|
| 12 |
A' |
1156 |
1137 |
1.24 |
|
|
|
| 13 |
A' |
1149 |
1131 |
6.76 |
|
|
|
| 14 |
A' |
904 |
889 |
21.69 |
|
|
|
| 15 |
A' |
830 |
816 |
25.49 |
|
|
|
| 16 |
A' |
531 |
522 |
4.82 |
|
|
|
| 17 |
A' |
340 |
335 |
5.67 |
|
|
|
| 18 |
A' |
322 |
316 |
0.26 |
|
|
|
| 19 |
A' |
223 |
219 |
0.77 |
|
|
|
| 20 |
A" |
3068 |
3017 |
17.16 |
|
|
|
| 21 |
A" |
3052 |
3002 |
1.98 |
|
|
|
| 22 |
A" |
2980 |
2931 |
22.18 |
|
|
|
| 23 |
A" |
1471 |
1446 |
1.58 |
|
|
|
| 24 |
A" |
1468 |
1444 |
1.29 |
|
|
|
| 25 |
A" |
1371 |
1348 |
3.24 |
|
|
|
| 26 |
A" |
1312 |
1291 |
1.59 |
|
|
|
| 27 |
A" |
1108 |
1089 |
0.32 |
|
|
|
| 28 |
A" |
969 |
953 |
0.23 |
|
|
|
| 29 |
A" |
929 |
914 |
0.70 |
|
|
|
| 30 |
A" |
911 |
896 |
1.65 |
|
|
|
| 31 |
A" |
317 |
312 |
0.45 |
|
|
|
| 32 |
A" |
207 |
204 |
0.10 |
|
|
|
| 33 |
A" |
63 |
62 |
3.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24183.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 23784.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.176 |
|
|
|
| 2 |
H |
0.176 |
|
|
|
| 3 |
H |
0.163 |
|
|
|
| 4 |
H |
0.163 |
|
|
|
| 5 |
H |
0.171 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
| 7 |
C |
-0.497 |
|
|
|
| 8 |
C |
-0.497 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
C |
-0.165 |
|
|
|
| 11 |
C |
0.235 |
|
|
|
| 12 |
H |
0.094 |
|
|
|
| 13 |
O |
-0.355 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.352 |
2.645 |
0.000 |
2.668 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.719 |
0.844 |
0.000 |
| y |
0.844 |
-37.287 |
-0.008 |
| z |
0.000 |
-0.008 |
-30.477 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.164 |
0.844 |
0.000 |
| y |
0.844 |
-6.689 |
-0.008 |
| z |
0.000 |
-0.008 |
3.526 |
|
| Polar |
|---|
| 3z2-r2 | 7.052 |
|---|
| x2-y2 | 6.569 |
|---|
| xy | 0.844 |
|---|
| xz | 0.000 |
|---|
| yz | -0.008 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.454 |
-0.076 |
-0.003 |
| y |
-0.076 |
7.846 |
0.002 |
| z |
-0.003 |
0.002 |
6.291 |
<r2> (average value of r
2) Å
2
| <r2> |
135.989 |
| (<r2>)1/2 |
11.661 |