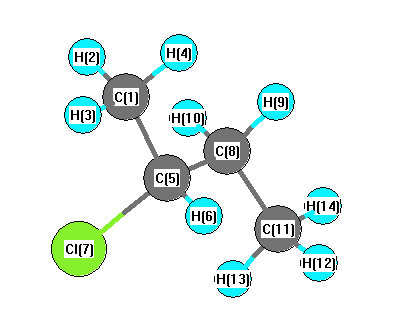
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3162 |
3025 |
18.36 |
|
|
|
| 2 |
A |
3153 |
3017 |
22.10 |
|
|
|
| 3 |
A |
3140 |
3004 |
26.28 |
|
|
|
| 4 |
A |
3134 |
2999 |
30.58 |
|
|
|
| 5 |
A |
3107 |
2973 |
14.30 |
|
|
|
| 6 |
A |
3089 |
2955 |
1.89 |
|
|
|
| 7 |
A |
3062 |
2930 |
15.58 |
|
|
|
| 8 |
A |
3062 |
2929 |
30.06 |
|
|
|
| 9 |
A |
3043 |
2911 |
19.43 |
|
|
|
| 10 |
A |
1531 |
1465 |
6.92 |
|
|
|
| 11 |
A |
1523 |
1457 |
11.13 |
|
|
|
| 12 |
A |
1521 |
1455 |
0.72 |
|
|
|
| 13 |
A |
1511 |
1446 |
7.59 |
|
|
|
| 14 |
A |
1497 |
1433 |
0.82 |
|
|
|
| 15 |
A |
1438 |
1375 |
6.80 |
|
|
|
| 16 |
A |
1434 |
1372 |
5.95 |
|
|
|
| 17 |
A |
1411 |
1350 |
0.38 |
|
|
|
| 18 |
A |
1340 |
1282 |
18.07 |
|
|
|
| 19 |
A |
1333 |
1275 |
5.63 |
|
|
|
| 20 |
A |
1280 |
1224 |
14.49 |
|
|
|
| 21 |
A |
1191 |
1140 |
7.61 |
|
|
|
| 22 |
A |
1142 |
1093 |
2.70 |
|
|
|
| 23 |
A |
1108 |
1060 |
4.49 |
|
|
|
| 24 |
A |
1054 |
1008 |
1.84 |
|
|
|
| 25 |
A |
1015 |
971 |
11.29 |
|
|
|
| 26 |
A |
982 |
939 |
7.76 |
|
|
|
| 27 |
A |
862 |
825 |
9.63 |
|
|
|
| 28 |
A |
804 |
769 |
12.79 |
|
|
|
| 29 |
A |
621 |
594 |
25.51 |
|
|
|
| 30 |
A |
461 |
441 |
1.40 |
|
|
|
| 31 |
A |
390 |
373 |
2.50 |
|
|
|
| 32 |
A |
326 |
312 |
1.37 |
|
|
|
| 33 |
A |
254 |
243 |
0.20 |
|
|
|
| 34 |
A |
233 |
223 |
0.07 |
|
|
|
| 35 |
A |
215 |
206 |
1.20 |
|
|
|
| 36 |
A |
115 |
110 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27270.9 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 26090.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.500 |
|
|
-0.470 |
| 2 |
H |
0.189 |
|
|
0.139 |
| 3 |
H |
0.193 |
|
|
0.146 |
| 4 |
H |
0.176 |
|
|
0.137 |
| 5 |
C |
-0.243 |
|
|
0.087 |
| 6 |
H |
0.209 |
|
|
0.105 |
| 7 |
Cl |
-0.096 |
|
|
-0.210 |
| 8 |
C |
-0.299 |
|
|
0.043 |
| 9 |
H |
0.170 |
|
|
0.039 |
| 10 |
H |
0.181 |
|
|
0.049 |
| 11 |
C |
-0.513 |
|
|
-0.347 |
| 12 |
H |
0.166 |
|
|
0.087 |
| 13 |
H |
0.195 |
|
|
0.096 |
| 14 |
H |
0.172 |
|
|
0.100 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.862 |
2.101 |
0.371 |
2.301 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.874 |
2.169 |
0.420 |
2.376 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.355 |
-1.360 |
-0.314 |
| y |
-1.360 |
-39.870 |
0.327 |
| z |
-0.314 |
0.327 |
-38.710 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.065 |
-1.360 |
-0.314 |
| y |
-1.360 |
-0.837 |
0.327 |
| z |
-0.314 |
0.327 |
0.902 |
|
| Polar |
|---|
| 3z2-r2 | 1.804 |
|---|
| x2-y2 | 0.515 |
|---|
| xy | -1.360 |
|---|
| xz | -0.314 |
|---|
| yz | 0.327 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
181.934 |
| (<r2>)1/2 |
13.488 |