return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH2CH3 (Butane, 2-chloro-)

using model chemistry: QCISD/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G*
 hartrees
Energy at 0K-616.924237
Energy at 298.15K-616.934249
HF Energy-616.202451
Nuclear repulsion energy233.497855
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3166 3015 19.96      
2 A 3162 3011 23.50      
3 A 3148 2998 24.81      
4 A 3139 2990 33.46      
5 A 3115 2967 14.26      
6 A 3098 2950 2.53      
7 A 3071 2925 18.16      
8 A 3069 2923 23.51      
9 A 3054 2908 18.37      
10 A 1557 1483 6.45      
11 A 1548 1474 9.31      
12 A 1546 1472 0.71      
13 A 1539 1465 6.27      
14 A 1530 1457 0.95      
15 A 1470 1400 4.04      
16 A 1467 1397 7.04      
17 A 1444 1375 0.30      
18 A 1372 1307 24.78      
19 A 1362 1297 1.65      
20 A 1313 1250 9.89      
21 A 1221 1163 5.05      
22 A 1168 1113 1.80      
23 A 1137 1083 3.77      
24 A 1075 1023 1.30      
25 A 1039 990 8.56      
26 A 1001 953 5.11      
27 A 884 842 9.04      
28 A 823 783 9.04      
29 A 650 619 19.03      
30 A 473 450 1.15      
31 A 401 382 1.59      
32 A 337 321 0.98      
33 A 263 251 0.16      
34 A 240 228 0.10      
35 A 227 216 0.87      
36 A 118 112 0.21      

Unscaled Zero Point Vibrational Energy (zpe) 27612.1 cm-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 26295.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G*
ABC
0.15314 0.10346 0.06686

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.335 1.456 -0.008
H2 -1.506 1.462 -1.091
H3 -2.284 1.235 0.491
H4 -1.004 2.457 0.299
C5 -0.269 0.433 0.359
H6 -0.130 0.407 1.446
Cl7 -0.873 -1.222 -0.070
C8 1.068 0.699 -0.326
H9 1.335 1.744 -0.106
H10 0.925 0.633 -1.414
C11 2.204 -0.222 0.119
H12 2.360 -0.156 1.204
H13 1.988 -1.266 -0.131
H14 3.143 0.060 -0.373

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 Cl7 C8 H9 H10 C11 H12 H13 H14
C11.09621.09501.09791.52242.16042.71912.53982.68732.78623.91864.20934.29734.7046
H21.09621.77801.78142.16553.07262.94102.79163.02012.58884.24964.77754.53534.9081
H31.09501.77801.78032.17252.49752.88903.49193.70313.78044.73304.89954.98915.6194
H41.09791.78141.78032.15422.50663.70042.78842.47843.15954.18304.35444.79544.8365
C51.52242.16552.17252.15421.09601.81341.52622.12342.14672.56962.82352.86723.5096
H62.16043.07262.49752.50661.09602.34612.15862.51793.05602.75692.56353.12533.7599
Cl72.71912.94102.88903.70041.81342.34612.74343.69822.91193.24093.63422.86184.2262
C82.53982.79163.49192.78841.52622.15862.74341.10081.09901.52802.17702.17822.1711
H92.68733.02013.70312.47842.12342.51793.69821.10081.76482.16082.52493.08002.4851
H102.78622.58883.78043.15952.14673.05602.91191.09901.76482.17093.08712.52562.5155
C113.91864.24964.73304.18302.56962.75693.24091.52802.16082.17091.09811.09511.0967
H124.20934.77754.89954.35442.82352.56353.63422.17702.52493.08711.09811.77491.7739
H134.29734.53534.98914.79542.86723.12532.86182.17823.08002.52561.09511.77491.7747
H144.70464.90815.61944.83653.50963.75994.22622.17112.48512.51551.09671.77391.7747

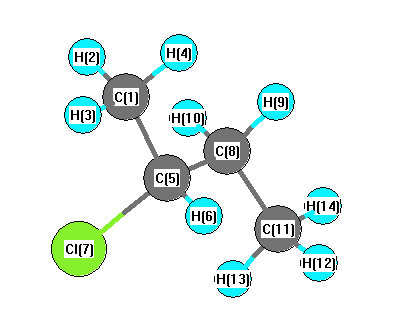
picture of Butane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.129 C1 C5 Cl7 108.887
C1 C5 C8 112.841 H2 C1 H3 108.465
H2 C1 H4 108.555 H2 C1 C5 110.523
H3 C1 H4 108.548 H3 C1 C5 111.155
H4 C1 C5 109.529 C5 C8 H9 106.742
C5 C8 H10 108.625 C5 C8 C11 114.558
H6 C5 Cl7 104.787 H6 C5 C8 109.718
Cl7 C5 C8 110.167 C8 C11 H12 110.925
C8 C11 H13 111.201 C8 C11 H14 110.542
H9 C8 H10 106.690 H9 C8 C11 109.489
H10 C8 C11 110.390 H12 C11 H13 108.059
H12 C11 H14 107.852 H13 C11 H14 108.137
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability