return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH2CH3 (Butane, 2-chloro-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-617.675583
Energy at 298.15K-617.685531
HF Energy-617.458993
Nuclear repulsion energy233.185376
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3182 3021 19.04      
2 A 3177 3016 22.29      
3 A 3162 3001 25.53      
4 A 3155 2995 32.66      
5 A 3131 2972 13.31      
6 A 3112 2954 2.99      
7 A 3086 2929 14.22      
8 A 3085 2929 28.50      
9 A 3066 2910 18.75      
10 A 1557 1478 5.93      
11 A 1548 1470 8.87      
12 A 1546 1468 0.81      
13 A 1537 1459 6.18      
14 A 1525 1447 0.63      
15 A 1462 1388 5.22      
16 A 1459 1385 3.90      
17 A 1432 1360 0.24      
18 A 1362 1293 19.37      
19 A 1355 1286 6.77      
20 A 1302 1236 14.63      
21 A 1210 1148 6.71      
22 A 1155 1096 2.44      
23 A 1126 1069 3.90      
24 A 1059 1006 1.34      
25 A 1031 979 8.99      
26 A 995 944 6.35      
27 A 871 826 9.25      
28 A 817 775 11.07      
29 A 627 595 25.28      
30 A 467 443 1.36      
31 A 395 375 2.34      
32 A 331 314 1.27      
33 A 259 246 0.18      
34 A 238 225 0.09      
35 A 222 210 1.10      
36 A 117 111 0.22      

Unscaled Zero Point Vibrational Energy (zpe) 27579.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 26178.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.15237 0.10321 0.06657

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.315 1.476 -0.013
H2 -1.482 1.481 -1.092
H3 -2.268 1.287 0.485
H4 -0.954 2.463 0.291
C5 -0.270 0.415 0.355
H6 -0.133 0.417 1.439
Cl7 -0.915 -1.206 -0.069
C8 1.092 0.686 -0.313
H9 1.369 1.711 -0.046
H10 0.956 0.654 -1.398
C11 2.238 -0.253 0.110
H12 2.393 -0.208 1.191
H13 2.021 -1.285 -0.162
H14 3.175 0.033 -0.376

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 Cl7 C8 H9 H10 C11 H12 H13 H14
C11.09191.09241.09361.53362.15052.71172.55042.69382.78393.95284.24634.33264.7294
H21.09191.77231.77622.16743.05872.93012.80453.04542.59304.27694.80454.55934.9292
H31.09241.77231.77422.18412.49532.88973.50563.70023.78724.77704.94585.04315.6517
H41.09361.77621.77422.16032.48533.68612.77612.46463.12634.19504.37584.80674.8370
C51.53362.16742.18412.16031.09291.79431.54102.12812.15232.60722.86012.89933.5423
H62.15053.05872.49532.48531.09292.34892.15422.47713.04782.79942.61373.17823.7924
Cl72.71172.93012.88973.68611.79432.34892.76883.70462.95383.29873.67742.93834.2841
C82.55042.80453.50562.77611.54102.15422.76881.09461.09421.54092.18062.18462.1838
H92.69383.04543.70022.46462.12812.47713.70461.09461.76502.15362.50333.06902.4874
H102.78392.59303.78723.12632.15233.04782.95381.09421.76502.17713.08432.53392.5203
C113.95284.27694.77704.19502.60722.79943.29871.54092.15362.17711.09341.08981.0933
H124.24634.80454.94584.37582.86012.61373.67742.18062.50333.08431.09341.76951.7684
H134.33264.55935.04314.80672.89933.17822.93832.18463.06902.53391.08981.76951.7649
H144.72944.92925.65174.83703.54233.79244.28412.18382.48742.52031.09331.76841.7649

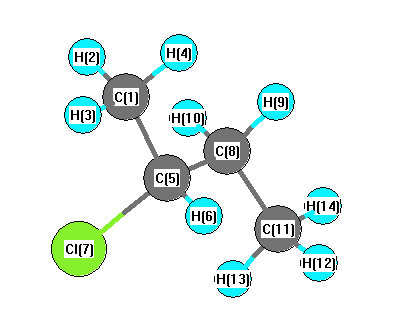
picture of Butane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 108.760 C1 C5 Cl7 108.892
C1 C5 C8 112.096 H2 C1 H3 108.466
H2 C1 H4 108.730 H2 C1 C5 110.151
H3 C1 H4 108.502 H3 C1 C5 111.445
H4 C1 C5 109.487 C5 C8 H9 106.457
C5 C8 H10 108.327 C5 C8 C11 115.550
H6 C5 Cl7 106.335 H6 C5 C8 108.547
Cl7 C5 C8 112.001 C8 C11 H12 110.599
C8 C11 H13 111.132 C8 C11 H14 110.852
H9 C8 H10 107.483 H9 C8 C11 108.414
H10 C8 C11 110.270 H12 C11 H13 108.301
H12 C11 H14 107.942 H13 C11 H14 107.896
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.453      
2 H 0.174      
3 H 0.178      
4 H 0.160      
5 C -0.207      
6 H 0.194      
7 Cl -0.118      
8 C -0.270      
9 H 0.154      
10 H 0.166      
11 C -0.467      
12 H 0.150      
13 H 0.181      
14 H 0.156      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.089 0.347 0.050
y 0.347 8.555 0.222
z 0.050 0.222 6.513


<r2> (average value of r2) Å2
<r2> 182.225
(<r2>)1/2 13.499