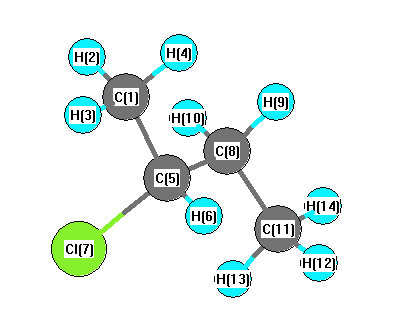
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3088 |
3037 |
16.49 |
|
|
|
| 2 |
A |
3078 |
3027 |
19.30 |
|
|
|
| 3 |
A |
3064 |
3014 |
25.91 |
|
|
|
| 4 |
A |
3061 |
3011 |
25.51 |
|
|
|
| 5 |
A |
3024 |
2975 |
19.19 |
|
|
|
| 6 |
A |
3007 |
2958 |
0.73 |
|
|
|
| 7 |
A |
2989 |
2940 |
18.04 |
|
|
|
| 8 |
A |
2988 |
2939 |
24.51 |
|
|
|
| 9 |
A |
2963 |
2915 |
19.70 |
|
|
|
| 10 |
A |
1489 |
1464 |
6.73 |
|
|
|
| 11 |
A |
1480 |
1455 |
10.56 |
|
|
|
| 12 |
A |
1478 |
1453 |
0.66 |
|
|
|
| 13 |
A |
1467 |
1443 |
7.20 |
|
|
|
| 14 |
A |
1451 |
1427 |
0.76 |
|
|
|
| 15 |
A |
1390 |
1368 |
7.44 |
|
|
|
| 16 |
A |
1387 |
1364 |
4.95 |
|
|
|
| 17 |
A |
1363 |
1341 |
0.41 |
|
|
|
| 18 |
A |
1296 |
1275 |
16.29 |
|
|
|
| 19 |
A |
1289 |
1268 |
6.23 |
|
|
|
| 20 |
A |
1237 |
1216 |
14.67 |
|
|
|
| 21 |
A |
1155 |
1136 |
8.68 |
|
|
|
| 22 |
A |
1112 |
1093 |
3.10 |
|
|
|
| 23 |
A |
1073 |
1055 |
3.56 |
|
|
|
| 24 |
A |
1027 |
1010 |
2.13 |
|
|
|
| 25 |
A |
983 |
967 |
11.24 |
|
|
|
| 26 |
A |
953 |
937 |
7.91 |
|
|
|
| 27 |
A |
837 |
823 |
9.15 |
|
|
|
| 28 |
A |
779 |
766 |
12.86 |
|
|
|
| 29 |
A |
594 |
584 |
25.56 |
|
|
|
| 30 |
A |
450 |
443 |
1.41 |
|
|
|
| 31 |
A |
379 |
373 |
2.73 |
|
|
|
| 32 |
A |
319 |
313 |
1.50 |
|
|
|
| 33 |
A |
251 |
247 |
0.23 |
|
|
|
| 34 |
A |
229 |
226 |
0.09 |
|
|
|
| 35 |
A |
213 |
209 |
1.31 |
|
|
|
| 36 |
A |
116 |
114 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26529.6 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 26091.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.472 |
|
|
|
| 2 |
H |
0.179 |
|
|
|
| 3 |
H |
0.182 |
|
|
|
| 4 |
H |
0.166 |
|
|
|
| 5 |
C |
-0.227 |
|
|
|
| 6 |
H |
0.194 |
|
|
|
| 7 |
Cl |
-0.094 |
|
|
|
| 8 |
C |
-0.272 |
|
|
|
| 9 |
H |
0.159 |
|
|
|
| 10 |
H |
0.169 |
|
|
|
| 11 |
C |
-0.487 |
|
|
|
| 12 |
H |
0.157 |
|
|
|
| 13 |
H |
0.185 |
|
|
|
| 14 |
H |
0.162 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.841 |
2.069 |
0.360 |
2.262 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-39.453 |
-1.338 |
-0.305 |
| y |
-1.338 |
-39.959 |
0.317 |
| z |
-0.305 |
0.317 |
-38.902 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.022 |
-1.338 |
-0.305 |
| y |
-1.338 |
-0.782 |
0.317 |
| z |
-0.305 |
0.317 |
0.804 |
|
| Polar |
|---|
| 3z2-r2 | 1.607 |
|---|
| x2-y2 | 0.506 |
|---|
| xy | -1.338 |
|---|
| xz | -0.305 |
|---|
| yz | 0.317 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.580 |
0.295 |
0.037 |
| y |
0.295 |
8.928 |
0.193 |
| z |
0.037 |
0.193 |
6.726 |
<r2> (average value of r
2) Å
2
| <r2> |
183.547 |
| (<r2>)1/2 |
13.548 |