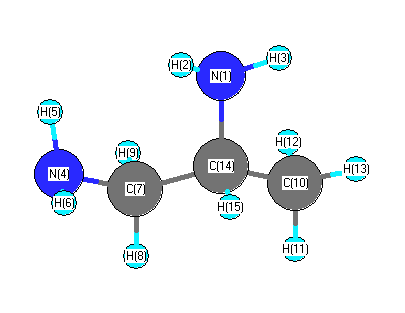
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3588 |
3433 |
0.02 |
|
|
|
| 2 |
A |
3575 |
3420 |
0.43 |
|
|
|
| 3 |
A |
3490 |
3339 |
2.85 |
|
|
|
| 4 |
A |
3489 |
3338 |
1.37 |
|
|
|
| 5 |
A |
3141 |
3005 |
28.89 |
|
|
|
| 6 |
A |
3125 |
2989 |
32.34 |
|
|
|
| 7 |
A |
3105 |
2971 |
28.54 |
|
|
|
| 8 |
A |
3055 |
2923 |
44.73 |
|
|
|
| 9 |
A |
3050 |
2918 |
19.96 |
|
|
|
| 10 |
A |
2918 |
2792 |
91.11 |
|
|
|
| 11 |
A |
1699 |
1625 |
4.30 |
|
|
|
| 12 |
A |
1688 |
1615 |
60.65 |
|
|
|
| 13 |
A |
1527 |
1461 |
3.09 |
|
|
|
| 14 |
A |
1515 |
1450 |
5.27 |
|
|
|
| 15 |
A |
1498 |
1433 |
0.79 |
|
|
|
| 16 |
A |
1437 |
1374 |
4.05 |
|
|
|
| 17 |
A |
1426 |
1364 |
11.93 |
|
|
|
| 18 |
A |
1413 |
1352 |
4.34 |
|
|
|
| 19 |
A |
1391 |
1331 |
10.51 |
|
|
|
| 20 |
A |
1355 |
1297 |
10.85 |
|
|
|
| 21 |
A |
1279 |
1223 |
1.16 |
|
|
|
| 22 |
A |
1220 |
1167 |
5.42 |
|
|
|
| 23 |
A |
1184 |
1132 |
5.05 |
|
|
|
| 24 |
A |
1131 |
1082 |
10.82 |
|
|
|
| 25 |
A |
1062 |
1016 |
13.49 |
|
|
|
| 26 |
A |
1033 |
988 |
1.33 |
|
|
|
| 27 |
A |
968 |
926 |
13.07 |
|
|
|
| 28 |
A |
921 |
881 |
109.01 |
|
|
|
| 29 |
A |
896 |
857 |
91.63 |
|
|
|
| 30 |
A |
852 |
815 |
89.87 |
|
|
|
| 31 |
A |
806 |
771 |
68.11 |
|
|
|
| 32 |
A |
495 |
473 |
3.09 |
|
|
|
| 33 |
A |
472 |
451 |
24.82 |
|
|
|
| 34 |
A |
371 |
355 |
57.32 |
|
|
|
| 35 |
A |
366 |
350 |
1.01 |
|
|
|
| 36 |
A |
257 |
245 |
8.87 |
|
|
|
| 37 |
A |
246 |
235 |
38.33 |
|
|
|
| 38 |
A |
223 |
214 |
24.00 |
|
|
|
| 39 |
A |
120 |
114 |
6.38 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30691.4 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 29362.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.746 |
|
|
-0.932 |
| 2 |
H |
0.299 |
|
|
0.310 |
| 3 |
H |
0.313 |
|
|
0.369 |
| 4 |
N |
-0.747 |
|
|
-0.982 |
| 5 |
H |
0.318 |
|
|
0.380 |
| 6 |
H |
0.306 |
|
|
0.351 |
| 7 |
C |
-0.170 |
|
|
0.311 |
| 8 |
H |
0.159 |
|
|
0.000 |
| 9 |
H |
0.173 |
|
|
0.022 |
| 10 |
C |
-0.513 |
|
|
-0.488 |
| 11 |
H |
0.168 |
|
|
0.093 |
| 12 |
H |
0.182 |
|
|
0.135 |
| 13 |
H |
0.160 |
|
|
0.141 |
| 14 |
C |
-0.022 |
|
|
0.319 |
| 15 |
H |
0.123 |
|
|
-0.029 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.014 |
1.206 |
1.770 |
2.370 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.004 |
1.204 |
1.735 |
2.338 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
143.449 |
| (<r2>)1/2 |
11.977 |