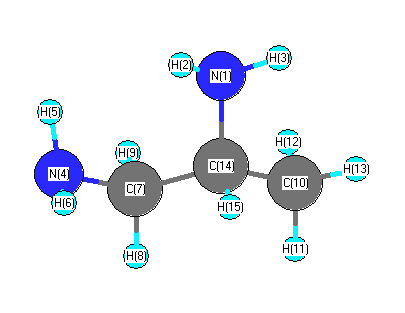
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3606 |
3430 |
0.02 |
|
|
|
| 2 |
A |
3595 |
3419 |
0.34 |
|
|
|
| 3 |
A |
3509 |
3337 |
2.18 |
|
|
|
| 4 |
A |
3506 |
3334 |
1.06 |
|
|
|
| 5 |
A |
3156 |
3001 |
26.90 |
|
|
|
| 6 |
A |
3140 |
2986 |
30.81 |
|
|
|
| 7 |
A |
3118 |
2965 |
28.20 |
|
|
|
| 8 |
A |
3068 |
2917 |
41.20 |
|
|
|
| 9 |
A |
3062 |
2912 |
20.46 |
|
|
|
| 10 |
A |
2932 |
2788 |
88.74 |
|
|
|
| 11 |
A |
1704 |
1621 |
4.40 |
|
|
|
| 12 |
A |
1692 |
1609 |
62.76 |
|
|
|
| 13 |
A |
1529 |
1454 |
3.01 |
|
|
|
| 14 |
A |
1517 |
1443 |
5.58 |
|
|
|
| 15 |
A |
1499 |
1426 |
0.80 |
|
|
|
| 16 |
A |
1444 |
1373 |
2.81 |
|
|
|
| 17 |
A |
1429 |
1359 |
15.43 |
|
|
|
| 18 |
A |
1417 |
1347 |
3.93 |
|
|
|
| 19 |
A |
1396 |
1328 |
9.91 |
|
|
|
| 20 |
A |
1358 |
1292 |
10.42 |
|
|
|
| 21 |
A |
1283 |
1220 |
1.23 |
|
|
|
| 22 |
A |
1226 |
1166 |
5.12 |
|
|
|
| 23 |
A |
1193 |
1135 |
6.10 |
|
|
|
| 24 |
A |
1143 |
1087 |
10.99 |
|
|
|
| 25 |
A |
1066 |
1014 |
13.99 |
|
|
|
| 26 |
A |
1042 |
991 |
1.42 |
|
|
|
| 27 |
A |
975 |
927 |
11.76 |
|
|
|
| 28 |
A |
926 |
881 |
116.13 |
|
|
|
| 29 |
A |
901 |
857 |
86.44 |
|
|
|
| 30 |
A |
857 |
815 |
85.28 |
|
|
|
| 31 |
A |
814 |
774 |
71.11 |
|
|
|
| 32 |
A |
499 |
475 |
3.11 |
|
|
|
| 33 |
A |
474 |
451 |
24.41 |
|
|
|
| 34 |
A |
374 |
356 |
58.70 |
|
|
|
| 35 |
A |
369 |
351 |
1.03 |
|
|
|
| 36 |
A |
259 |
246 |
6.86 |
|
|
|
| 37 |
A |
249 |
236 |
37.85 |
|
|
|
| 38 |
A |
226 |
215 |
26.66 |
|
|
|
| 39 |
A |
122 |
116 |
6.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30838.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 29326.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.749 |
|
|
|
| 2 |
H |
0.300 |
|
|
|
| 3 |
H |
0.316 |
|
|
|
| 4 |
N |
-0.750 |
|
|
|
| 5 |
H |
0.319 |
|
|
|
| 6 |
H |
0.308 |
|
|
|
| 7 |
C |
-0.173 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.174 |
|
|
|
| 10 |
C |
-0.517 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.183 |
|
|
|
| 13 |
H |
0.162 |
|
|
|
| 14 |
C |
-0.029 |
|
|
|
| 15 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.020 |
1.217 |
1.762 |
2.372 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.597 |
-2.826 |
-0.917 |
| y |
-2.826 |
-30.739 |
1.783 |
| z |
-0.917 |
1.783 |
-33.124 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.666 |
-2.826 |
-0.917 |
| y |
-2.826 |
3.621 |
1.783 |
| z |
-0.917 |
1.783 |
0.044 |
|
| Polar |
|---|
| 3z2-r2 | 0.088 |
|---|
| x2-y2 | -4.858 |
|---|
| xy | -2.826 |
|---|
| xz | -0.917 |
|---|
| yz | 1.783 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.595 |
-0.231 |
-0.117 |
| y |
-0.231 |
7.041 |
-0.087 |
| z |
-0.117 |
-0.087 |
6.159 |
<r2> (average value of r
2) Å
2
| <r2> |
142.739 |
| (<r2>)1/2 |
11.947 |