return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H10N2 (1,2-Diaminopropane)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-229.071986
Energy at 298.15K-229.084662
HF Energy-228.302362
Nuclear repulsion energy195.141085
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3619 3397 0.03      
2 A 3608 3387 0.20      
3 A 3526 3310 0.77      
4 A 3521 3305 0.61      
5 A 3188 2992 25.13      
6 A 3172 2978 28.50      
7 A 3154 2960 24.52      
8 A 3102 2911 37.46      
9 A 3096 2906 12.58      
10 A 2991 2807 69.95      
11 A 1737 1631 9.78      
12 A 1725 1619 60.67      
13 A 1562 1466 3.65      
14 A 1553 1458 5.33      
15 A 1542 1447 0.68      
16 A 1483 1392 2.41      
17 A 1465 1375 13.45      
18 A 1448 1359 8.54      
19 A 1427 1339 3.68      
20 A 1388 1303 9.70      
21 A 1307 1227 0.79      
22 A 1257 1180 5.86      
23 A 1213 1139 6.82      
24 A 1156 1085 7.05      
25 A 1087 1020 15.99      
26 A 1065 1000 2.87      
27 A 1000 939 19.70      
28 A 967 908 150.35      
29 A 933 875 89.61      
30 A 889 834 58.39      
31 A 835 784 41.01      
32 A 508 477 2.31      
33 A 486 456 22.83      
34 A 382 359 8.47      
35 A 373 350 51.22      
36 A 267 251 5.77      
37 A 258 242 34.37      
38 A 235 220 30.95      
39 A 125 117 6.93      

Unscaled Zero Point Vibrational Energy (zpe) 31324.3 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 29401.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.26636 0.11697 0.08929

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.472 1.385 -0.222
H2 -0.286 1.926 0.188
H3 1.329 1.862 0.052
N4 -2.052 -0.133 0.024
H5 -2.111 0.754 -0.471
H6 -2.177 0.067 1.015
C7 -0.739 -0.737 -0.209
H8 -0.747 -1.747 0.219
H9 -0.611 -0.841 -1.293
C10 1.769 -0.672 -0.032
H11 1.776 -1.705 0.330
H12 1.884 -0.678 -1.120
H13 2.634 -0.157 0.401
C14 0.467 0.028 0.345
H15 0.382 0.037 1.448

Atom - Atom Distances (Å)
  N1 H2 H3 N4 H5 H6 C7 H8 H9 C10 H11 H12 H13 C14 H15
N11.01781.01852.95572.67063.20732.44273.38922.69702.43853.39832.65582.72741.47042.1477
H21.01781.62242.71722.26632.77772.72973.70133.15493.31944.17763.63303.59322.04802.3669
H31.01851.62243.92633.65184.05563.33164.16713.58882.57333.60572.85202.42962.04792.4856
N42.95572.71723.92631.01721.01871.46442.08462.07673.85934.14954.13494.70142.54512.8257
H52.67062.26633.65181.01721.63832.04302.93072.33904.15674.66834.29294.90962.80013.2268
H63.20732.77774.05561.01871.63832.05282.44282.93304.14884.38564.64804.85542.72872.5963
C72.44272.72973.33161.46442.04302.05281.09701.09572.51462.74772.77663.47601.53172.1450
H83.38923.70134.16712.08462.93072.44281.09701.76712.74752.52573.13963.74052.15412.4427
H92.69703.15493.58882.07672.33902.93301.09571.76712.69823.01222.50563.72342.14433.0441
C102.43853.31942.57333.85934.15674.14882.51462.74752.69821.09461.09451.09571.52462.1477
H113.39834.17763.60574.14954.66834.38562.74772.52573.01221.09461.78031.77102.17112.4945
H122.65583.63302.85204.13494.29294.64802.77663.13962.50561.09451.78031.77432.15623.0591
H132.72743.59322.42964.70144.90964.85543.47603.74053.72341.09571.77101.77432.17502.4903
C141.47042.04802.04792.54512.80012.72871.53172.15412.14431.52462.17112.15622.17501.1064
H152.14772.36692.48562.82573.22682.59632.14502.44273.04412.14772.49453.05912.49031.1064

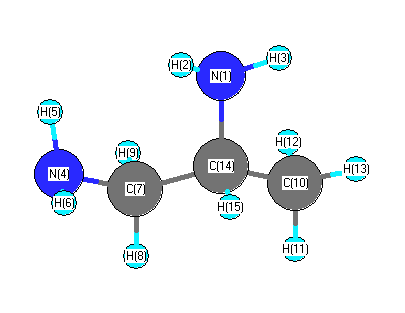
picture of 1,2-Diaminopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 C7 108.896 N1 C14 C10 109.002
N1 C14 H15 112.146 H2 N1 H3 105.636
H2 N1 C14 109.446 H3 N1 C14 109.389
N4 C7 H8 108.091 N4 C7 H9 107.545
N4 C7 C14 116.288 H5 N4 H6 107.170
H5 N4 C7 109.505 H6 N4 C7 110.230
C7 C14 C10 110.719 C7 C14 H15 107.704
H8 C7 H9 107.397 H8 C7 C14 108.937
H9 C7 C14 108.251 C10 C14 H15 108.379
H11 C10 H12 108.833 H11 C10 H13 107.917
H11 C10 C14 110.905 H12 C10 H13 108.219
H12 C10 C14 109.730 H13 C10 C14 111.156
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability