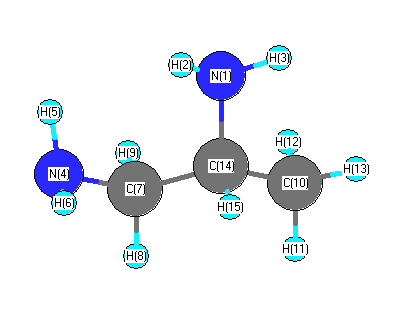
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3464 |
3407 |
0.75 |
|
|
|
| 2 |
A |
3452 |
3395 |
1.24 |
|
|
|
| 3 |
A |
3366 |
3310 |
9.11 |
|
|
|
| 4 |
A |
3365 |
3309 |
1.63 |
|
|
|
| 5 |
A |
3066 |
3016 |
26.20 |
|
|
|
| 6 |
A |
3050 |
3000 |
30.14 |
|
|
|
| 7 |
A |
3025 |
2975 |
27.79 |
|
|
|
| 8 |
A |
2980 |
2931 |
33.86 |
|
|
|
| 9 |
A |
2972 |
2923 |
29.52 |
|
|
|
| 10 |
A |
2828 |
2781 |
93.05 |
|
|
|
| 11 |
A |
1648 |
1621 |
1.09 |
|
|
|
| 12 |
A |
1636 |
1609 |
55.62 |
|
|
|
| 13 |
A |
1484 |
1459 |
3.08 |
|
|
|
| 14 |
A |
1471 |
1447 |
4.87 |
|
|
|
| 15 |
A |
1450 |
1426 |
0.70 |
|
|
|
| 16 |
A |
1388 |
1365 |
6.18 |
|
|
|
| 17 |
A |
1379 |
1356 |
8.70 |
|
|
|
| 18 |
A |
1372 |
1349 |
1.89 |
|
|
|
| 19 |
A |
1348 |
1325 |
12.85 |
|
|
|
| 20 |
A |
1313 |
1291 |
11.71 |
|
|
|
| 21 |
A |
1245 |
1224 |
1.62 |
|
|
|
| 22 |
A |
1185 |
1165 |
4.65 |
|
|
|
| 23 |
A |
1146 |
1127 |
3.44 |
|
|
|
| 24 |
A |
1097 |
1079 |
7.31 |
|
|
|
| 25 |
A |
1033 |
1016 |
13.06 |
|
|
|
| 26 |
A |
1007 |
990 |
1.56 |
|
|
|
| 27 |
A |
943 |
928 |
18.53 |
|
|
|
| 28 |
A |
902 |
887 |
126.40 |
|
|
|
| 29 |
A |
876 |
862 |
74.20 |
|
|
|
| 30 |
A |
829 |
815 |
79.93 |
|
|
|
| 31 |
A |
788 |
775 |
45.74 |
|
|
|
| 32 |
A |
483 |
475 |
5.11 |
|
|
|
| 33 |
A |
463 |
455 |
25.00 |
|
|
|
| 34 |
A |
373 |
367 |
50.88 |
|
|
|
| 35 |
A |
357 |
351 |
1.44 |
|
|
|
| 36 |
A |
254 |
249 |
13.76 |
|
|
|
| 37 |
A |
242 |
238 |
33.46 |
|
|
|
| 38 |
A |
222 |
218 |
23.47 |
|
|
|
| 39 |
A |
115 |
113 |
5.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29806.7 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 29314.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.696 |
|
|
|
| 2 |
H |
0.279 |
|
|
|
| 3 |
H |
0.296 |
|
|
|
| 4 |
N |
-0.695 |
|
|
|
| 5 |
H |
0.301 |
|
|
|
| 6 |
H |
0.290 |
|
|
|
| 7 |
C |
-0.168 |
|
|
|
| 8 |
H |
0.147 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
| 10 |
C |
-0.485 |
|
|
|
| 11 |
H |
0.158 |
|
|
|
| 12 |
H |
0.171 |
|
|
|
| 13 |
H |
0.150 |
|
|
|
| 14 |
C |
-0.016 |
|
|
|
| 15 |
H |
0.109 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.964 |
1.188 |
1.736 |
2.314 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.714 |
-2.787 |
-0.862 |
| y |
-2.787 |
-30.940 |
1.815 |
| z |
-0.862 |
1.815 |
-33.305 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.591 |
-2.787 |
-0.862 |
| y |
-2.787 |
3.570 |
1.815 |
| z |
-0.862 |
1.815 |
0.022 |
|
| Polar |
|---|
| 3z2-r2 | 0.043 |
|---|
| x2-y2 | -4.774 |
|---|
| xy | -2.787 |
|---|
| xz | -0.862 |
|---|
| yz | 1.815 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.036 |
-0.221 |
-0.106 |
| y |
-0.221 |
7.384 |
-0.070 |
| z |
-0.106 |
-0.070 |
6.369 |
<r2> (average value of r
2) Å
2
| <r2> |
144.760 |
| (<r2>)1/2 |
12.032 |