Jump to
S1C2
Vibrational Frequencies calculated at QCISD/6-31G*
Geometric Data calculated at QCISD/6-31G*
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/6-31G*
| | hartrees |
|---|
| Energy at 0K | -231.740547 |
| Energy at 298.15K | -231.748845 |
| Nuclear repulsion energy | 175.595642 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3186 |
3034 |
9.70 |
|
|
|
| 2 |
A |
3166 |
3015 |
22.84 |
|
|
|
| 3 |
A |
3156 |
3005 |
22.86 |
|
|
|
| 4 |
A |
3137 |
2988 |
12.24 |
|
|
|
| 5 |
A |
3084 |
2937 |
10.81 |
|
|
|
| 6 |
A |
3081 |
2934 |
19.81 |
|
|
|
| 7 |
A |
3070 |
2923 |
8.49 |
|
|
|
| 8 |
A |
3050 |
2904 |
14.56 |
|
|
|
| 9 |
A |
1820 |
1733 |
105.11 |
|
|
|
| 10 |
A |
1548 |
1474 |
9.06 |
|
|
|
| 11 |
A |
1545 |
1471 |
5.85 |
|
|
|
| 12 |
A |
1526 |
1453 |
10.54 |
|
|
|
| 13 |
A |
1524 |
1452 |
17.85 |
|
|
|
| 14 |
A |
1517 |
1444 |
0.15 |
|
|
|
| 15 |
A |
1470 |
1400 |
4.90 |
|
|
|
| 16 |
A |
1447 |
1378 |
48.38 |
|
|
|
| 17 |
A |
1423 |
1355 |
7.49 |
|
|
|
| 18 |
A |
1318 |
1255 |
0.16 |
|
|
|
| 19 |
A |
1233 |
1174 |
59.81 |
|
|
|
| 20 |
A |
1168 |
1113 |
0.19 |
|
|
|
| 21 |
A |
1140 |
1086 |
1.48 |
|
|
|
| 22 |
A |
1030 |
981 |
3.49 |
|
|
|
| 23 |
A |
984 |
937 |
3.01 |
|
|
|
| 24 |
A |
979 |
932 |
5.32 |
|
|
|
| 25 |
A |
789 |
751 |
1.57 |
|
|
|
| 26 |
A |
784 |
746 |
2.94 |
|
|
|
| 27 |
A |
599 |
571 |
10.83 |
|
|
|
| 28 |
A |
478 |
455 |
0.25 |
|
|
|
| 29 |
A |
409 |
390 |
4.36 |
|
|
|
| 30 |
A |
257 |
245 |
4.95 |
|
|
|
| 31 |
A |
217 |
207 |
0.15 |
|
|
|
| 32 |
A |
108 |
103 |
0.04 |
|
|
|
| 33 |
A |
39 |
37 |
0.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25140.0 cm
-1
Scaled (by 0.9523) Zero Point Vibrational Energy (zpe) 23940.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.892 |
-0.497 |
0.000 |
| C2 |
-0.525 |
0.164 |
0.000 |
| C3 |
0.681 |
-0.764 |
-0.000 |
| C4 |
2.012 |
-0.020 |
0.000 |
| O5 |
-0.402 |
1.381 |
-0.000 |
| H6 |
-2.669 |
0.271 |
-0.002 |
| H7 |
-2.005 |
-1.140 |
-0.883 |
| H8 |
-2.006 |
-1.136 |
0.885 |
| H9 |
2.849 |
-0.728 |
0.001 |
| H10 |
2.098 |
0.622 |
-0.883 |
| H11 |
2.098 |
0.623 |
0.883 |
| H12 |
0.597 |
-1.428 |
0.875 |
| H13 |
0.597 |
-1.427 |
-0.876 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5178 | 2.5862 | 3.9327 | 2.3973 | 1.0933 | 1.0979 | 1.0979 | 4.7467 | 4.2371 | 4.2368 | 2.7974 | 2.7973 |
C2 | 1.5178 | | 1.5219 | 2.5441 | 1.2234 | 2.1463 | 2.1606 | 2.1605 | 3.4907 | 2.8061 | 2.8056 | 2.1351 | 2.1348 | C3 | 2.5862 | 1.5219 | | 1.5251 | 2.4029 | 3.5062 | 2.8519 | 2.8538 | 2.1690 | 2.1703 | 2.1704 | 1.1012 | 1.1012 | C4 | 3.9327 | 2.5441 | 1.5251 | | 2.7911 | 4.6902 | 4.2625 | 4.2637 | 1.0965 | 1.0951 | 1.0951 | 2.1790 | 2.1789 | O5 | 2.3973 | 1.2234 | 2.4029 | 2.7911 | | 2.5241 | 3.1150 | 3.1137 | 3.8754 | 2.7582 | 2.7575 | 3.1068 | 3.1062 | H6 | 1.0933 | 2.1463 | 3.5062 | 4.6902 | 2.5241 | | 1.7912 | 1.7912 | 5.6082 | 4.8608 | 4.8612 | 3.7848 | 3.7835 | H7 | 1.0979 | 2.1606 | 2.8519 | 4.2625 | 3.1150 | 1.7912 | | 1.7686 | 4.9512 | 4.4654 | 4.8017 | 3.1535 | 2.6178 | H8 | 1.0979 | 2.1605 | 2.8538 | 4.2637 | 3.1137 | 1.7912 | 1.7686 | | 4.9528 | 4.8030 | 4.4655 | 2.6201 | 3.1568 | H9 | 4.7467 | 3.4907 | 2.1690 | 1.0965 | 3.8754 | 5.6082 | 4.9512 | 4.9528 | | 1.7794 | 1.7794 | 2.5152 | 2.5157 | H10 | 4.2371 | 2.8061 | 2.1703 | 1.0951 | 2.7582 | 4.8608 | 4.4654 | 4.8030 | 1.7794 | | 1.7656 | 3.0893 | 2.5397 | H11 | 4.2368 | 2.8056 | 2.1704 | 1.0951 | 2.7575 | 4.8612 | 4.8017 | 4.4655 | 1.7794 | 1.7656 | | 2.5407 | 3.0892 | H12 | 2.7974 | 2.1351 | 1.1012 | 2.1790 | 3.1068 | 3.7848 | 3.1535 | 2.6201 | 2.5152 | 3.0893 | 2.5407 | | 1.7506 | H13 | 2.7973 | 2.1348 | 1.1012 | 2.1789 | 3.1062 | 3.7835 | 2.6178 | 3.1568 | 2.5157 | 2.5397 | 3.0892 | 1.7506 | |
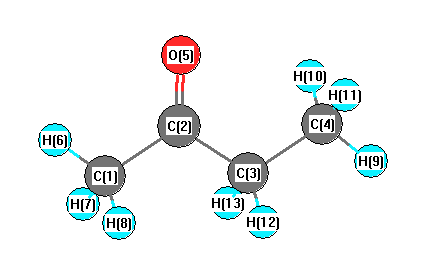 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
116.603 |
|
C1 |
C2 |
O5 |
121.618 |
| C2 |
C1 |
H6 |
109.501 |
|
C2 |
C1 |
H7 |
110.348 |
| C2 |
C1 |
H8 |
110.343 |
|
C2 |
C3 |
C4 |
113.223 |
| C2 |
C3 |
H12 |
107.892 |
|
C2 |
C3 |
H13 |
107.869 |
| C3 |
C2 |
O5 |
121.779 |
|
C3 |
C4 |
H9 |
110.587 |
| C3 |
C4 |
H10 |
110.781 |
|
C3 |
C4 |
H11 |
110.784 |
| C4 |
C3 |
H12 |
111.105 |
|
C4 |
C3 |
H13 |
111.090 |
| H6 |
C1 |
H7 |
109.659 |
|
H6 |
C1 |
H8 |
109.660 |
| H7 |
C1 |
H8 |
107.301 |
|
H9 |
C4 |
H10 |
108.573 |
| H9 |
C4 |
H11 |
108.572 |
|
H10 |
C4 |
H11 |
107.445 |
| H12 |
C3 |
H13 |
105.278 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
