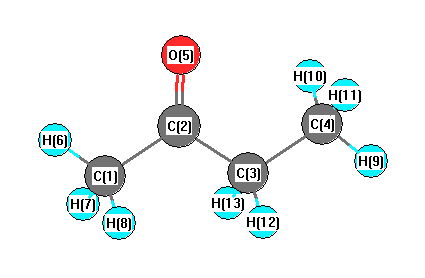
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3199 |
3040 |
7.77 |
|
|
|
| 2 |
A' |
3169 |
3011 |
19.19 |
|
|
|
| 3 |
A' |
3089 |
2935 |
19.74 |
|
|
|
| 4 |
A' |
3074 |
2921 |
6.06 |
|
|
|
| 5 |
A' |
3053 |
2901 |
15.50 |
|
|
|
| 6 |
A' |
1855 |
1763 |
135.97 |
|
|
|
| 7 |
A' |
1525 |
1449 |
10.07 |
|
|
|
| 8 |
A' |
1493 |
1418 |
19.78 |
|
|
|
| 9 |
A' |
1478 |
1404 |
3.08 |
|
|
|
| 10 |
A' |
1438 |
1367 |
6.91 |
|
|
|
| 11 |
A' |
1415 |
1345 |
63.22 |
|
|
|
| 12 |
A' |
1391 |
1322 |
9.84 |
|
|
|
| 13 |
A' |
1209 |
1149 |
68.03 |
|
|
|
| 14 |
A' |
1127 |
1071 |
1.04 |
|
|
|
| 15 |
A' |
1023 |
972 |
5.10 |
|
|
|
| 16 |
A' |
956 |
909 |
6.02 |
|
|
|
| 17 |
A' |
782 |
743 |
2.74 |
|
|
|
| 18 |
A' |
595 |
566 |
8.62 |
|
|
|
| 19 |
A' |
403 |
383 |
4.21 |
|
|
|
| 20 |
A' |
248 |
236 |
5.27 |
|
|
|
| 21 |
A" |
3175 |
3017 |
18.36 |
|
|
|
| 22 |
A" |
3143 |
2987 |
8.33 |
|
|
|
| 23 |
A" |
3087 |
2934 |
8.64 |
|
|
|
| 24 |
A" |
1521 |
1445 |
7.89 |
|
|
|
| 25 |
A" |
1502 |
1428 |
12.02 |
|
|
|
| 26 |
A" |
1295 |
1231 |
0.01 |
|
|
|
| 27 |
A" |
1143 |
1086 |
0.50 |
|
|
|
| 28 |
A" |
958 |
910 |
5.19 |
|
|
|
| 29 |
A" |
760 |
722 |
3.78 |
|
|
|
| 30 |
A" |
475 |
452 |
0.00 |
|
|
|
| 31 |
A" |
204 |
194 |
0.18 |
|
|
|
| 32 |
A" |
106 |
101 |
0.00 |
|
|
|
| 33 |
A" |
25i |
24i |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24931.8 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 23692.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.611 |
|
|
|
| 2 |
C |
0.456 |
|
|
|
| 3 |
C |
-0.421 |
|
|
|
| 4 |
C |
-0.515 |
|
|
|
| 5 |
O |
-0.433 |
|
|
|
| 6 |
H |
0.209 |
|
|
|
| 7 |
H |
0.194 |
|
|
|
| 8 |
H |
0.194 |
|
|
|
| 9 |
H |
0.169 |
|
|
|
| 10 |
H |
0.190 |
|
|
|
| 11 |
H |
0.190 |
|
|
|
| 12 |
H |
0.189 |
|
|
|
| 13 |
H |
0.189 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.661 |
-0.591 |
0.000 |
2.726 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.366 |
2.287 |
0.000 |
| y |
2.287 |
-30.743 |
0.000 |
| z |
0.000 |
0.000 |
-30.165 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.912 |
2.287 |
0.000 |
| y |
2.287 |
1.023 |
0.000 |
| z |
0.000 |
0.000 |
1.889 |
|
| Polar |
|---|
| 3z2-r2 | 3.778 |
|---|
| x2-y2 | -2.623 |
|---|
| xy | 2.287 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.058 |
0.221 |
0.000 |
| y |
0.221 |
7.189 |
0.000 |
| z |
0.000 |
0.000 |
5.314 |
<r2> (average value of r
2) Å
2
| <r2> |
136.226 |
| (<r2>)1/2 |
11.672 |