return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COCH2CH3 (2-Butanone)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-232.198360
Energy at 298.15K-232.206574
HF Energy-231.968107
Nuclear repulsion energy175.772153
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3201 3038 9.60      
2 A' 3172 3011 22.47      
3 A' 3098 2941 20.82      
4 A' 3083 2926 7.55      
5 A' 3061 2905 15.14      
6 A' 1800 1708 107.90      
7 A' 1549 1470 7.89      
8 A' 1518 1441 18.12      
9 A' 1506 1430 2.27      
10 A' 1461 1387 4.88      
11 A' 1433 1360 39.91      
12 A' 1409 1337 11.98      
13 A' 1218 1156 70.86      
14 A' 1132 1075 1.69      
15 A' 1021 969 2.80      
16 A' 965 916 8.78      
17 A' 777 737 2.26      
18 A' 593 563 9.14      
19 A' 405 384 4.21      
20 A' 252 240 4.99      
21 A" 3181 3019 22.13      
22 A" 3147 2987 11.45      
23 A" 3093 2936 10.72      
24 A" 1545 1467 5.87      
25 A" 1527 1449 10.19      
26 A" 1312 1246 0.10      
27 A" 1159 1101 0.29      
28 A" 976 926 3.57      
29 A" 775 736 3.13      
30 A" 476 451 0.07      
31 A" 210 200 0.17      
32 A" 108 102 0.00      
33 A" 19 18 0.10      

Unscaled Zero Point Vibrational Energy (zpe) 25089.5 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 23815.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.31726 0.11980 0.09135

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.042 1.654 0.000
C2 0.000 0.551 0.000
C3 0.523 -0.878 0.000
C4 -0.584 -1.926 0.000
O5 -1.195 0.799 0.000
H6 0.547 2.626 0.000
H7 1.687 1.571 0.881
H8 1.687 1.571 -0.881
H9 -0.162 -2.934 0.000
H10 -1.222 -1.816 0.880
H11 -1.222 -1.816 -0.880
H12 1.180 -0.997 -0.872
H13 1.180 -0.997 0.872

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13
C11.51752.58533.93202.39511.08991.09491.09494.74344.23574.23572.79422.7942
C21.51751.52212.54471.22072.14562.15922.15923.48862.80532.80532.13262.1326
C32.58531.52211.52412.40113.50412.85122.85122.16662.16732.16731.09811.0981
C43.93202.54471.52412.79224.68994.26154.26151.09291.09181.09182.17612.1761
O52.39511.22072.40112.79222.52463.11133.11133.87302.75912.75913.10263.1026
H61.08992.14563.50414.68992.52461.78531.78535.60454.86134.86133.77923.7792
H71.09492.15922.85124.26153.11131.78531.76114.94844.46474.79913.14982.6173
H81.09492.15922.85124.26153.11131.78531.76114.94844.79914.46472.61733.1498
H94.74343.48862.16661.09293.87305.60454.94844.94841.77361.77362.51262.5126
H104.23572.80532.16731.09182.75914.86134.46474.79911.77361.75913.08332.5376
H114.23572.80532.16731.09182.75914.86134.79914.46471.77361.75912.53763.0833
H122.79422.13261.09812.17613.10263.77923.14982.61732.51263.08332.53761.7438
H132.79422.13261.09812.17613.10263.77922.61733.14982.51262.53763.08331.7438

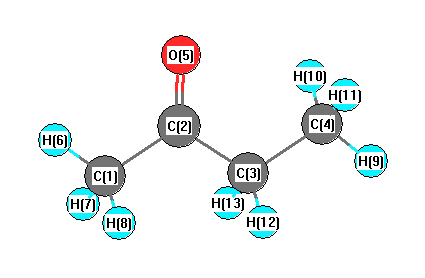
picture of 2-Butanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 116.543 C1 C2 O5 121.647
C2 C1 H6 109.658 C2 C1 H7 110.440
C2 C1 H8 110.440 C2 C3 C4 113.310
C2 C3 H12 107.869 C2 C3 H13 107.869
C3 C2 O5 121.810 C3 C4 H9 110.684
C3 C4 H10 110.808 C3 C4 H11 110.808
C4 C3 H12 111.132 C4 C3 H13 111.132
H6 C1 H7 109.596 H6 C1 H8 109.596
H7 C1 H8 107.070 H9 C4 H10 108.548
H9 C4 H11 108.548 H10 C4 H11 107.338
H12 C3 H13 105.124
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.545      
2 C 0.446      
3 C -0.368      
4 C -0.449      
5 O -0.428      
6 H 0.187      
7 H 0.173      
8 H 0.173      
9 H 0.147      
10 H 0.167      
11 H 0.167      
12 H 0.165      
13 H 0.165      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.005 0.191 0.000
y 0.191 7.097 0.000
z 0.000 0.000 5.278


<r2> (average value of r2) Å2
<r2> 137.154
(<r2>)1/2 11.711