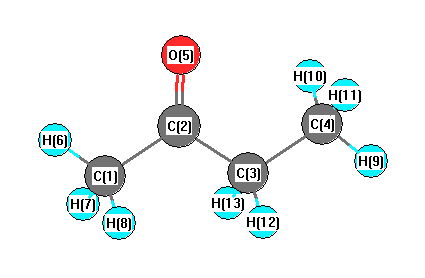
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3198 |
3041 |
8.38 |
|
|
|
| 2 |
A' |
3168 |
3013 |
20.18 |
|
|
|
| 3 |
A' |
3088 |
2937 |
20.41 |
|
|
|
| 4 |
A' |
3072 |
2922 |
6.39 |
|
|
|
| 5 |
A' |
3051 |
2901 |
15.97 |
|
|
|
| 6 |
A' |
1852 |
1761 |
135.82 |
|
|
|
| 7 |
A' |
1525 |
1451 |
9.92 |
|
|
|
| 8 |
A' |
1493 |
1420 |
19.48 |
|
|
|
| 9 |
A' |
1478 |
1405 |
3.06 |
|
|
|
| 10 |
A' |
1438 |
1367 |
6.70 |
|
|
|
| 11 |
A' |
1415 |
1346 |
61.48 |
|
|
|
| 12 |
A' |
1391 |
1323 |
9.98 |
|
|
|
| 13 |
A' |
1209 |
1150 |
68.64 |
|
|
|
| 14 |
A' |
1126 |
1071 |
1.09 |
|
|
|
| 15 |
A' |
1022 |
972 |
4.91 |
|
|
|
| 16 |
A' |
956 |
909 |
6.27 |
|
|
|
| 17 |
A' |
781 |
743 |
2.72 |
|
|
|
| 18 |
A' |
595 |
566 |
8.65 |
|
|
|
| 19 |
A' |
404 |
384 |
4.20 |
|
|
|
| 20 |
A' |
248 |
236 |
5.30 |
|
|
|
| 21 |
A" |
3173 |
3018 |
19.34 |
|
|
|
| 22 |
A" |
3141 |
2987 |
8.95 |
|
|
|
| 23 |
A" |
3085 |
2934 |
9.17 |
|
|
|
| 24 |
A" |
1521 |
1446 |
7.69 |
|
|
|
| 25 |
A" |
1503 |
1429 |
11.84 |
|
|
|
| 26 |
A" |
1295 |
1232 |
0.01 |
|
|
|
| 27 |
A" |
1143 |
1087 |
0.49 |
|
|
|
| 28 |
A" |
958 |
911 |
5.05 |
|
|
|
| 29 |
A" |
760 |
723 |
3.71 |
|
|
|
| 30 |
A" |
475 |
452 |
0.00 |
|
|
|
| 31 |
A" |
203 |
193 |
0.18 |
|
|
|
| 32 |
A" |
107 |
102 |
0.00 |
|
|
|
| 33 |
A" |
27i |
25i |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24923.2 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 23701.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.604 |
|
|
|
| 2 |
C |
0.456 |
|
|
|
| 3 |
C |
-0.414 |
|
|
|
| 4 |
C |
-0.508 |
|
|
|
| 5 |
O |
-0.433 |
|
|
|
| 6 |
H |
0.207 |
|
|
|
| 7 |
H |
0.192 |
|
|
|
| 8 |
H |
0.192 |
|
|
|
| 9 |
H |
0.167 |
|
|
|
| 10 |
H |
0.187 |
|
|
|
| 11 |
H |
0.187 |
|
|
|
| 12 |
H |
0.186 |
|
|
|
| 13 |
H |
0.186 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.660 |
-0.589 |
0.000 |
2.724 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.381 |
2.289 |
0.000 |
| y |
2.289 |
-30.751 |
0.000 |
| z |
0.000 |
0.000 |
-30.169 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.921 |
2.289 |
0.000 |
| y |
2.289 |
1.024 |
0.000 |
| z |
0.000 |
0.000 |
1.897 |
|
| Polar |
|---|
| 3z2-r2 | 3.794 |
|---|
| x2-y2 | -2.630 |
|---|
| xy | 2.289 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.066 |
0.221 |
0.000 |
| y |
0.221 |
7.200 |
0.000 |
| z |
0.000 |
0.000 |
5.319 |
<r2> (average value of r
2) Å
2
| <r2> |
136.206 |
| (<r2>)1/2 |
11.671 |