Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -1072.555963 |
| Energy at 298.15K | -1072.558453 |
| HF Energy | -1072.555963 |
| Nuclear repulsion energy | 263.519235 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3202 |
3045 |
0.75 |
|
|
|
| 2 |
A' |
3014 |
2866 |
46.16 |
|
|
|
| 3 |
A' |
1880 |
1787 |
161.04 |
|
|
|
| 4 |
A' |
1417 |
1348 |
3.84 |
|
|
|
| 5 |
A' |
1235 |
1174 |
8.09 |
|
|
|
| 6 |
A' |
1081 |
1028 |
26.92 |
|
|
|
| 7 |
A' |
799 |
760 |
25.68 |
|
|
|
| 8 |
A' |
445 |
423 |
4.52 |
|
|
|
| 9 |
A' |
326 |
310 |
25.33 |
|
|
|
| 10 |
A' |
259 |
246 |
2.76 |
|
|
|
| 11 |
A" |
1276 |
1213 |
25.72 |
|
|
|
| 12 |
A" |
1025 |
975 |
19.74 |
|
|
|
| 13 |
A" |
747 |
710 |
131.58 |
|
|
|
| 14 |
A" |
275 |
262 |
3.22 |
|
|
|
| 15 |
A" |
82 |
78 |
7.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8531.0 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 8113.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.385 |
-0.022 |
0.000 |
| C2 |
-0.171 |
1.397 |
0.000 |
| H3 |
1.474 |
-0.003 |
0.000 |
| Cl4 |
-0.171 |
-0.841 |
1.475 |
| Cl5 |
-0.171 |
-0.841 |
-1.475 |
| O6 |
0.544 |
2.360 |
0.000 |
| H7 |
-1.277 |
1.451 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5234 | 1.0892 | 1.7767 | 1.7767 | 2.3873 | 2.2208 |
C2 | 1.5234 | | 2.1599 | 2.6801 | 2.6801 | 1.2002 | 1.1073 | H3 | 1.0892 | 2.1599 | | 2.3631 | 2.3631 | 2.5396 | 3.1118 | Cl4 | 1.7767 | 2.6801 | 2.3631 | | 2.9510 | 3.5965 | 2.9417 | Cl5 | 1.7767 | 2.6801 | 2.3631 | 2.9510 | | 3.5965 | 2.9417 | O6 | 2.3873 | 1.2002 | 2.5396 | 3.5965 | 3.5965 | | 2.0357 | H7 | 2.2208 | 1.1073 | 3.1118 | 2.9417 | 2.9417 | 2.0357 | |
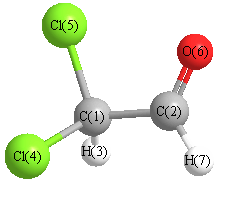 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.003 |
|
C1 |
C2 |
H7 |
114.234 |
| C2 |
C1 |
H3 |
110.427 |
|
C2 |
C1 |
Cl4 |
108.359 |
| C2 |
C1 |
Cl5 |
108.359 |
|
H3 |
C1 |
Cl4 |
108.705 |
| H3 |
C1 |
Cl5 |
108.705 |
|
Cl4 |
C1 |
Cl5 |
112.289 |
| O6 |
C2 |
H7 |
123.762 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.457 |
|
|
|
| 2 |
C |
0.279 |
|
|
|
| 3 |
H |
0.288 |
|
|
|
| 4 |
Cl |
0.030 |
|
|
|
| 5 |
Cl |
0.030 |
|
|
|
| 6 |
O |
-0.347 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.259 |
-0.575 |
0.000 |
0.630 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.453 |
-2.817 |
0.000 |
| y |
-2.817 |
-47.941 |
0.000 |
| z |
0.000 |
0.000 |
-42.652 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.843 |
-2.817 |
0.000 |
| y |
-2.817 |
-6.389 |
0.000 |
| z |
0.000 |
0.000 |
1.545 |
|
| Polar |
|---|
| 3z2-r2 | 3.090 |
|---|
| x2-y2 | 7.488 |
|---|
| xy | -2.817 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.961 |
0.703 |
0.000 |
| y |
0.703 |
6.811 |
0.000 |
| z |
0.000 |
0.000 |
7.057 |
<r2> (average value of r
2) Å
2
| <r2> |
191.947 |
| (<r2>)1/2 |
13.854 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G*
| | hartrees |
|---|
| Energy at 0K | -1072.553705 |
| Energy at 298.15K | |
| HF Energy | -1072.553705 |
| Nuclear repulsion energy | 266.250661 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3140 |
2986 |
4.26 |
84.01 |
0.24 |
0.38 |
| 2 |
A |
2996 |
2849 |
54.27 |
130.57 |
0.32 |
0.48 |
| 3 |
A |
1893 |
1800 |
133.75 |
14.55 |
0.44 |
0.61 |
| 4 |
A |
1413 |
1344 |
14.08 |
4.75 |
0.67 |
0.80 |
| 5 |
A |
1287 |
1224 |
15.27 |
5.90 |
0.73 |
0.85 |
| 6 |
A |
1250 |
1189 |
11.78 |
10.89 |
0.74 |
0.85 |
| 7 |
A |
1045 |
994 |
13.13 |
2.22 |
0.53 |
0.69 |
| 8 |
A |
950 |
903 |
10.11 |
6.84 |
0.60 |
0.75 |
| 9 |
A |
827 |
787 |
88.29 |
6.36 |
0.67 |
0.81 |
| 10 |
A |
661 |
628 |
40.13 |
8.70 |
0.11 |
0.19 |
| 11 |
A |
619 |
588 |
32.55 |
6.77 |
0.43 |
0.60 |
| 12 |
A |
349 |
332 |
1.49 |
3.38 |
0.24 |
0.39 |
| 13 |
A |
279 |
265 |
3.73 |
5.60 |
0.65 |
0.79 |
| 14 |
A |
221 |
210 |
2.69 |
1.47 |
0.71 |
0.83 |
| 15 |
A |
88 |
84 |
10.15 |
2.06 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8508.8 cm
-1
Scaled (by 0.951) Zero Point Vibrational Energy (zpe) 8091.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.101 |
-0.025 |
0.511 |
| C2 |
-0.709 |
-1.281 |
0.183 |
| H3 |
0.223 |
0.053 |
1.595 |
| Cl4 |
1.739 |
-0.255 |
-0.165 |
| Cl5 |
-0.679 |
1.443 |
-0.058 |
| O6 |
-1.802 |
-1.275 |
-0.305 |
| H7 |
-0.175 |
-2.213 |
0.459 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5306 | 1.0934 | 1.7874 | 1.7569 | 2.4185 | 2.2059 |
C2 | 1.5306 | | 2.1550 | 2.6774 | 2.7354 | 1.1970 | 1.1086 | H3 | 1.0934 | 2.1550 | | 2.3430 | 2.3403 | 3.0782 | 2.5657 | Cl4 | 1.7874 | 2.6774 | 2.3430 | | 2.9573 | 3.6878 | 2.8081 | Cl5 | 1.7569 | 2.7354 | 2.3403 | 2.9573 | | 2.9510 | 3.7265 | O6 | 2.4185 | 1.1970 | 3.0782 | 3.6878 | 2.9510 | | 2.0277 | H7 | 2.2059 | 1.1086 | 2.5657 | 2.8081 | 3.7265 | 2.0277 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.467 |
|
C1 |
C2 |
H7 |
112.417 |
| C2 |
C1 |
H3 |
109.290 |
|
C2 |
C1 |
Cl4 |
107.344 |
| C2 |
C1 |
Cl5 |
112.437 |
|
H3 |
C1 |
Cl4 |
106.334 |
| H3 |
C1 |
Cl5 |
108.117 |
|
Cl4 |
C1 |
Cl5 |
113.103 |
| O6 |
C2 |
H7 |
123.112 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.466 |
|
|
|
| 2 |
C |
0.265 |
|
|
|
| 3 |
H |
0.281 |
|
|
|
| 4 |
Cl |
0.017 |
|
|
|
| 5 |
Cl |
0.052 |
|
|
|
| 6 |
O |
-0.325 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.150 |
-0.893 |
1.964 |
2.445 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.299 |
-1.930 |
-1.087 |
| y |
-1.930 |
-41.308 |
-1.434 |
| z |
-1.087 |
-1.434 |
-39.923 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.684 |
-1.930 |
-1.087 |
| y |
-1.930 |
2.303 |
-1.434 |
| z |
-1.087 |
-1.434 |
4.381 |
|
| Polar |
|---|
| 3z2-r2 | 8.761 |
|---|
| x2-y2 | -5.992 |
|---|
| xy | -1.930 |
|---|
| xz | -1.087 |
|---|
| yz | -1.434 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.653 |
-0.719 |
0.178 |
| y |
-0.719 |
6.625 |
-0.317 |
| z |
0.178 |
-0.317 |
4.261 |
<r2> (average value of r
2) Å
2
| <r2> |
185.220 |
| (<r2>)1/2 |
13.610 |
