Jump to
S1C2
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
Geometric Data calculated at PBEPBEultrafine/6-31G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBEultrafine/6-31G*
| | hartrees |
|---|
| Energy at 0K | -1072.428953 |
| Energy at 298.15K | |
| HF Energy | -1072.428953 |
| Nuclear repulsion energy | 263.512179 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3040 |
2990 |
3.55 |
90.66 |
0.22 |
0.36 |
| 2 |
A |
2870 |
2823 |
65.27 |
146.49 |
0.33 |
0.49 |
| 3 |
A |
1801 |
1772 |
121.52 |
17.62 |
0.40 |
0.57 |
| 4 |
A |
1362 |
1339 |
11.36 |
5.87 |
0.71 |
0.83 |
| 5 |
A |
1229 |
1208 |
16.54 |
7.35 |
0.75 |
0.86 |
| 6 |
A |
1192 |
1173 |
9.61 |
11.62 |
0.73 |
0.85 |
| 7 |
A |
989 |
973 |
13.28 |
2.54 |
0.59 |
0.74 |
| 8 |
A |
900 |
885 |
11.63 |
6.40 |
0.56 |
0.71 |
| 9 |
A |
765 |
752 |
86.23 |
5.33 |
0.61 |
0.76 |
| 10 |
A |
625 |
615 |
39.27 |
9.37 |
0.10 |
0.17 |
| 11 |
A |
590 |
580 |
43.19 |
7.94 |
0.43 |
0.60 |
| 12 |
A |
330 |
324 |
2.30 |
3.90 |
0.21 |
0.35 |
| 13 |
A |
266 |
262 |
3.39 |
6.72 |
0.64 |
0.78 |
| 14 |
A |
211 |
208 |
1.92 |
1.83 |
0.71 |
0.83 |
| 15 |
A |
84 |
82 |
8.20 |
2.14 |
0.69 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 8126.5 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 7992.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBEultrafine/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.099 |
-0.015 |
0.522 |
| C2 |
-0.692 |
-1.300 |
0.203 |
| H3 |
0.234 |
0.078 |
1.613 |
| Cl4 |
1.757 |
-0.235 |
-0.171 |
| Cl5 |
-0.715 |
1.450 |
-0.059 |
| O6 |
-1.782 |
-1.327 |
-0.323 |
| H7 |
-0.142 |
-2.222 |
0.522 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5428 | 1.1026 | 1.8105 | 1.7740 | 2.4444 | 2.2196 |
C2 | 1.5428 | | 2.1784 | 2.6971 | 2.7629 | 1.2101 | 1.1200 | H3 | 1.1026 | 2.1784 | | 2.3660 | 2.3616 | 3.1283 | 2.5726 | Cl4 | 1.8105 | 2.6971 | 2.3660 | | 2.9938 | 3.7069 | 2.8339 | Cl5 | 1.7740 | 2.7629 | 2.3616 | 2.9938 | | 2.9871 | 3.7613 | O6 | 2.4444 | 1.2101 | 3.1283 | 3.7069 | 2.9871 | | 2.0507 | H7 | 2.2196 | 1.1200 | 2.5726 | 2.8339 | 3.7613 | 2.0507 | |
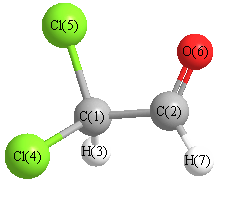 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.786 |
|
C1 |
C2 |
H7 |
111.954 |
| C2 |
C1 |
H3 |
109.747 |
|
C2 |
C1 |
Cl4 |
106.817 |
| C2 |
C1 |
Cl5 |
112.631 |
|
H3 |
C1 |
Cl4 |
106.062 |
| H3 |
C1 |
Cl5 |
108.087 |
|
Cl4 |
C1 |
Cl5 |
113.275 |
| O6 |
C2 |
H7 |
123.259 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.441 |
|
|
|
| 2 |
C |
0.247 |
|
|
|
| 3 |
H |
0.265 |
|
|
|
| 4 |
Cl |
0.014 |
|
|
|
| 5 |
Cl |
0.050 |
|
|
|
| 6 |
O |
-0.293 |
|
|
|
| 7 |
H |
0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.014 |
-0.794 |
1.937 |
2.326 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.790 |
-1.854 |
-1.031 |
| y |
-1.854 |
-41.499 |
-1.423 |
| z |
-1.031 |
-1.423 |
-39.967 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.057 |
-1.854 |
-1.031 |
| y |
-1.854 |
1.880 |
-1.423 |
| z |
-1.031 |
-1.423 |
4.177 |
|
| Polar |
|---|
| 3z2-r2 | 8.355 |
|---|
| x2-y2 | -5.291 |
|---|
| xy | -1.854 |
|---|
| xz | -1.031 |
|---|
| yz | -1.423 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.079 |
-0.689 |
0.205 |
| y |
-0.689 |
7.018 |
-0.349 |
| z |
0.205 |
-0.349 |
4.400 |
<r2> (average value of r
2) Å
2
| <r2> |
188.303 |
| (<r2>)1/2 |
13.722 |
