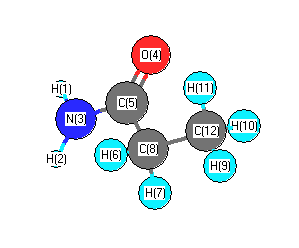
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3743 |
3581 |
32.72 |
|
|
|
| 2 |
A |
3613 |
3456 |
31.38 |
|
|
|
| 3 |
A |
3169 |
3031 |
15.75 |
|
|
|
| 4 |
A |
3147 |
3011 |
26.93 |
|
|
|
| 5 |
A |
3086 |
2953 |
18.78 |
|
|
|
| 6 |
A |
3075 |
2941 |
23.34 |
|
|
|
| 7 |
A |
3045 |
2913 |
20.13 |
|
|
|
| 8 |
A |
1825 |
1746 |
274.12 |
|
|
|
| 9 |
A |
1646 |
1574 |
113.19 |
|
|
|
| 10 |
A |
1528 |
1462 |
10.45 |
|
|
|
| 11 |
A |
1518 |
1453 |
7.55 |
|
|
|
| 12 |
A |
1492 |
1428 |
5.55 |
|
|
|
| 13 |
A |
1437 |
1374 |
5.67 |
|
|
|
| 14 |
A |
1424 |
1362 |
76.85 |
|
|
|
| 15 |
A |
1306 |
1249 |
63.82 |
|
|
|
| 16 |
A |
1297 |
1240 |
28.73 |
|
|
|
| 17 |
A |
1135 |
1086 |
1.69 |
|
|
|
| 18 |
A |
1110 |
1062 |
0.36 |
|
|
|
| 19 |
A |
1077 |
1030 |
2.46 |
|
|
|
| 20 |
A |
1017 |
973 |
4.04 |
|
|
|
| 21 |
A |
825 |
790 |
11.57 |
|
|
|
| 22 |
A |
799 |
765 |
3.88 |
|
|
|
| 23 |
A |
659 |
630 |
6.42 |
|
|
|
| 24 |
A |
604 |
578 |
12.73 |
|
|
|
| 25 |
A |
505 |
483 |
6.04 |
|
|
|
| 26 |
A |
437 |
418 |
3.19 |
|
|
|
| 27 |
A |
258 |
247 |
10.45 |
|
|
|
| 28 |
A |
208 |
199 |
3.03 |
|
|
|
| 29 |
A |
182 |
174 |
242.40 |
|
|
|
| 30 |
A |
36 |
34 |
3.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22600.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 21621.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.359 |
|
|
0.428 |
| 2 |
H |
0.351 |
|
|
0.394 |
| 3 |
N |
-0.790 |
|
|
-0.973 |
| 4 |
O |
-0.506 |
|
|
-0.524 |
| 5 |
C |
0.609 |
|
|
0.672 |
| 6 |
H |
0.193 |
|
|
0.035 |
| 7 |
H |
0.170 |
|
|
0.036 |
| 8 |
C |
-0.424 |
|
|
-0.034 |
| 9 |
H |
0.162 |
|
|
0.071 |
| 10 |
H |
0.179 |
|
|
0.078 |
| 11 |
H |
0.198 |
|
|
0.080 |
| 12 |
C |
-0.503 |
|
|
-0.263 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.969 |
-3.508 |
0.194 |
3.645 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.956 |
-3.506 |
0.212 |
3.640 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.812 |
2.680 |
-0.006 |
| y |
2.680 |
-32.198 |
-0.399 |
| z |
-0.006 |
-0.399 |
-30.914 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.744 |
2.680 |
-0.006 |
| y |
2.680 |
-4.335 |
-0.399 |
| z |
-0.006 |
-0.399 |
-2.409 |
|
| Polar |
|---|
| 3z2-r2 | -4.818 |
|---|
| x2-y2 | 7.386 |
|---|
| xy | 2.680 |
|---|
| xz | -0.006 |
|---|
| yz | -0.399 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
127.703 |
| (<r2>)1/2 |
11.301 |