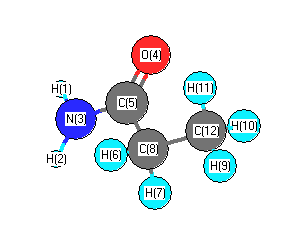
Vibrational Frequencies calculated at PBE1PBE/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3766 |
3578 |
36.45 |
|
|
|
| 2 |
A |
3634 |
3454 |
35.56 |
|
|
|
| 3 |
A |
3185 |
3027 |
13.37 |
|
|
|
| 4 |
A |
3166 |
3008 |
23.90 |
|
|
|
| 5 |
A |
3104 |
2950 |
16.58 |
|
|
|
| 6 |
A |
3089 |
2936 |
21.56 |
|
|
|
| 7 |
A |
3061 |
2909 |
18.50 |
|
|
|
| 8 |
A |
1846 |
1754 |
279.56 |
|
|
|
| 9 |
A |
1652 |
1570 |
121.94 |
|
|
|
| 10 |
A |
1530 |
1454 |
11.71 |
|
|
|
| 11 |
A |
1520 |
1445 |
7.99 |
|
|
|
| 12 |
A |
1494 |
1420 |
5.93 |
|
|
|
| 13 |
A |
1444 |
1372 |
25.87 |
|
|
|
| 14 |
A |
1432 |
1360 |
63.25 |
|
|
|
| 15 |
A |
1312 |
1247 |
68.02 |
|
|
|
| 16 |
A |
1301 |
1236 |
13.89 |
|
|
|
| 17 |
A |
1142 |
1085 |
1.57 |
|
|
|
| 18 |
A |
1113 |
1058 |
0.35 |
|
|
|
| 19 |
A |
1087 |
1033 |
1.95 |
|
|
|
| 20 |
A |
1022 |
971 |
4.43 |
|
|
|
| 21 |
A |
829 |
788 |
11.84 |
|
|
|
| 22 |
A |
808 |
768 |
3.99 |
|
|
|
| 23 |
A |
662 |
629 |
6.59 |
|
|
|
| 24 |
A |
608 |
578 |
13.30 |
|
|
|
| 25 |
A |
508 |
483 |
5.99 |
|
|
|
| 26 |
A |
440 |
418 |
3.24 |
|
|
|
| 27 |
A |
261 |
248 |
10.61 |
|
|
|
| 28 |
A |
208 |
198 |
3.17 |
|
|
|
| 29 |
A |
185 |
175 |
244.53 |
|
|
|
| 30 |
A |
38 |
36 |
3.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22721.9 cm
-1
Scaled (by 0.9503) Zero Point Vibrational Energy (zpe) 21592.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.516 |
|
|
|
| 2 |
C |
-0.438 |
|
|
|
| 3 |
C |
0.611 |
|
|
|
| 4 |
O |
-0.505 |
|
|
|
| 5 |
N |
-0.798 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
| 7 |
H |
0.203 |
|
|
|
| 8 |
H |
0.184 |
|
|
|
| 9 |
H |
0.176 |
|
|
|
| 10 |
H |
0.198 |
|
|
|
| 11 |
H |
0.355 |
|
|
|
| 12 |
H |
0.362 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.983 |
-3.507 |
0.205 |
3.647 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.890 |
-2.637 |
0.038 |
| y |
-2.637 |
-32.249 |
-0.407 |
| z |
0.038 |
-0.407 |
-30.982 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.726 |
-2.637 |
0.038 |
| y |
-2.637 |
-4.313 |
-0.407 |
| z |
0.038 |
-0.407 |
-2.412 |
|
| Polar |
|---|
| 3z2-r2 | -4.825 |
|---|
| x2-y2 | 7.359 |
|---|
| xy | -2.637 |
|---|
| xz | 0.038 |
|---|
| yz | -0.407 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.955 |
-0.323 |
-0.154 |
| y |
-0.323 |
6.447 |
0.029 |
| z |
-0.154 |
0.029 |
4.342 |
<r2> (average value of r
2) Å
2
| <r2> |
127.170 |
| (<r2>)1/2 |
11.277 |