return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (Propanamide)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-248.252723
Energy at 298.15K-248.260901
HF Energy-248.011978
Nuclear repulsion energy178.413243
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3739 3549 31.31      
2 A 3610 3426 33.70      
3 A 3191 3029 16.98      
4 A 3170 3009 26.81      
5 A 3109 2951 18.14      
6 A 3098 2941 22.11      
7 A 3068 2912 18.80      
8 A 1805 1713 248.51      
9 A 1660 1576 96.83      
10 A 1553 1474 8.56      
11 A 1544 1466 6.00      
12 A 1521 1444 4.25      
13 A 1461 1387 3.28      
14 A 1439 1366 64.66      
15 A 1320 1253 27.95      
16 A 1313 1246 79.47      
17 A 1148 1090 2.50      
18 A 1128 1070 0.40      
19 A 1084 1028 3.45      
20 A 1027 975 2.71      
21 A 834 792 10.33      
22 A 806 765 3.61      
23 A 650 617 9.32      
24 A 608 577 19.89      
25 A 503 477 8.97      
26 A 443 420 3.17      
27 A 268 254 247.32      
28 A 263 250 11.87      
29 A 214 203 0.71      
30 A 33 32 2.98      

Unscaled Zero Point Vibrational Energy (zpe) 22803.7 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 21645.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.32305 0.12655 0.09513

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.965 0.038 -0.123
C2 -0.684 -0.740 0.156
C3 0.559 0.135 0.031
O4 0.522 1.359 0.066
N5 1.734 -0.552 -0.135
H6 -2.842 -0.594 0.039
H7 -2.030 0.908 0.532
H8 -1.985 0.399 -1.153
H9 -0.592 -1.601 -0.516
H10 -0.701 -1.147 1.175
H11 1.785 -1.547 0.017
H12 2.591 -0.023 -0.060

Atom - Atom Distances (Å)
  C1 C2 C3 O4 N5 H6 H7 H8 H9 H10 H11 H12
C11.52522.53072.82263.74631.09301.09061.09232.17422.16514.07364.5569
C21.52521.52512.42262.44252.16702.16172.16941.09591.09772.60043.3590
C32.53071.52511.22481.37183.47862.74812.81852.15402.13062.08202.0400
O42.82262.42261.22482.27223.89022.63342.94853.21603.00083.16942.4910
N53.74632.44251.37182.27224.58014.09273.97182.58022.82831.00821.0098
H61.09302.16703.47863.89024.58011.77721.77252.52712.48704.72435.4640
H71.09062.16172.74812.63344.09271.77721.76083.07632.53094.56614.7509
H81.09232.16942.81852.94853.97181.77251.76082.51933.07614.40114.7236
H92.17421.09592.15403.21602.58022.52713.07632.51931.75502.43643.5820
H102.16511.09772.13063.00082.82832.48702.53093.07611.75502.77093.6908
H114.07362.60042.08203.16941.00824.72434.56614.40112.43642.77091.7266
H124.55693.35902.04002.49101.00985.46404.75094.72363.58203.69081.7266

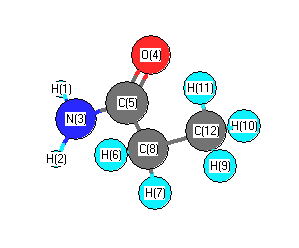
picture of Propanamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 N3 H2 33.938 H1 N3 C5 145.889
H2 N3 C5 114.841 N3 C5 O4 27.199
N3 C5 C8 26.961 O4 C5 C8 47.283
C5 C8 H6 98.418 C5 C8 H7 81.261
C5 C8 C12 8.927 H6 C8 H7 60.393
H6 C8 C12 105.239 H7 C8 C12 80.165
C8 C12 H9 31.685 C8 C12 H10 40.605
C8 C12 H11 68.704 H9 C12 H10 27.876
H9 C12 H11 37.020 H10 C12 H11 45.551
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.453      
2 C -0.391      
3 C 0.659      
4 O -0.549      
5 N -0.818      
6 H 0.148      
7 H 0.184      
8 H 0.166      
9 H 0.163      
10 H 0.176      
11 H 0.354      
12 H 0.362      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 6.273 0.250 0.129
y 0.250 4.484 -0.606
z 0.129 -0.606 6.951


<r2> (average value of r2) Å2
<r2> 128.168
(<r2>)1/2 11.321