return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (Propanamide)

using model chemistry: MP3/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP3/6-31G*
 hartrees
Energy at 0K-247.762352
Energy at 298.15K-247.770662
HF Energy-247.009976
Nuclear repulsion energy178.720287
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3765 3534 34.79      
2 A 3645 3421 39.76      
3 A 3211 3014 17.60      
4 A 3198 3002 20.15      
5 A 3133 2940 11.34      
6 A 3120 2928 18.15      
7 A 3091 2902 18.06      
8 A 1885 1769 270.34      
9 A 1683 1580 119.18      
10 A 1560 1464 11.47      
11 A 1553 1458 5.70      
12 A 1538 1444 4.80      
13 A 1485 1393 18.53      
14 A 1466 1376 69.15      
15 A 1333 1251 94.41      
16 A 1328 1246 2.26      
17 A 1172 1100 2.71      
18 A 1143 1073 0.21      
19 A 1110 1042 3.13      
20 A 1041 977 3.21      
21 A 843 792 11.56      
22 A 830 779 3.65      
23 A 634 595 35.08      
24 A 618 580 6.85      
25 A 500 469 32.44      
26 A 459 431 10.65      
27 A 352 331 263.20      
28 A 266 249 9.17      
29 A 215 202 0.21      
30 A 29 27 1.19      

Unscaled Zero Point Vibrational Energy (zpe) 23102.4 cm-1
Scaled (by 0.9386) Zero Point Vibrational Energy (zpe) 21684.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/6-31G*
ABC
0.33026 0.12660 0.09483

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.582 0.024 0.104
H2 -1.806 -1.506 0.195
N3 -1.756 -0.533 -0.064
O4 -0.497 1.352 0.013
C5 -0.557 0.135 0.004
H6 0.627 -1.401 0.900
H7 0.613 -1.432 -0.854
C8 0.673 -0.759 0.011
H9 2.833 -0.634 0.002
H10 2.026 0.664 -0.903
H11 2.031 0.701 0.859
C12 1.971 0.040 -0.009

Atom - Atom Distances (Å)
  H1 H2 N3 O4 C5 H6 H7 C8 H9 H10 H11 C12
H11.71811.01012.47382.03053.60063.63953.34965.45594.75974.72284.5549
H21.71811.00863.14852.07102.53492.63752.59574.72394.53814.47524.0861
N31.01011.00862.26751.37382.71352.65432.44104.59064.05424.08783.7713
O42.47383.14852.26751.21813.10353.11952.41353.87712.77042.74362.7954
C52.03052.07101.37381.21812.13732.13551.52123.47642.78792.78352.5304
H63.60062.53492.71353.10352.13731.75441.09842.50243.07812.52822.1704
H73.63952.63752.65433.11952.13551.75441.09712.50942.52793.08042.1735
C83.34962.59572.44102.41351.52121.09841.09712.16332.16532.16601.5241
H95.45594.72394.59063.87713.47642.50242.50942.16331.77671.77721.0939
H104.75974.53814.05422.77042.78793.07812.52792.16531.77671.76181.0921
H114.72284.47524.08782.74362.78352.52823.08042.16601.77721.76181.0920
C124.55494.08613.77132.79542.53042.17042.17351.52411.09391.09211.0920

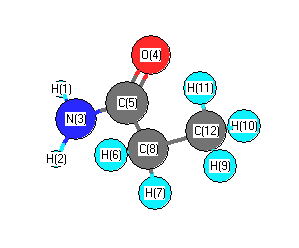
picture of Propanamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 N3 H2 116.657 H1 N3 C5 115.962
H2 N3 C5 119.973 N3 C5 O4 121.940
N3 C5 C8 114.856 O4 C5 C8 123.164
C5 C8 H6 108.275 C5 C8 H7 108.203
C5 C8 C12 112.385 H6 C8 H7 106.087
H6 C8 C12 110.661 H7 C8 C12 110.981
C8 C12 H9 110.365 C8 C12 H10 110.631
C8 C12 H11 110.695 H9 C12 H10 108.739
H9 C12 H11 108.787 H10 C12 H11 107.545
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability