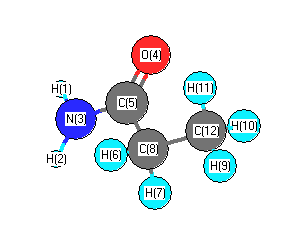
Vibrational Frequencies calculated at B3LYP/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3712 |
3565 |
26.61 |
|
|
|
| 2 |
A |
3586 |
3444 |
27.26 |
|
|
|
| 3 |
A |
3153 |
3027 |
17.84 |
|
|
|
| 4 |
A |
3129 |
3005 |
30.77 |
|
|
|
| 5 |
A |
3070 |
2948 |
22.60 |
|
|
|
| 6 |
A |
3065 |
2943 |
24.04 |
|
|
|
| 7 |
A |
3032 |
2912 |
20.33 |
|
|
|
| 8 |
A |
1808 |
1736 |
264.82 |
|
|
|
| 9 |
A |
1645 |
1580 |
101.09 |
|
|
|
| 10 |
A |
1534 |
1473 |
8.46 |
|
|
|
| 11 |
A |
1524 |
1463 |
6.20 |
|
|
|
| 12 |
A |
1499 |
1440 |
4.17 |
|
|
|
| 13 |
A |
1441 |
1383 |
3.74 |
|
|
|
| 14 |
A |
1419 |
1362 |
61.86 |
|
|
|
| 15 |
A |
1307 |
1255 |
47.61 |
|
|
|
| 16 |
A |
1297 |
1245 |
64.54 |
|
|
|
| 17 |
A |
1135 |
1090 |
2.45 |
|
|
|
| 18 |
A |
1113 |
1068 |
0.50 |
|
|
|
| 19 |
A |
1070 |
1027 |
3.76 |
|
|
|
| 20 |
A |
1013 |
973 |
2.49 |
|
|
|
| 21 |
A |
825 |
793 |
10.50 |
|
|
|
| 22 |
A |
794 |
762 |
3.84 |
|
|
|
| 23 |
A |
656 |
630 |
7.31 |
|
|
|
| 24 |
A |
604 |
580 |
15.58 |
|
|
|
| 25 |
A |
503 |
483 |
5.91 |
|
|
|
| 26 |
A |
437 |
420 |
3.00 |
|
|
|
| 27 |
A |
261 |
250 |
10.34 |
|
|
|
| 28 |
A |
217 |
208 |
218.18 |
|
|
|
| 29 |
A |
207 |
199 |
19.00 |
|
|
|
| 30 |
A |
34 |
33 |
3.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22543.6 cm
-1
Scaled (by 0.9603) Zero Point Vibrational Energy (zpe) 21648.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.338 |
0.407 |
0.425 |
0.410 |
| 2 |
H |
0.331 |
0.375 |
0.409 |
0.377 |
| 3 |
N |
-0.746 |
-0.928 |
-1.222 |
-0.928 |
| 4 |
O |
-0.501 |
-0.534 |
-1.215 |
-0.522 |
| 5 |
C |
0.594 |
0.681 |
1.518 |
0.656 |
| 6 |
H |
0.165 |
0.019 |
0.023 |
0.024 |
| 7 |
H |
0.144 |
0.023 |
0.006 |
0.027 |
| 8 |
C |
-0.359 |
-0.026 |
-0.001 |
-0.017 |
| 9 |
H |
0.138 |
0.041 |
0.006 |
0.053 |
| 10 |
H |
0.155 |
0.045 |
0.016 |
0.061 |
| 11 |
H |
0.175 |
0.050 |
0.038 |
0.063 |
| 12 |
C |
-0.434 |
-0.152 |
-0.005 |
-0.203 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.933 |
-3.460 |
0.279 |
3.594 |
| CHELPG |
-0.933 |
-3.458 |
0.288 |
3.593 |
| AIM |
-0.789 |
1.290 |
-0.076 |
1.514 |
| ESP |
-0.920 |
-3.457 |
0.293 |
3.589 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.141 |
2.662 |
-0.197 |
| y |
2.662 |
-32.458 |
-0.504 |
| z |
-0.197 |
-0.504 |
-31.052 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.614 |
2.662 |
-0.197 |
| y |
2.662 |
-4.362 |
-0.504 |
| z |
-0.197 |
-0.504 |
-2.253 |
|
| Polar |
|---|
| 3z2-r2 | -4.506 |
|---|
| x2-y2 | 7.317 |
|---|
| xy | 2.662 |
|---|
| xz | -0.197 |
|---|
| yz | -0.504 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.061 |
0.319 |
0.168 |
| y |
0.319 |
6.496 |
0.027 |
| z |
0.168 |
0.027 |
4.355 |
<r2> (average value of r
2) Å
2
| <r2> |
128.425 |
| (<r2>)1/2 |
11.332 |