return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HOCH2COOH (Hydroxyacetic acid)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-304.001041
Energy at 298.15K-304.006534
HF Energy-303.740402
Nuclear repulsion energy179.540225
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3701 3513 20.01      
2 A 3700 3512 114.85      
3 A 3093 2936 24.61      
4 A 3062 2907 27.09      
5 A 1834 1741 243.65      
6 A 1550 1471 5.43      
7 A 1514 1437 19.46      
8 A 1389 1318 142.47      
9 A 1328 1260 22.83      
10 A 1266 1202 0.78      
11 A 1203 1142 148.06      
12 A 1136 1079 212.20      
13 A 1066 1012 0.83      
14 A 874 829 36.30      
15 A 677 643 147.46      
16 A 652 619 19.16      
17 A 510 484 3.98      
18 A 476 452 27.57      
19 A 297 282 75.92      
20 A 289 274 11.34      
21 A 32 30 20.63      

Unscaled Zero Point Vibrational Energy (zpe) 14823.9 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 14070.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.35477 0.13531 0.09977

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.742 -0.752 0.004
C2 0.513 0.095 0.000
O3 1.639 -0.644 -0.003
O4 0.499 1.311 0.002
O5 -1.889 0.057 -0.005
H6 -0.716 -1.411 -0.873
H7 -0.717 -1.397 0.892
H8 -1.569 0.977 0.007
H9 2.385 -0.015 -0.003

Atom - Atom Distances (Å)
  C1 C2 O3 O4 O5 H6 H7 H8 H9
C11.51362.38282.40691.40401.09741.09791.91693.2125
C21.51361.34631.21622.40232.13072.12962.26171.8753
O32.38281.34632.26293.59672.62532.63043.59440.9760
O42.40691.21622.26292.69673.10613.09912.09482.3059
O51.40402.40233.59672.69672.06942.07220.97384.2748
H61.09742.13072.62533.10612.06941.76532.68393.5105
H71.09792.12962.63043.09912.07221.76532.67323.5124
H81.91692.26173.59442.09480.97382.68392.67324.0771
H93.21251.87530.97602.30594.27483.51053.51244.0771

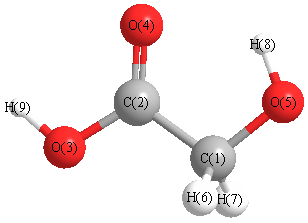
picture of Hydroxyacetic acid state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 112.723 C1 C2 O4 123.330
C1 O5 H8 106.027 C2 C1 O5 110.795
C2 C1 H6 108.326 C2 C1 H7 108.215
C2 O3 H9 106.622 O3 C2 O4 123.947
O5 C1 H6 111.050 O5 C1 H7 111.255
H6 C1 H7 107.054
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.055      
2 C 0.611      
3 O -0.616      
4 O -0.516      
5 O -0.677      
6 H 0.185      
7 H 0.184      
8 H 0.440      
9 H 0.445      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 5.073 -0.145 -0.001
y -0.145 5.094 0.001
z -0.001 0.001 2.938


<r2> (average value of r2) Å2
<r2> 111.219
(<r2>)1/2 10.546