return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COOCH3 (methyl acetate)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-268.109320
Energy at 298.15K-268.116079
HF Energy-267.863569
Nuclear repulsion energy181.166881
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3221 3057 19.33      
2 A' 3220 3056 4.00      
3 A' 3109 2951 28.09      
4 A' 3105 2948 1.22      
5 A' 1830 1737 203.92      
6 A' 1546 1468 7.62      
7 A' 1522 1445 14.84      
8 A' 1510 1433 9.39      
9 A' 1442 1369 46.57      
10 A' 1306 1240 332.69      
11 A' 1232 1169 1.51      
12 A' 1098 1042 72.91      
13 A' 1006 955 2.24      
14 A' 869 825 17.88      
15 A' 649 616 6.55      
16 A' 425 403 6.40      
17 A' 287 273 12.11      
18 A" 3190 3028 21.72      
19 A" 3176 3014 5.52      
20 A" 1535 1457 6.17      
21 A" 1525 1448 7.09      
22 A" 1198 1138 1.18      
23 A" 1090 1034 7.33      
24 A" 607 576 6.64      
25 A" 182 173 6.85      
26 A" 140 133 0.30      
27 A" 57 54 0.50      

Unscaled Zero Point Vibrational Energy (zpe) 20038.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 19020.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.33800 0.13898 0.10225

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.126 1.496 0.000
C2 0.000 0.493 0.000
O3 -1.186 0.754 0.000
O4 0.486 -0.772 0.000
C5 -0.518 -1.801 0.000
H6 0.711 2.503 0.000
H7 1.755 1.351 0.881
H8 1.755 1.351 -0.881
H9 0.030 -2.741 0.000
H10 -1.147 -1.722 0.888
H11 -1.147 -1.722 -0.888

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 H6 H7 H8 H9 H10 H11
C11.50792.42772.35683.68441.08841.09251.09254.37624.03884.0388
C21.50791.21401.35492.35192.13182.14322.14323.23382.64782.6478
O32.42771.21402.26292.64052.58003.12763.12763.69992.63042.6304
O42.35681.35492.26291.43773.28232.62602.62602.02062.08742.0874
C53.68442.35192.64051.43774.47593.98523.98521.08811.09091.0909
H61.08842.13182.58003.28234.47591.78661.78665.28734.69994.6999
H71.09252.14323.12762.62603.98521.78661.76234.52714.22714.5824
H81.09252.14323.12762.62603.98521.78661.76234.52714.58244.2271
H94.37623.23383.69992.02061.08815.28734.52714.52711.79241.7924
H104.03882.64782.63042.08741.09094.69994.22714.58241.79241.7757
H114.03882.64782.63042.08741.09094.69994.58244.22711.79241.7757

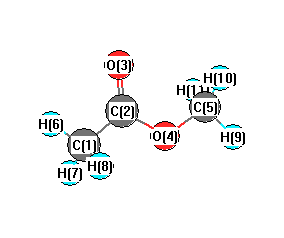
picture of methyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 125.889 C1 C2 O4 110.706
C2 C1 H6 109.320 C2 C1 H7 109.987
C2 C1 H8 109.987 C2 O4 C5 114.705
O3 C2 O4 123.405 O4 C5 H9 105.420
O4 C5 H10 110.531 O4 C5 H11 110.531
H6 C1 H7 110.007 H6 C1 H8 110.007
H7 C1 H8 107.514 H9 C5 H10 110.690
H9 C5 H11 110.690 H10 C5 H11 108.954
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.540      
2 C 0.670      
3 O -0.514      
4 O -0.516      
5 C -0.205      
6 H 0.194      
7 H 0.192      
8 H 0.192      
9 H 0.172      
10 H 0.177      
11 H 0.177      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 5.718 0.238 0.000
y 0.238 6.446 0.000
z 0.000 0.000 4.177


<r2> (average value of r2) Å2
<r2> 120.032
(<r2>)1/2 10.956