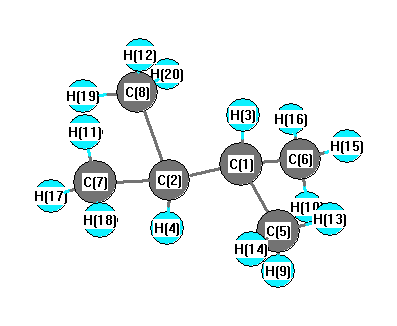
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3146 |
3010 |
0.00 |
|
|
|
| 2 |
Ag |
3120 |
2985 |
0.00 |
|
|
|
| 3 |
Ag |
3052 |
2920 |
0.00 |
|
|
|
| 4 |
Ag |
3003 |
2873 |
0.00 |
|
|
|
| 5 |
Ag |
1551 |
1484 |
0.00 |
|
|
|
| 6 |
Ag |
1529 |
1463 |
0.00 |
|
|
|
| 7 |
Ag |
1444 |
1382 |
0.00 |
|
|
|
| 8 |
Ag |
1395 |
1334 |
0.00 |
|
|
|
| 9 |
Ag |
1224 |
1171 |
0.00 |
|
|
|
| 10 |
Ag |
1205 |
1153 |
0.00 |
|
|
|
| 11 |
Ag |
962 |
920 |
0.00 |
|
|
|
| 12 |
Ag |
766 |
733 |
0.00 |
|
|
|
| 13 |
Ag |
497 |
475 |
0.00 |
|
|
|
| 14 |
Ag |
391 |
374 |
0.00 |
|
|
|
| 15 |
Ag |
241 |
231 |
0.00 |
|
|
|
| 16 |
Au |
3146 |
3009 |
61.01 |
|
|
|
| 17 |
Au |
3115 |
2980 |
0.87 |
|
|
|
| 18 |
Au |
3047 |
2915 |
46.72 |
|
|
|
| 19 |
Au |
1532 |
1465 |
5.25 |
|
|
|
| 20 |
Au |
1507 |
1441 |
1.37 |
|
|
|
| 21 |
Au |
1425 |
1363 |
10.14 |
|
|
|
| 22 |
Au |
1351 |
1292 |
3.59 |
|
|
|
| 23 |
Au |
1100 |
1053 |
1.69 |
|
|
|
| 24 |
Au |
981 |
939 |
0.61 |
|
|
|
| 25 |
Au |
935 |
895 |
2.82 |
|
|
|
| 26 |
Au |
314 |
300 |
0.01 |
|
|
|
| 27 |
Au |
197 |
189 |
0.04 |
|
|
|
| 28 |
Au |
59 |
56 |
0.00 |
|
|
|
| 29 |
Bg |
3133 |
2998 |
0.00 |
|
|
|
| 30 |
Bg |
3115 |
2980 |
0.00 |
|
|
|
| 31 |
Bg |
3046 |
2914 |
0.00 |
|
|
|
| 32 |
Bg |
1516 |
1450 |
0.00 |
|
|
|
| 33 |
Bg |
1511 |
1445 |
0.00 |
|
|
|
| 34 |
Bg |
1420 |
1359 |
0.00 |
|
|
|
| 35 |
Bg |
1365 |
1306 |
0.00 |
|
|
|
| 36 |
Bg |
1188 |
1137 |
0.00 |
|
|
|
| 37 |
Bg |
971 |
929 |
0.00 |
|
|
|
| 38 |
Bg |
927 |
887 |
0.00 |
|
|
|
| 39 |
Bg |
426 |
407 |
0.00 |
|
|
|
| 40 |
Bg |
203 |
194 |
0.00 |
|
|
|
| 41 |
Bu |
3136 |
3001 |
76.44 |
|
|
|
| 42 |
Bu |
3121 |
2986 |
141.63 |
|
|
|
| 43 |
Bu |
3050 |
2918 |
67.90 |
|
|
|
| 44 |
Bu |
3017 |
2887 |
37.71 |
|
|
|
| 45 |
Bu |
1536 |
1469 |
18.44 |
|
|
|
| 46 |
Bu |
1526 |
1460 |
17.49 |
|
|
|
| 47 |
Bu |
1437 |
1374 |
13.92 |
|
|
|
| 48 |
Bu |
1317 |
1260 |
6.87 |
|
|
|
| 49 |
Bu |
1187 |
1136 |
8.41 |
|
|
|
| 50 |
Bu |
1009 |
965 |
7.69 |
|
|
|
| 51 |
Bu |
885 |
846 |
2.68 |
|
|
|
| 52 |
Bu |
405 |
388 |
0.11 |
|
|
|
| 53 |
Bu |
350 |
335 |
0.14 |
|
|
|
| 54 |
Bu |
219 |
210 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41622.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 39820.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.102 |
|
|
0.310 |
| 2 |
C |
-0.102 |
|
|
0.318 |
| 3 |
H |
0.145 |
|
|
-0.019 |
| 4 |
H |
0.145 |
|
|
-0.017 |
| 5 |
C |
-0.513 |
|
|
-0.479 |
| 6 |
C |
-0.513 |
|
|
-0.479 |
| 7 |
C |
-0.513 |
|
|
-0.502 |
| 8 |
C |
-0.513 |
|
|
-0.502 |
| 9 |
H |
0.160 |
|
|
0.106 |
| 10 |
H |
0.160 |
|
|
0.106 |
| 11 |
H |
0.160 |
|
|
0.113 |
| 12 |
H |
0.160 |
|
|
0.113 |
| 13 |
H |
0.163 |
|
|
0.119 |
| 14 |
H |
0.168 |
|
|
0.108 |
| 15 |
H |
0.163 |
|
|
0.119 |
| 16 |
H |
0.168 |
|
|
0.108 |
| 17 |
H |
0.163 |
|
|
0.125 |
| 18 |
H |
0.168 |
|
|
0.114 |
| 19 |
H |
0.163 |
|
|
0.125 |
| 20 |
H |
0.168 |
|
|
0.114 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.003 |
0.002 |
0.000 |
0.003 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.121 |
-0.402 |
0.000 |
| y |
-0.402 |
-41.131 |
0.000 |
| z |
0.000 |
0.000 |
-41.660 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.275 |
-0.402 |
0.000 |
| y |
-0.402 |
0.260 |
0.000 |
| z |
0.000 |
0.000 |
-0.534 |
|
| Polar |
|---|
| 3z2-r2 | -1.069 |
|---|
| x2-y2 | 0.010 |
|---|
| xy | -0.402 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
208.235 |
| (<r2>)1/2 |
14.430 |