return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H14 (Butane, 2,3-dimethyl-)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-236.750517
Energy at 298.15K-236.765670
HF Energy-236.488401
Nuclear repulsion energy262.311837
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3169 3008 0.00      
2 Ag 3141 2982 0.00      
3 Ag 3076 2919 0.00      
4 Ag 3025 2872 0.00      
5 Ag 1576 1496 0.00      
6 Ag 1554 1475 0.00      
7 Ag 1469 1395 0.00      
8 Ag 1415 1343 0.00      
9 Ag 1243 1180 0.00      
10 Ag 1223 1161 0.00      
11 Ag 973 924 0.00      
12 Ag 773 734 0.00      
13 Ag 507 481 0.00      
14 Ag 397 377 0.00      
15 Ag 242 230 0.00      
16 Au 3169 3008 60.46      
17 Au 3135 2976 0.72      
18 Au 3070 2914 39.36      
19 Au 1557 1478 4.47      
20 Au 1534 1456 0.97      
21 Au 1450 1376 6.82      
22 Au 1375 1305 4.39      
23 Au 1109 1052 1.71      
24 Au 996 945 0.41      
25 Au 951 902 2.03      
26 Au 315 299 0.00      
27 Au 201 191 0.02      
28 Au 62 59 0.00      
29 Bg 3154 2994 0.00      
30 Bg 3136 2976 0.00      
31 Bg 3069 2913 0.00      
32 Bg 1542 1464 0.00      
33 Bg 1537 1459 0.00      
34 Bg 1445 1371 0.00      
35 Bg 1390 1320 0.00      
36 Bg 1202 1141 0.00      
37 Bg 984 934 0.00      
38 Bg 945 897 0.00      
39 Bg 433 411 0.00      
40 Bg 205 195 0.00      
41 Bu 3157 2997 79.06      
42 Bu 3143 2983 141.83      
43 Bu 3073 2917 61.24      
44 Bu 3039 2884 39.00      
45 Bu 1560 1480 17.56      
46 Bu 1553 1474 12.12      
47 Bu 1463 1388 10.66      
48 Bu 1339 1271 7.22      
49 Bu 1206 1145 7.49      
50 Bu 1023 971 5.41      
51 Bu 891 846 1.98      
52 Bu 417 395 0.06      
53 Bu 356 338 0.11      
54 Bu 223 212 0.03      

Unscaled Zero Point Vibrational Energy (zpe) 42094.6 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 39956.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.14054 0.09606 0.06247

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.233 0.738 0.000
C2 0.233 -0.738 0.000
H3 -1.333 0.731 0.000
H4 1.333 -0.731 0.000
C5 0.233 1.494 1.251
C6 0.233 1.494 -1.251
C7 -0.233 -1.494 1.251
C8 -0.233 -1.494 -1.251
H9 1.325 1.466 1.337
H10 1.325 1.466 -1.337
H11 -1.325 -1.466 1.337
H12 -1.325 -1.466 -1.337
H13 -0.067 2.545 1.199
H14 -0.187 1.079 2.170
H15 -0.067 2.545 -1.199
H16 -0.187 1.079 -2.170
H17 0.067 -2.545 1.199
H18 0.187 -1.079 2.170
H19 0.067 -2.545 -1.199
H20 0.187 -1.079 -2.170

Atom - Atom Distances (Å)
  C1 C2 H3 H4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20
C11.54711.10042.14681.53471.53472.55872.55872.17852.17852.79942.79942.17542.19742.17542.19743.50792.86133.50792.8613
C21.54712.14681.10042.55872.55871.53471.53472.79942.79942.17852.17853.50792.86133.50792.86132.17542.19742.17542.1974
H31.10042.14683.04062.14522.14522.77962.77963.06543.06542.57132.57132.51682.47892.51682.47893.75893.20873.75893.2087
H42.14681.10043.04062.77962.77962.14522.14522.57132.57133.06543.06543.75893.20873.75893.20872.51682.47892.51682.4789
C51.53472.55872.14522.77962.50293.02433.92571.09622.80973.34624.22961.09431.09212.68313.47224.04312.73274.72744.2815
C61.53472.55872.14522.77962.50293.92573.02432.80971.09624.22963.34622.68313.47221.09431.09214.72744.28154.04312.7327
C72.55871.53472.77962.14523.02433.92572.50293.34624.22961.09622.80974.04312.73274.72744.28151.09431.09212.68313.4722
C82.55871.53472.77962.14523.92573.02432.50294.22963.34622.80971.09624.72744.28154.04312.73272.68313.47221.09431.0921
H92.17852.79943.06542.57131.09622.80973.34624.22962.67403.95244.77191.76721.76953.08793.83894.20612.90984.90954.4803
H102.17852.79943.06542.57132.80971.09624.22963.34622.67404.77193.95243.08793.83891.76721.76954.90954.48034.20612.9098
H112.79942.17852.57133.06543.34624.22961.09622.80973.95244.77192.67404.20612.90984.90954.48031.76721.76953.08793.8389
H122.79942.17852.57133.06544.22963.34622.80971.09624.77193.95242.67404.90954.48034.20612.90983.08793.83891.76721.7695
H132.17543.50792.51683.75891.09432.68314.04314.72741.76723.08794.20614.90951.76272.39793.67635.09223.76085.62864.9550
H142.19742.86132.47893.20871.09213.47222.73274.28151.76953.83892.90984.48031.76273.67634.34043.76082.19044.95504.8618
H152.17543.50792.51683.75892.68311.09434.72744.04313.08791.76724.90954.20612.39793.67631.76275.62864.95505.09223.7608
H162.19742.86132.47893.20873.47221.09214.28152.73273.83891.76954.48032.90983.67634.34041.76274.95504.86183.76082.1904
H173.50792.17543.75892.51684.04314.72741.09432.68314.20614.90951.76723.08795.09223.76085.62864.95501.76272.39793.6763
H182.86132.19743.20872.47892.73274.28151.09213.47222.90984.48031.76953.83893.76082.19044.95504.86181.76273.67634.3404
H193.50792.17543.75892.51684.72744.04312.68311.09434.90954.20613.08791.76725.62864.95505.09223.76082.39793.67631.7627
H202.86132.19743.20872.47894.28152.73273.47221.09214.48032.90983.83891.76954.95504.86183.76082.19043.67634.34041.7627

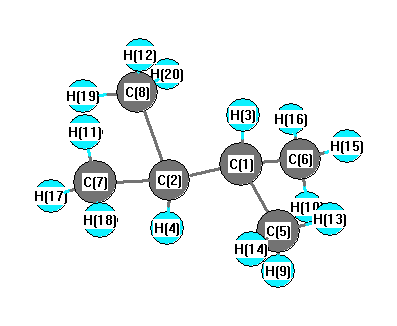
picture of Butane, 2,3-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 H4 107.148 C1 C2 C7 112.251
C1 C2 C8 112.251 C1 C5 H9 110.694
C1 C5 H13 110.557 C1 C5 H14 112.460
C1 C6 H10 110.694 C1 C6 H15 110.557
C1 C6 H16 112.460 C2 C1 H3 107.148
C2 C1 C5 112.251 C2 C1 C6 112.251
C2 C7 H11 110.694 C2 C7 H17 110.557
C2 C7 H18 112.460 C2 C8 H12 110.694
C2 C8 H19 110.557 C2 C8 H20 112.460
H3 C1 C5 107.860 H3 C1 C6 107.860
H4 C2 C7 107.860 H4 C2 C8 107.860
C5 C1 C6 109.263 C7 C2 C8 109.263
H9 C5 H13 107.557 H9 C5 H14 107.924
H10 C6 H15 107.557 H10 C6 H16 107.924
H11 C7 H17 107.557 H11 C7 H18 107.924
H12 C8 H19 107.557 H12 C8 H20 107.924
H13 C5 H14 107.455 H15 C6 H16 107.455
H17 C7 H18 107.455 H19 C8 H20 107.455
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.092      
2 C -0.092      
3 H 0.130      
4 H 0.130      
5 C -0.466      
6 C -0.466      
7 C -0.466      
8 C -0.466      
9 H 0.145      
10 H 0.145      
11 H 0.145      
12 H 0.145      
13 H 0.148      
14 H 0.154      
15 H 0.148      
16 H 0.154      
17 H 0.148      
18 H 0.154      
19 H 0.148      
20 H 0.154      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 8.472 0.169 0.000
y 0.169 10.181 0.000
z 0.000 0.000 9.584


<r2> (average value of r2) Å2
<r2> 208.549
(<r2>)1/2 14.441