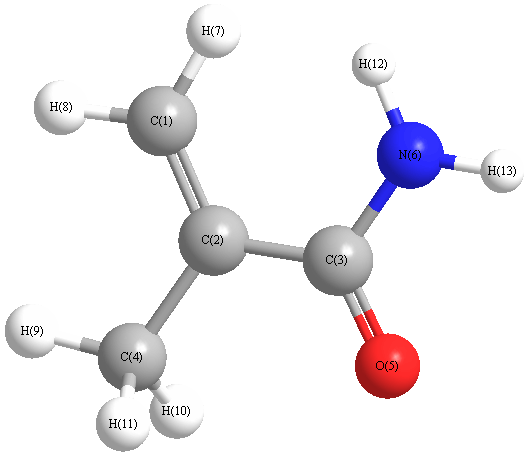
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3732 |
3571 |
32.14 |
|
|
|
| 2 |
A |
3606 |
3450 |
33.12 |
|
|
|
| 3 |
A |
3255 |
3114 |
11.39 |
|
|
|
| 4 |
A |
3173 |
3036 |
6.92 |
|
|
|
| 5 |
A |
3161 |
3025 |
12.20 |
|
|
|
| 6 |
A |
3131 |
2995 |
11.76 |
|
|
|
| 7 |
A |
3063 |
2931 |
15.66 |
|
|
|
| 8 |
A |
1805 |
1727 |
259.11 |
|
|
|
| 9 |
A |
1722 |
1648 |
29.28 |
|
|
|
| 10 |
A |
1640 |
1569 |
110.54 |
|
|
|
| 11 |
A |
1516 |
1451 |
18.50 |
|
|
|
| 12 |
A |
1495 |
1431 |
9.77 |
|
|
|
| 13 |
A |
1464 |
1400 |
15.54 |
|
|
|
| 14 |
A |
1433 |
1371 |
8.53 |
|
|
|
| 15 |
A |
1400 |
1340 |
80.68 |
|
|
|
| 16 |
A |
1249 |
1195 |
47.26 |
|
|
|
| 17 |
A |
1109 |
1061 |
2.26 |
|
|
|
| 18 |
A |
1080 |
1033 |
1.05 |
|
|
|
| 19 |
A |
1029 |
984 |
5.72 |
|
|
|
| 20 |
A |
966 |
924 |
5.25 |
|
|
|
| 21 |
A |
949 |
908 |
39.26 |
|
|
|
| 22 |
A |
825 |
789 |
18.45 |
|
|
|
| 23 |
A |
756 |
723 |
5.29 |
|
|
|
| 24 |
A |
683 |
654 |
14.99 |
|
|
|
| 25 |
A |
580 |
554 |
24.40 |
|
|
|
| 26 |
A |
567 |
543 |
7.07 |
|
|
|
| 27 |
A |
509 |
487 |
5.65 |
|
|
|
| 28 |
A |
394 |
377 |
172.55 |
|
|
|
| 29 |
A |
367 |
351 |
42.17 |
|
|
|
| 30 |
A |
321 |
307 |
4.40 |
|
|
|
| 31 |
A |
262 |
250 |
12.18 |
|
|
|
| 32 |
A |
165 |
158 |
0.26 |
|
|
|
| 33 |
A |
76 |
72 |
3.92 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23741.3 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 22713.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.787 |
|
|
-0.894 |
| 2 |
H |
0.359 |
|
|
0.404 |
| 3 |
H |
0.352 |
|
|
0.374 |
| 4 |
C |
-0.567 |
|
|
-0.356 |
| 5 |
H |
0.187 |
|
|
0.102 |
| 6 |
H |
0.214 |
|
|
0.122 |
| 7 |
H |
0.171 |
|
|
0.119 |
| 8 |
C |
-0.434 |
|
|
-0.494 |
| 9 |
H |
0.172 |
|
|
0.183 |
| 10 |
H |
0.174 |
|
|
0.199 |
| 11 |
C |
0.554 |
|
|
0.645 |
| 12 |
O |
-0.503 |
|
|
-0.510 |
| 13 |
C |
0.108 |
|
|
0.106 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.281 |
2.918 |
1.600 |
3.339 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.323 |
2.915 |
1.606 |
3.343 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.679 |
-4.153 |
-2.605 |
| y |
-4.153 |
-36.636 |
0.037 |
| z |
-2.605 |
0.037 |
-36.488 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.883 |
-4.153 |
-2.605 |
| y |
-4.153 |
-3.052 |
0.037 |
| z |
-2.605 |
0.037 |
-2.831 |
|
| Polar |
|---|
| 3z2-r2 | -5.661 |
|---|
| x2-y2 | 5.957 |
|---|
| xy | -4.153 |
|---|
| xz | -2.605 |
|---|
| yz | 0.037 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
163.325 |
| (<r2>)1/2 |
12.780 |