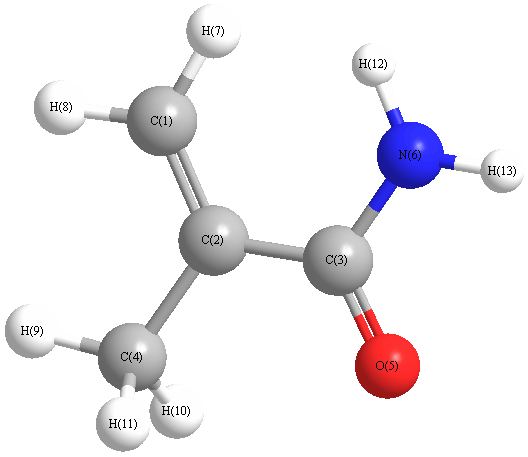
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3618 |
3558 |
22.06 |
|
|
|
| 2 |
A |
3493 |
3435 |
20.17 |
|
|
|
| 3 |
A |
3173 |
3121 |
12.49 |
|
|
|
| 4 |
A |
3092 |
3041 |
7.58 |
|
|
|
| 5 |
A |
3087 |
3036 |
11.25 |
|
|
|
| 6 |
A |
3052 |
3002 |
10.02 |
|
|
|
| 7 |
A |
2988 |
2938 |
14.70 |
|
|
|
| 8 |
A |
1732 |
1703 |
212.15 |
|
|
|
| 9 |
A |
1663 |
1635 |
26.60 |
|
|
|
| 10 |
A |
1585 |
1559 |
93.76 |
|
|
|
| 11 |
A |
1472 |
1448 |
16.97 |
|
|
|
| 12 |
A |
1449 |
1425 |
9.57 |
|
|
|
| 13 |
A |
1418 |
1395 |
13.19 |
|
|
|
| 14 |
A |
1382 |
1360 |
5.94 |
|
|
|
| 15 |
A |
1352 |
1329 |
73.88 |
|
|
|
| 16 |
A |
1211 |
1192 |
43.14 |
|
|
|
| 17 |
A |
1072 |
1054 |
3.47 |
|
|
|
| 18 |
A |
1045 |
1028 |
1.23 |
|
|
|
| 19 |
A |
996 |
979 |
4.82 |
|
|
|
| 20 |
A |
939 |
924 |
3.89 |
|
|
|
| 21 |
A |
901 |
886 |
38.04 |
|
|
|
| 22 |
A |
794 |
781 |
13.46 |
|
|
|
| 23 |
A |
733 |
721 |
5.70 |
|
|
|
| 24 |
A |
656 |
646 |
13.18 |
|
|
|
| 25 |
A |
563 |
554 |
25.45 |
|
|
|
| 26 |
A |
549 |
540 |
3.51 |
|
|
|
| 27 |
A |
494 |
486 |
7.66 |
|
|
|
| 28 |
A |
395 |
389 |
179.08 |
|
|
|
| 29 |
A |
361 |
355 |
15.74 |
|
|
|
| 30 |
A |
311 |
306 |
4.56 |
|
|
|
| 31 |
A |
258 |
253 |
10.61 |
|
|
|
| 32 |
A |
157 |
155 |
0.29 |
|
|
|
| 33 |
A |
75 |
73 |
3.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23033.0 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 22653.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.724 |
|
|
|
| 2 |
H |
0.337 |
|
|
|
| 3 |
H |
0.330 |
|
|
|
| 4 |
C |
-0.551 |
|
|
|
| 5 |
H |
0.177 |
|
|
|
| 6 |
H |
0.202 |
|
|
|
| 7 |
H |
0.160 |
|
|
|
| 8 |
C |
-0.410 |
|
|
|
| 9 |
H |
0.154 |
|
|
|
| 10 |
H |
0.156 |
|
|
|
| 11 |
C |
0.490 |
|
|
|
| 12 |
O |
-0.465 |
|
|
|
| 13 |
C |
0.143 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.203 |
2.751 |
1.561 |
3.169 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.876 |
-4.096 |
-2.549 |
| y |
-4.096 |
-36.736 |
0.173 |
| z |
-2.549 |
0.173 |
-36.594 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.789 |
-4.096 |
-2.549 |
| y |
-4.096 |
-3.001 |
0.173 |
| z |
-2.549 |
0.173 |
-2.788 |
|
| Polar |
|---|
| 3z2-r2 | -5.576 |
|---|
| x2-y2 | 5.861 |
|---|
| xy | -4.096 |
|---|
| xz | -2.549 |
|---|
| yz | 0.173 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.822 |
0.362 |
-0.193 |
| y |
0.362 |
9.711 |
-0.156 |
| z |
-0.193 |
-0.156 |
4.443 |
<r2> (average value of r
2) Å
2
| <r2> |
165.088 |
| (<r2>)1/2 |
12.849 |