return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H12N2 (2-Methyl-1,2-propanediamine)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-268.807765
Energy at 298.15K-268.822568
HF Energy-268.524657
Nuclear repulsion energy268.588759
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3506 3328 0.37      
2 A' 3461 3285 2.53      
3 A' 3171 3010 40.77      
4 A' 3143 2983 52.19      
5 A' 3075 2918 0.67      
6 A' 3052 2897 44.69      
7 A' 1717 1630 18.34      
8 A' 1710 1623 30.08      
9 A' 1558 1479 8.89      
10 A' 1551 1472 4.61      
11 A' 1523 1446 0.55      
12 A' 1462 1388 9.78      
13 A' 1405 1333 8.68      
14 A' 1296 1230 8.02      
15 A' 1252 1188 25.57      
16 A' 1126 1069 7.95      
17 A' 1046 993 1.66      
18 A' 955 907 173.20      
19 A' 924 877 102.04      
20 A' 892 847 152.46      
21 A' 742 705 10.28      
22 A' 518 492 12.34      
23 A' 429 407 11.02      
24 A' 367 348 0.10      
25 A' 265 251 0.53      
26 A' 262 249 5.72      
27 A" 3599 3416 0.01      
28 A" 3552 3372 1.35      
29 A" 3166 3005 0.77      
30 A" 3140 2980 15.15      
31 A" 3098 2940 41.53      
32 A" 3069 2913 37.74      
33 A" 1542 1464 0.61      
34 A" 1533 1455 0.14      
35 A" 1450 1376 7.39      
36 A" 1431 1358 3.99      
37 A" 1379 1309 0.23      
38 A" 1209 1147 0.37      
39 A" 1080 1025 1.12      
40 A" 1024 972 0.78      
41 A" 966 917 0.21      
42 A" 874 830 0.09      
43 A" 458 434 1.36      
44 A" 357 339 21.42      
45 A" 321 304 0.97      
46 A" 294 279 81.33      
47 A" 224 213 0.31      
48 A" 122 115 1.80      

Unscaled Zero Point Vibrational Energy (zpe) 37146.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 35259.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.14720 0.08519 0.08443

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.377 0.221 0.000
H2 1.495 0.819 0.892
H3 1.495 0.819 -0.892
N4 1.949 -0.999 0.000
H5 1.704 -1.544 0.820
H6 1.704 -1.544 -0.820
N7 -0.917 1.590 0.000
H8 -0.554 2.078 -0.823
H9 -0.554 2.078 0.823
C10 1.181 0.264 0.000
C11 -0.917 -0.487 -1.253
C12 -0.917 -0.487 1.253
H13 -0.574 0.029 -2.156
H14 -0.574 0.029 2.156
H15 -0.582 -1.516 1.303
H16 -0.582 -1.516 -1.303
H17 -2.009 -0.469 -1.240
H18 -2.009 -0.469 1.240

Atom - Atom Distances (Å)
  C1 H2 H3 N4 H5 H6 N7 H8 H9 C10 C11 C12 H13 H14 H15 H16 H17 H18
C12.15752.15752.62652.84912.84911.47162.03862.03861.55871.53751.53752.17312.17312.18162.18162.16252.1625
H22.15751.78392.07512.37312.92522.68462.95362.40571.09603.48192.76623.76692.54933.15183.81894.29833.7487
H32.15751.78392.07512.92522.37312.68462.40572.95361.09602.76623.48192.54933.76693.81893.15183.74874.2983
N42.62652.07512.07511.01461.01463.86244.05094.05091.47813.17033.17033.47403.47402.89372.89374.18164.1816
H52.84912.37312.92521.01461.63974.16714.57334.26812.05303.50542.85974.06403.07362.33683.12014.38013.8885
H62.84912.92522.37311.01461.63974.16714.26814.57332.05302.85973.50543.07364.06403.12012.33683.88854.3801
N71.47162.68462.68463.86244.16714.16711.02331.02332.48222.42532.42532.68352.68353.38523.38522.63912.6391
H82.03862.95362.40574.05094.57334.26811.02331.64592.64172.62543.31942.44423.61524.17603.62612.96193.5852
H92.03862.40572.95364.05094.26814.57331.02331.64592.64173.31942.62543.61522.44423.62614.17603.58522.9619
C101.55871.09601.09601.47812.05302.05302.48222.64172.64172.55712.55712.78972.78972.82462.82463.50023.5002
C111.53753.48192.76623.17033.50542.85972.42532.62543.31942.55712.50671.09453.46482.77661.08411.09182.7216
C121.53752.76623.48193.17032.85973.50542.42533.31942.62542.55712.50673.46481.09451.08412.77662.72161.0918
H132.17313.76692.54933.47404.06403.07362.68352.44423.61522.78971.09453.46484.31133.78851.76471.77373.7196
H142.17312.54933.76693.47403.07364.06402.68353.61522.44422.78973.46481.09454.31131.76473.78853.71961.7737
H152.18163.15183.81892.89372.33683.12013.38524.17603.62612.82462.77661.08413.78851.76472.60683.09871.7716
H162.18163.81893.15182.89373.12012.33683.38523.62614.17602.82461.08412.77661.76473.78852.60681.77163.0987
H172.16254.29833.74874.18164.38013.88852.63912.96193.58523.50021.09182.72161.77373.71963.09871.77162.4795
H182.16253.74874.29834.18163.88854.38012.63913.58522.96193.50022.72161.09183.71961.77371.77163.09872.4795

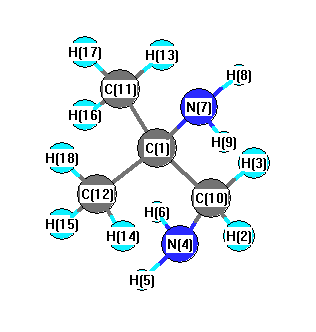
picture of 2-Methyl-1,2-propanediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N7 H8 108.250 C1 N7 H9 108.250
C1 C10 H2 107.441 C1 C10 H3 107.441
C1 C10 N4 119.714 C1 C11 H13 110.176
C1 C11 H16 111.470 C1 C11 H17 109.495
C1 C12 H14 110.176 C1 C12 H15 111.470
C1 C12 H18 109.495 H2 C10 H3 108.937
H2 C10 N4 106.491 H3 C10 N4 106.491
H5 N4 H6 107.810 H5 N4 C10 109.486
H6 N4 C10 109.486 N7 C1 C10 109.962
N7 C1 C11 107.393 N7 C1 C12 107.393
H8 N7 H9 107.071 C10 C1 C11 111.360
C10 C1 C12 111.360 C11 C1 C12 109.213
H13 C11 H16 108.191 H13 C11 H17 108.436
H14 C12 H15 108.191 H14 C12 H18 108.436
H15 C12 H18 109.008 H16 C11 H17 109.008
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.148      
2 H 0.139      
3 H 0.139      
4 N -0.766      
5 H 0.314      
6 H 0.314      
7 N -0.755      
8 H 0.306      
9 H 0.306      
10 C -0.136      
11 C -0.456      
12 C -0.456      
13 H 0.142      
14 H 0.142      
15 H 0.138      
16 H 0.138      
17 H 0.172      
18 H 0.172      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 7.927 -0.289 0.000
y -0.289 8.584 0.000
z 0.000 0.000 8.416


<r2> (average value of r2) Å2
<r2> 183.503
(<r2>)1/2 13.546