Jump to
S1C2
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -216.423318 |
| Energy at 298.15K | |
| HF Energy | -215.917178 |
| Nuclear repulsion energy | 116.889868 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3375 |
3117 |
6.60 |
|
|
|
| 2 |
A' |
3290 |
3039 |
5.72 |
|
|
|
| 3 |
A' |
3268 |
3019 |
10.12 |
|
|
|
| 4 |
A' |
3155 |
2914 |
38.74 |
|
|
|
| 5 |
A' |
1827 |
1687 |
1.73 |
|
|
|
| 6 |
A' |
1605 |
1483 |
0.96 |
|
|
|
| 7 |
A' |
1529 |
1412 |
11.39 |
|
|
|
| 8 |
A' |
1506 |
1391 |
15.15 |
|
|
|
| 9 |
A' |
1383 |
1277 |
0.33 |
|
|
|
| 10 |
A' |
1205 |
1113 |
50.90 |
|
|
|
| 11 |
A' |
1072 |
990 |
33.51 |
|
|
|
| 12 |
A' |
963 |
889 |
2.52 |
|
|
|
| 13 |
A' |
633 |
585 |
6.46 |
|
|
|
| 14 |
A' |
287 |
265 |
3.06 |
|
|
|
| 15 |
A" |
3198 |
2954 |
40.36 |
|
|
|
| 16 |
A" |
1325 |
1224 |
0.08 |
|
|
|
| 17 |
A" |
1116 |
1031 |
15.41 |
|
|
|
| 18 |
A" |
1069 |
988 |
12.05 |
|
|
|
| 19 |
A" |
1008 |
931 |
43.00 |
|
|
|
| 20 |
A" |
589 |
544 |
8.14 |
|
|
|
| 21 |
A" |
172 |
159 |
3.51 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16787.6 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 15506.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.946 |
-0.205 |
0.000 |
| C2 |
0.000 |
0.953 |
0.000 |
| C3 |
1.319 |
0.836 |
0.000 |
| F4 |
-0.261 |
-1.405 |
0.000 |
| H5 |
1.958 |
1.706 |
0.000 |
| H6 |
1.798 |
-0.131 |
0.000 |
| H7 |
-0.471 |
1.930 |
0.000 |
| H8 |
-1.586 |
-0.184 |
0.883 |
| H9 |
-1.586 |
-0.184 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4949 | 2.4921 | 1.3819 | 3.4764 | 2.7442 | 2.1866 | 1.0914 | 1.0914 |
C2 | 1.4949 | | 1.3240 | 2.3728 | 2.0983 | 2.0993 | 1.0844 | 2.1423 | 2.1423 | C3 | 2.4921 | 1.3240 | | 2.7422 | 1.0801 | 1.0793 | 2.0977 | 3.2031 | 3.2031 | F4 | 1.3819 | 2.3728 | 2.7422 | | 3.8222 | 2.4211 | 3.3417 | 2.0070 | 2.0070 | H5 | 3.4764 | 2.0983 | 1.0801 | 3.8222 | | 1.8446 | 2.4399 | 4.1132 | 4.1132 | H6 | 2.7442 | 2.0993 | 1.0793 | 2.4211 | 1.8446 | | 3.0652 | 3.4977 | 3.4977 | H7 | 2.1866 | 1.0844 | 2.0977 | 3.3417 | 2.4399 | 3.0652 | | 2.5478 | 2.5478 | H8 | 1.0914 | 2.1423 | 3.2031 | 2.0070 | 4.1132 | 3.4977 | 2.5478 | | 1.7667 | H9 | 1.0914 | 2.1423 | 3.2031 | 2.0070 | 4.1132 | 3.4977 | 2.5478 | 1.7667 | |
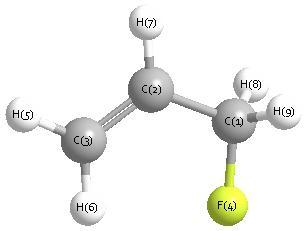 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.167 |
|
C1 |
C2 |
H7 |
115.005 |
| C2 |
C1 |
F4 |
111.075 |
|
C2 |
C1 |
H8 |
110.889 |
| C2 |
C1 |
H9 |
110.889 |
|
C2 |
C3 |
H5 |
121.244 |
| C2 |
C3 |
H6 |
121.407 |
|
C3 |
C2 |
H7 |
120.828 |
| F4 |
C1 |
H8 |
107.897 |
|
F4 |
C1 |
H9 |
107.897 |
| H5 |
C3 |
H6 |
117.349 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CID/6-31G*
| | hartrees |
|---|
| Energy at 0K | -216.422273 |
| Energy at 298.15K | |
| HF Energy | -215.916457 |
| Nuclear repulsion energy | 114.493381 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CID/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3353 |
3097 |
13.31 |
|
|
|
| 2 |
A |
3287 |
3036 |
5.13 |
|
|
|
| 3 |
A |
3265 |
3016 |
9.91 |
|
|
|
| 4 |
A |
3220 |
2974 |
32.27 |
|
|
|
| 5 |
A |
3165 |
2924 |
36.82 |
|
|
|
| 6 |
A |
1824 |
1685 |
0.37 |
|
|
|
| 7 |
A |
1608 |
1485 |
0.61 |
|
|
|
| 8 |
A |
1546 |
1428 |
24.03 |
|
|
|
| 9 |
A |
1476 |
1363 |
21.55 |
|
|
|
| 10 |
A |
1380 |
1275 |
0.06 |
|
|
|
| 11 |
A |
1330 |
1229 |
2.85 |
|
|
|
| 12 |
A |
1245 |
1150 |
2.43 |
|
|
|
| 13 |
A |
1140 |
1053 |
114.15 |
|
|
|
| 14 |
A |
1082 |
1000 |
24.85 |
|
|
|
| 15 |
A |
1044 |
965 |
2.59 |
|
|
|
| 16 |
A |
1016 |
939 |
45.43 |
|
|
|
| 17 |
A |
976 |
902 |
3.68 |
|
|
|
| 18 |
A |
679 |
627 |
6.94 |
|
|
|
| 19 |
A |
457 |
422 |
2.39 |
|
|
|
| 20 |
A |
341 |
315 |
7.13 |
|
|
|
| 21 |
A |
116 |
107 |
1.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16774.4 cm
-1
Scaled (by 0.9237) Zero Point Vibrational Energy (zpe) 15494.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CID/6-31G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.604 |
0.448 |
0.290 |
| C2 |
0.640 |
-0.375 |
0.237 |
| C3 |
1.804 |
0.082 |
-0.202 |
| F4 |
-1.646 |
-0.238 |
-0.319 |
| H5 |
2.694 |
-0.532 |
-0.197 |
| H6 |
1.916 |
1.089 |
-0.582 |
| H7 |
0.545 |
-1.389 |
0.605 |
| H8 |
-0.906 |
0.643 |
1.320 |
| H9 |
-0.473 |
1.400 |
-0.224 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4927 | 2.4853 | 1.3882 | 3.4750 | 2.7426 | 2.1903 | 1.0907 | 1.0894 |
C2 | 1.4927 | | 1.3250 | 2.3561 | 2.1052 | 2.1071 | 1.0835 | 2.1442 | 2.1446 | C3 | 2.4853 | 1.3250 | | 3.4665 | 1.0805 | 1.0825 | 2.0971 | 3.1583 | 2.6314 | F4 | 1.3882 | 2.3561 | 3.4665 | | 4.3510 | 3.8098 | 2.6419 | 2.0027 | 2.0169 | H5 | 3.4750 | 2.1052 | 1.0805 | 4.3510 | | 1.8382 | 2.4485 | 4.0791 | 3.7095 | H6 | 2.7426 | 2.1071 | 1.0825 | 3.8098 | 1.8382 | | 3.0704 | 3.4320 | 2.4355 | H7 | 2.1903 | 1.0835 | 2.0971 | 2.6419 | 2.4485 | 3.0704 | | 2.5971 | 3.0827 | H8 | 1.0907 | 2.1442 | 3.1583 | 2.0027 | 4.0791 | 3.4320 | 2.5971 | | 1.7734 | H9 | 1.0894 | 2.1446 | 2.6314 | 2.0169 | 3.7095 | 2.4355 | 3.0827 | 1.7734 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.673 |
|
C1 |
C2 |
H7 |
115.556 |
| C2 |
C1 |
F4 |
109.684 |
|
C2 |
C1 |
H8 |
111.245 |
| C2 |
C1 |
H9 |
111.360 |
|
C2 |
C3 |
H5 |
121.795 |
| C2 |
C3 |
H6 |
121.819 |
|
C3 |
C2 |
H7 |
120.759 |
| F4 |
C1 |
H8 |
107.170 |
|
F4 |
C1 |
H9 |
108.385 |
| H5 |
C3 |
H6 |
116.386 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
