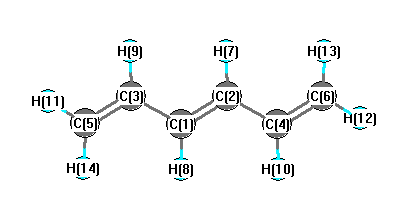
Vibrational Frequencies calculated at B3PW91/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3262 |
3120 |
0.00 |
|
|
|
| 2 |
Ag |
3175 |
3038 |
0.00 |
|
|
|
| 3 |
Ag |
3164 |
3027 |
0.00 |
|
|
|
| 4 |
Ag |
3157 |
3020 |
0.00 |
|
|
|
| 5 |
Ag |
1721 |
1646 |
0.00 |
|
|
|
| 6 |
Ag |
1662 |
1590 |
0.00 |
|
|
|
| 7 |
Ag |
1451 |
1388 |
0.00 |
|
|
|
| 8 |
Ag |
1340 |
1282 |
0.00 |
|
|
|
| 9 |
Ag |
1325 |
1268 |
0.00 |
|
|
|
| 10 |
Ag |
1234 |
1181 |
0.00 |
|
|
|
| 11 |
Ag |
952 |
911 |
0.00 |
|
|
|
| 12 |
Ag |
445 |
426 |
0.00 |
|
|
|
| 13 |
Ag |
348 |
333 |
0.00 |
|
|
|
| 14 |
Au |
1062 |
1016 |
60.43 |
|
|
|
| 15 |
Au |
971 |
929 |
5.80 |
|
|
|
| 16 |
Au |
921 |
881 |
84.58 |
|
|
|
| 17 |
Au |
710 |
680 |
9.62 |
|
|
|
| 18 |
Au |
254 |
243 |
1.52 |
|
|
|
| 19 |
Au |
99 |
95 |
0.33 |
|
|
|
| 20 |
Bg |
1028 |
984 |
0.00 |
|
|
|
| 21 |
Bg |
925 |
885 |
0.00 |
|
|
|
| 22 |
Bg |
896 |
858 |
0.00 |
|
|
|
| 23 |
Bg |
615 |
588 |
0.00 |
|
|
|
| 24 |
Bg |
225 |
215 |
0.00 |
|
|
|
| 25 |
Bu |
3262 |
3120 |
27.49 |
|
|
|
| 26 |
Bu |
3175 |
3037 |
11.58 |
|
|
|
| 27 |
Bu |
3169 |
3031 |
59.95 |
|
|
|
| 28 |
Bu |
3156 |
3020 |
0.05 |
|
|
|
| 29 |
Bu |
1715 |
1641 |
17.55 |
|
|
|
| 30 |
Bu |
1483 |
1419 |
3.01 |
|
|
|
| 31 |
Bu |
1337 |
1279 |
3.81 |
|
|
|
| 32 |
Bu |
1291 |
1235 |
2.40 |
|
|
|
| 33 |
Bu |
1174 |
1123 |
6.05 |
|
|
|
| 34 |
Bu |
987 |
944 |
3.84 |
|
|
|
| 35 |
Bu |
539 |
516 |
5.54 |
|
|
|
| 36 |
Bu |
146 |
139 |
1.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26186.5 cm
-1
Scaled (by 0.9567) Zero Point Vibrational Energy (zpe) 25052.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.153 |
|
|
-0.152 |
| 2 |
C |
-0.153 |
|
|
-0.189 |
| 3 |
C |
-0.094 |
|
|
-0.046 |
| 4 |
C |
-0.094 |
|
|
-0.022 |
| 5 |
C |
-0.403 |
|
|
-0.419 |
| 6 |
C |
-0.403 |
|
|
-0.429 |
| 7 |
H |
0.155 |
|
|
0.162 |
| 8 |
H |
0.155 |
|
|
0.155 |
| 9 |
H |
0.158 |
|
|
0.124 |
| 10 |
H |
0.158 |
|
|
0.118 |
| 11 |
H |
0.170 |
|
|
0.171 |
| 12 |
H |
0.170 |
|
|
0.173 |
| 13 |
H |
0.166 |
|
|
0.177 |
| 14 |
H |
0.166 |
|
|
0.175 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.008 |
0.003 |
0.000 |
0.008 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.704 |
-0.197 |
0.000 |
| y |
-0.197 |
-33.011 |
0.000 |
| z |
0.000 |
0.000 |
-40.963 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.283 |
-0.197 |
0.000 |
| y |
-0.197 |
3.822 |
0.000 |
| z |
0.000 |
0.000 |
-8.105 |
|
| Polar |
|---|
| 3z2-r2 | -16.210 |
|---|
| x2-y2 | 0.308 |
|---|
| xy | -0.197 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
256.359 |
| (<r2>)1/2 |
16.011 |