return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H8 ((E)-hexa-1,3,5-triene)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1Ag
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-233.106383
Energy at 298.15K-233.113487
HF Energy-232.853014
Nuclear repulsion energy193.722123
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3281 3114 0.00      
2 Ag 3195 3033 0.00      
3 Ag 3183 3021 0.00      
4 Ag 3176 3015 0.00      
5 Ag 1722 1635 0.00      
6 Ag 1659 1574 0.00      
7 Ag 1466 1392 0.00      
8 Ag 1355 1286 0.00      
9 Ag 1338 1270 0.00      
10 Ag 1243 1180 0.00      
11 Ag 962 913 0.00      
12 Ag 448 425 0.00      
13 Ag 352 334 0.00      
14 Au 1063 1009 57.51      
15 Au 976 926 2.90      
16 Au 919 873 77.79      
17 Au 712 676 7.91      
18 Au 255 242 1.45      
19 Au 99 94 0.26      
20 Bg 1032 980 0.00      
21 Bg 921 874 0.00      
22 Bg 888 843 0.00      
23 Bg 619 587 0.00      
24 Bg 218 207 0.00      
25 Bu 3281 3114 28.96      
26 Bu 3195 3033 10.71      
27 Bu 3187 3026 56.78      
28 Bu 3175 3014 0.03      
29 Bu 1712 1625 13.36      
30 Bu 1501 1425 2.03      
31 Bu 1349 1280 3.62      
32 Bu 1307 1240 2.37      
33 Bu 1178 1118 5.47      
34 Bu 995 944 2.59      
35 Bu 545 518 4.84      
36 Bu 148 141 1.91      

Unscaled Zero Point Vibrational Energy (zpe) 26326.5 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 24989.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
0.88627 0.04447 0.04235

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.185 0.638 0.000
C2 0.185 -0.638 0.000
C3 0.185 1.790 0.000
C4 -0.185 -1.790 0.000
C5 0.415 2.956 0.000
C6 -0.415 -2.956 0.000
H7 1.175 -0.869 0.000
H8 -1.175 0.869 0.000
H9 1.711 1.385 0.000
H10 -1.711 -1.385 0.000
H11 1.125 3.671 0.000
H12 -1.125 -3.671 0.000
H13 0.564 -3.237 0.000
H14 -0.564 3.237 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.32901.20962.42862.39453.60222.03001.01692.03772.53483.30354.41103.94722.6264
C21.32902.42861.20963.60222.39451.01692.03002.53482.03774.41103.30352.62643.9472
C31.20962.42863.59961.18884.78462.83781.64221.57983.69822.10295.61635.04181.6293
C42.42861.20963.59964.78461.18881.64222.83783.69821.57985.61632.10291.62935.0418
C52.39453.60221.18884.78465.97113.90032.62422.03704.83441.00706.80446.19541.0193
C63.60222.39454.78461.18885.97112.62423.90034.83442.03706.80441.00701.01936.1954
H72.03001.01692.83781.64223.90032.62422.92292.31702.93214.54063.62502.44554.4595
H81.01692.03001.64222.83782.62423.90032.92292.93212.31703.62504.54064.45952.4455
H92.03772.53481.57983.69822.03704.83442.31702.93214.40322.36045.79744.76232.9344
H102.53482.03773.69821.57984.83442.03702.93212.31704.40325.79742.36042.93444.7623
H113.30354.41102.10295.61631.00706.80444.54063.62502.36045.79747.67946.93111.7442
H124.41103.30355.61632.10296.80441.00703.62504.54065.79742.36047.67941.74426.9311
H133.94722.62645.04181.62936.19541.01932.44554.45954.76232.93446.93111.74426.5720
H142.62643.94721.62935.04181.01936.19544.45952.44552.93444.76231.74426.93116.5720

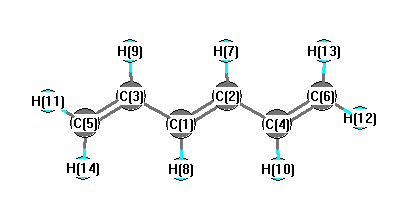
picture of (E)-hexa-1,3,5-triene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 146.107 C1 C2 H7 119.243
C1 C3 C5 173.433 C1 C3 H9 92.899
C2 C1 C3 146.107 C2 C1 H8 119.243
C2 C4 C6 173.433 C2 C4 H10 92.899
C3 C1 H8 94.650 C3 C5 H11 146.418
C3 C5 H14 94.782 C4 C2 H7 94.650
C4 C6 H12 146.418 C4 C6 H13 94.782
C5 C3 H9 93.668 C6 C4 H10 93.668
H11 C5 H14 118.799 H12 C6 H13 118.799
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.150      
2 C -0.150      
3 C -0.094      
4 C -0.094      
5 C -0.380      
6 C -0.380      
7 H 0.152      
8 H 0.152      
9 H 0.154      
10 H 0.154      
11 H 0.162      
12 H 0.162      
13 H 0.157      
14 H 0.157      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 9.550 4.158 0.000
y 4.158 20.471 0.000
z 0.000 0.000 3.276


<r2> (average value of r2) Å2
<r2> 256.912
(<r2>)1/2 16.028