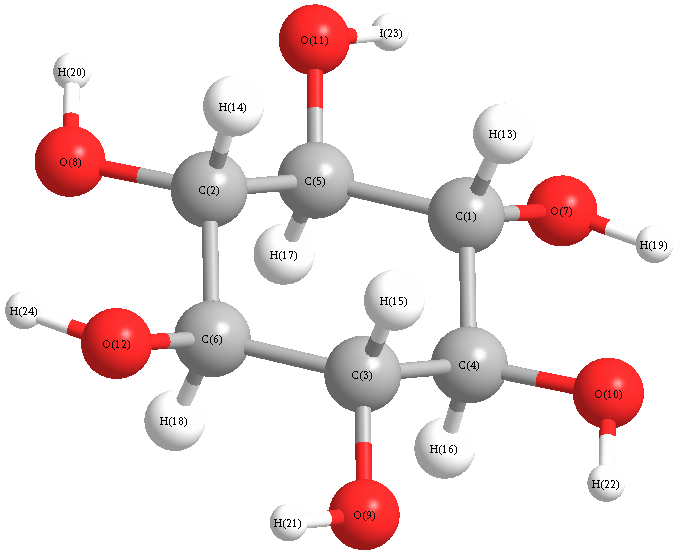
Vibrational Frequencies calculated at PBEPBEultrafine/6-31G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3614 |
3554 |
0.00 |
|
|
|
| 2 |
A1g |
2925 |
2877 |
0.00 |
|
|
|
| 3 |
A1g |
1416 |
1392 |
0.00 |
|
|
|
| 4 |
A1g |
1322 |
1300 |
0.00 |
|
|
|
| 5 |
A1g |
1156 |
1137 |
0.00 |
|
|
|
| 6 |
A1g |
983 |
966 |
0.00 |
|
|
|
| 7 |
A1g |
433 |
425 |
0.00 |
|
|
|
| 8 |
A1g |
120 |
118 |
0.00 |
|
|
|
| 9 |
A1u |
1288 |
1267 |
0.00 |
|
|
|
| 10 |
A1u |
975 |
959 |
0.00 |
|
|
|
| 11 |
A1u |
310 |
305 |
0.00 |
|
|
|
| 12 |
A1u |
7i |
7i |
0.00 |
|
|
|
| 13 |
A2g |
1141 |
1122 |
0.00 |
|
|
|
| 14 |
A2g |
625 |
615 |
0.00 |
|
|
|
| 15 |
A2g |
87 |
86 |
0.00 |
|
|
|
| 16 |
A2u |
3612 |
3552 |
33.20 |
|
|
|
| 17 |
A2u |
2955 |
2906 |
190.57 |
|
|
|
| 18 |
A2u |
1419 |
1396 |
28.29 |
|
|
|
| 19 |
A2u |
1259 |
1239 |
0.85 |
|
|
|
| 20 |
A2u |
1103 |
1085 |
23.22 |
|
|
|
| 21 |
A2u |
559 |
550 |
0.38 |
|
|
|
| 22 |
A2u |
224 |
220 |
20.78 |
|
|
|
| 23 |
Eg |
3612 |
3552 |
0.00 |
|
|
|
| 23 |
Eg |
3612 |
3552 |
0.00 |
|
|
|
| 24 |
Eg |
2931 |
2882 |
0.00 |
|
|
|
| 24 |
Eg |
2931 |
2882 |
0.00 |
|
|
|
| 25 |
Eg |
1403 |
1380 |
0.00 |
|
|
|
| 25 |
Eg |
1403 |
1380 |
0.00 |
|
|
|
| 26 |
Eg |
1343 |
1320 |
0.00 |
|
|
|
| 26 |
Eg |
1343 |
1320 |
0.00 |
|
|
|
| 27 |
Eg |
1190 |
1170 |
0.00 |
|
|
|
| 27 |
Eg |
1190 |
1170 |
0.00 |
|
|
|
| 28 |
Eg |
1105 |
1086 |
0.00 |
|
|
|
| 28 |
Eg |
1105 |
1086 |
0.00 |
|
|
|
| 29 |
Eg |
984 |
968 |
0.00 |
|
|
|
| 29 |
Eg |
984 |
968 |
0.00 |
|
|
|
| 30 |
Eg |
406 |
399 |
0.00 |
|
|
|
| 30 |
Eg |
406 |
399 |
0.00 |
|
|
|
| 31 |
Eg |
364 |
358 |
0.00 |
|
|
|
| 31 |
Eg |
364 |
358 |
0.00 |
|
|
|
| 32 |
Eg |
286 |
281 |
0.00 |
|
|
|
| 32 |
Eg |
286 |
281 |
0.00 |
|
|
|
| 33 |
Eg |
79 |
78 |
0.00 |
|
|
|
| 33 |
Eg |
79 |
78 |
0.00 |
|
|
|
| 34 |
Eu |
3613 |
3553 |
13.78 |
|
|
|
| 34 |
Eu |
3613 |
3553 |
13.78 |
|
|
|
| 35 |
Eu |
2918 |
2870 |
0.34 |
|
|
|
| 35 |
Eu |
2918 |
2870 |
0.34 |
|
|
|
| 36 |
Eu |
1398 |
1375 |
140.29 |
|
|
|
| 36 |
Eu |
1398 |
1375 |
140.30 |
|
|
|
| 37 |
Eu |
1337 |
1315 |
41.22 |
|
|
|
| 37 |
Eu |
1337 |
1315 |
41.22 |
|
|
|
| 38 |
Eu |
1167 |
1148 |
100.21 |
|
|
|
| 38 |
Eu |
1167 |
1148 |
100.21 |
|
|
|
| 39 |
Eu |
1118 |
1099 |
119.94 |
|
|
|
| 39 |
Eu |
1118 |
1099 |
119.95 |
|
|
|
| 40 |
Eu |
963 |
947 |
221.44 |
|
|
|
| 40 |
Eu |
963 |
947 |
221.43 |
|
|
|
| 41 |
Eu |
619 |
609 |
7.34 |
|
|
|
| 41 |
Eu |
619 |
609 |
7.34 |
|
|
|
| 42 |
Eu |
306 |
301 |
100.84 |
|
|
|
| 42 |
Eu |
306 |
301 |
100.83 |
|
|
|
| 43 |
Eu |
170 |
167 |
243.99 |
|
|
|
| 43 |
Eu |
170 |
167 |
244.00 |
|
|
|
| 44 |
Eu |
73 |
71 |
9.63 |
|
|
|
| 44 |
Eu |
73 |
71 |
9.63 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41139.2 cm
-1
Scaled (by 0.9835) Zero Point Vibrational Energy (zpe) 40460.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBEultrafine/6-31G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.052 |
|
|
|
| 2 |
C |
0.052 |
|
|
|
| 3 |
C |
0.052 |
|
|
|
| 4 |
C |
0.052 |
|
|
|
| 5 |
C |
0.052 |
|
|
|
| 6 |
C |
0.052 |
|
|
|
| 7 |
O |
-0.561 |
|
|
|
| 8 |
O |
-0.561 |
|
|
|
| 9 |
O |
-0.561 |
|
|
|
| 10 |
O |
-0.561 |
|
|
|
| 11 |
O |
-0.561 |
|
|
|
| 12 |
O |
-0.561 |
|
|
|
| 13 |
H |
0.124 |
|
|
|
| 14 |
H |
0.124 |
|
|
|
| 15 |
H |
0.124 |
|
|
|
| 16 |
H |
0.124 |
|
|
|
| 17 |
H |
0.124 |
|
|
|
| 18 |
H |
0.124 |
|
|
|
| 19 |
H |
0.385 |
|
|
|
| 20 |
H |
0.385 |
|
|
|
| 21 |
H |
0.385 |
|
|
|
| 22 |
H |
0.385 |
|
|
|
| 23 |
H |
0.385 |
|
|
|
| 24 |
H |
0.385 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-82.627 |
0.000 |
0.000 |
| y |
0.000 |
-82.627 |
0.000 |
| z |
0.000 |
0.000 |
-56.363 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-13.132 |
0.000 |
0.000 |
| y |
0.000 |
-13.132 |
0.000 |
| z |
0.000 |
0.000 |
26.263 |
|
| Polar |
|---|
| 3z2-r2 | 52.526 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.008 |
0.000 |
0.000 |
| y |
0.000 |
13.008 |
0.000 |
| z |
0.000 |
0.000 |
11.769 |
<r2> (average value of r
2) Å
2
| <r2> |
598.353 |
| (<r2>)1/2 |
24.461 |