return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O (allenol)

using model chemistry: B2PLYP/6-31G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP/6-31G*
 hartrees
Energy at 0K-191.658408
Energy at 298.15K-191.661889
HF Energy-191.473535
Nuclear repulsion energy101.798175
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3750 3560 30.52      
2 A' 3248 3083 5.65      
3 A' 3154 2993 18.81      
4 A' 2075 1970 39.06      
5 A' 1541 1463 39.68      
6 A' 1452 1378 67.81      
7 A' 1311 1245 2.11      
8 A' 1218 1156 130.89      
9 A' 1004 953 113.20      
10 A' 931 883 35.81      
11 A' 627 596 18.63      
12 A' 214 203 1.02      
13 A" 3223 3059 10.64      
14 A" 1062 1008 0.71      
15 A" 910 863 24.66      
16 A" 634 602 0.79      
17 A" 440 418 128.50      
18 A" 270 256 2.20      

Unscaled Zero Point Vibrational Energy (zpe) 13531.8 cm-1
Scaled (by 0.9492) Zero Point Vibrational Energy (zpe) 12844.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP/6-31G*
ABC
1.45211 0.14466 0.13518

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP/6-31G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.658 -0.491 0.000
C2 0.000 0.648 0.000
C3 -0.655 1.786 0.000
O4 0.121 -1.753 0.000
H5 1.743 -0.529 0.000
H6 -0.944 2.284 0.924
H7 -0.944 2.284 -0.924
H8 -0.845 -1.675 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8
C11.31512.62801.37191.08513.33463.33461.9137
C21.31511.31292.40382.10322.10282.10282.4718
C32.62801.31293.62273.33301.08881.08883.4661
O41.37192.40383.62272.03154.27604.27600.9687
H51.08512.10323.33302.03153.99853.99852.8297
H63.33462.10281.08884.27603.99851.84794.0668
H73.33462.10281.08884.27603.99851.84794.0668
H81.91372.47183.46610.96872.82974.06684.0668

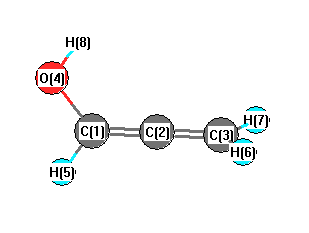
picture of allenol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.878 C1 O4 H8 108.460
C2 C1 O4 126.898 C2 C1 H5 122.091
C2 C3 H6 121.942 C2 C3 H7 121.942
O4 C1 H5 111.011 H6 C3 H7 116.115
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G* Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.007      
2 C 0.162      
3 C -0.489      
4 O -0.636      
5 H 0.188      
6 H 0.178      
7 H 0.178      
8 H 0.425      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 4.429 -2.188 0.000
y -2.188 8.080 0.000
z 0.000 0.000 2.853


<r2> (average value of r2) Å2
<r2> 88.042
(<r2>)1/2 9.383